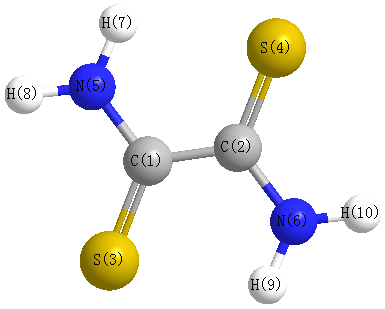
All results from a given calculation for C2H4N2S2 (Ethanedithioamide)
using model chemistry: QCISD(T)=FULL/aug-cc-pVDZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
C2H |
1AG |
Energy calculated at QCISD(T)=FULL/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -983.086451 |
| Energy at 298.15K | |
| HF Energy | -982.064179 |
| Nuclear repulsion energy | 334.440605 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)=FULL/aug-cc-pVDZ
Geometric Data calculated at QCISD(T)=FULL/aug-cc-pVDZ
Point Group is C2h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.048 |
0.768 |
0.000 |
| C2 |
0.048 |
-0.768 |
0.000 |
| S3 |
1.312 |
1.754 |
0.000 |
| S4 |
-1.312 |
-1.754 |
0.000 |
| N5 |
-1.312 |
1.223 |
0.000 |
| N6 |
1.312 |
-1.223 |
0.000 |
| H7 |
-2.074 |
0.544 |
0.000 |
| H8 |
-1.482 |
2.222 |
0.000 |
| H9 |
2.074 |
-0.544 |
0.000 |
| H10 |
1.482 |
-2.222 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
S3 |
S4 |
N5 |
N6 |
H7 |
H8 |
H9 |
H10 |
| C1 | | 1.5392 | 1.6803 | 2.8210 | 1.3434 | 2.4119 | 2.0381 | 2.0416 | 2.4951 | 3.3587 |
C2 | 1.5392 | | 2.8210 | 1.6803 | 2.4119 | 1.3434 | 2.4951 | 3.3587 | 2.0381 | 2.0416 | S3 | 1.6803 | 2.8210 | | 4.3811 | 2.6777 | 2.9773 | 3.5961 | 2.8330 | 2.4206 | 3.9793 | S4 | 2.8210 | 1.6803 | 4.3811 | | 2.9773 | 2.6777 | 2.4206 | 3.9793 | 3.5961 | 2.8330 | N5 | 1.3434 | 2.4119 | 2.6777 | 2.9773 | | 3.5882 | 1.0209 | 1.0127 | 3.8197 | 4.4358 | N6 | 2.4119 | 1.3434 | 2.9773 | 2.6777 | 3.5882 | | 3.8197 | 4.4358 | 1.0209 | 1.0127 | H7 | 2.0381 | 2.4951 | 3.5961 | 2.4206 | 1.0209 | 3.8197 | | 1.7795 | 4.2883 | 4.5047 | H8 | 2.0416 | 3.3587 | 2.8330 | 3.9793 | 1.0127 | 4.4358 | 1.7795 | | 4.5047 | 5.3412 | H9 | 2.4951 | 2.0381 | 2.4206 | 3.5961 | 3.8197 | 1.0209 | 4.2883 | 4.5047 | | 1.7795 | H10 | 3.3587 | 2.0416 | 3.9793 | 2.8330 | 4.4358 | 1.0127 | 4.5047 | 5.3412 | 1.7795 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
S4 |
122.324 |
|
C1 |
C2 |
N6 |
113.412 |
| C1 |
N5 |
H7 |
118.450 |
|
C1 |
N5 |
H8 |
119.450 |
| C2 |
C1 |
S3 |
122.324 |
|
C2 |
C1 |
N5 |
113.412 |
| C2 |
N6 |
H9 |
118.450 |
|
C2 |
N6 |
H10 |
119.450 |
| S3 |
C1 |
N5 |
124.264 |
|
S4 |
C2 |
N6 |
124.264 |
| H7 |
N5 |
H8 |
122.101 |
|
H9 |
N6 |
H10 |
122.101 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
