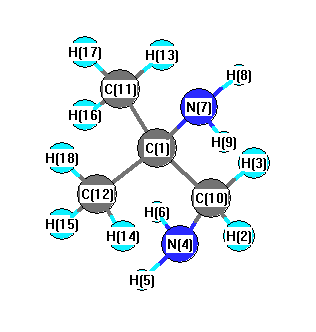
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3741 |
3407 |
1.45 |
|
|
|
| 2 |
A |
3710 |
3378 |
0.83 |
|
|
|
| 3 |
A |
3255 |
2964 |
69.22 |
|
|
|
| 4 |
A |
3229 |
2941 |
77.36 |
|
|
|
| 5 |
A |
3169 |
2886 |
8.24 |
|
|
|
| 6 |
A |
3160 |
2877 |
43.04 |
|
|
|
| 7 |
A |
1785 |
1625 |
22.23 |
|
|
|
| 8 |
A |
1780 |
1621 |
38.34 |
|
|
|
| 9 |
A |
1613 |
1469 |
6.14 |
|
|
|
| 10 |
A |
1604 |
1461 |
6.21 |
|
|
|
| 11 |
A |
1590 |
1448 |
2.07 |
|
|
|
| 12 |
A |
1534 |
1397 |
11.11 |
|
|
|
| 13 |
A |
1489 |
1356 |
3.58 |
|
|
|
| 14 |
A |
1366 |
1244 |
9.01 |
|
|
|
| 15 |
A |
1325 |
1206 |
28.06 |
|
|
|
| 16 |
A |
1178 |
1073 |
29.53 |
|
|
|
| 17 |
A |
1092 |
995 |
1.74 |
|
|
|
| 18 |
A |
1002 |
912 |
12.44 |
|
|
|
| 19 |
A |
937 |
853 |
263.06 |
|
|
|
| 20 |
A |
903 |
822 |
90.94 |
|
|
|
| 21 |
A |
774 |
705 |
14.84 |
|
|
|
| 22 |
A |
550 |
500 |
11.68 |
|
|
|
| 23 |
A |
460 |
419 |
9.51 |
|
|
|
| 24 |
A |
390 |
355 |
0.11 |
|
|
|
| 25 |
A |
286 |
260 |
0.29 |
|
|
|
| 26 |
A |
276 |
251 |
4.95 |
|
|
|
| 27 |
A |
3827 |
3485 |
2.94 |
|
|
|
| 28 |
A |
3792 |
3453 |
0.50 |
|
|
|
| 29 |
A |
3247 |
2957 |
0.74 |
|
|
|
| 30 |
A |
3225 |
2936 |
25.95 |
|
|
|
| 31 |
A |
3199 |
2913 |
47.37 |
|
|
|
| 32 |
A |
3159 |
2877 |
56.06 |
|
|
|
| 33 |
A |
1598 |
1455 |
0.35 |
|
|
|
| 34 |
A |
1585 |
1443 |
0.01 |
|
|
|
| 35 |
A |
1528 |
1392 |
10.66 |
|
|
|
| 36 |
A |
1499 |
1365 |
1.55 |
|
|
|
| 37 |
A |
1443 |
1314 |
0.60 |
|
|
|
| 38 |
A |
1265 |
1152 |
0.29 |
|
|
|
| 39 |
A |
1127 |
1026 |
0.63 |
|
|
|
| 40 |
A |
1076 |
980 |
0.66 |
|
|
|
| 41 |
A |
1011 |
920 |
0.10 |
|
|
|
| 42 |
A |
917 |
835 |
0.01 |
|
|
|
| 43 |
A |
483 |
440 |
1.74 |
|
|
|
| 44 |
A |
373 |
340 |
10.27 |
|
|
|
| 45 |
A |
312 |
284 |
0.81 |
|
|
|
| 46 |
A |
279 |
254 |
78.22 |
|
|
|
| 47 |
A |
232 |
211 |
0.11 |
|
|
|
| 48 |
A |
119 |
108 |
1.98 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 38745.8 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 35282.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-3.463 |
|
|
|
| 2 |
H |
-0.084 |
|
|
|
| 3 |
H |
-0.084 |
|
|
|
| 4 |
N |
-0.418 |
|
|
|
| 5 |
H |
-0.001 |
|
|
|
| 6 |
H |
-0.001 |
|
|
|
| 7 |
N |
0.007 |
|
|
|
| 8 |
H |
-0.045 |
|
|
|
| 9 |
H |
-0.045 |
|
|
|
| 10 |
C |
1.618 |
|
|
|
| 11 |
C |
1.307 |
|
|
|
| 12 |
C |
1.307 |
|
|
|
| 13 |
H |
-0.039 |
|
|
|
| 14 |
H |
-0.039 |
|
|
|
| 15 |
H |
-0.015 |
|
|
|
| 16 |
H |
-0.015 |
|
|
|
| 17 |
H |
0.004 |
|
|
|
| 18 |
H |
0.004 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.294 |
-0.064 |
0.000 |
0.301 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.087 |
4.311 |
0.000 |
| y |
4.311 |
-49.143 |
0.000 |
| z |
0.000 |
0.000 |
-37.717 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
5.343 |
4.311 |
0.000 |
| y |
4.311 |
-11.241 |
0.000 |
| z |
0.000 |
0.000 |
5.898 |
|
| Polar |
|---|
| 3z2-r2 | 11.796 |
|---|
| x2-y2 | 11.056 |
|---|
| xy | 4.311 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
0.000 |
0.000 |
0.000 |
| y |
0.000 |
0.000 |
0.000 |
| z |
0.000 |
0.000 |
0.000 |
<r2> (average value of r
2) Å
2
| <r2> |
183.322 |
| (<r2>)1/2 |
13.540 |