Jump to
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -217.110518 |
| Energy at 298.15K | |
| HF Energy | -217.110518 |
| Nuclear repulsion energy | 116.211501 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3282 |
3145 |
4.32 |
53.24 |
0.71 |
0.83 |
| 2 |
A' |
3188 |
3055 |
4.73 |
183.03 |
0.12 |
0.21 |
| 3 |
A' |
3175 |
3043 |
8.61 |
24.26 |
0.72 |
0.83 |
| 4 |
A' |
3049 |
2922 |
33.78 |
191.39 |
0.09 |
0.17 |
| 5 |
A' |
1737 |
1665 |
6.15 |
44.45 |
0.05 |
0.10 |
| 6 |
A' |
1471 |
1409 |
5.25 |
12.79 |
0.51 |
0.67 |
| 7 |
A' |
1439 |
1379 |
8.31 |
8.53 |
0.41 |
0.58 |
| 8 |
A' |
1395 |
1337 |
8.97 |
3.15 |
0.65 |
0.79 |
| 9 |
A' |
1310 |
1255 |
0.22 |
19.67 |
0.23 |
0.37 |
| 10 |
A' |
1136 |
1088 |
60.78 |
1.53 |
0.75 |
0.85 |
| 11 |
A' |
1008 |
966 |
36.47 |
4.69 |
0.65 |
0.79 |
| 12 |
A' |
923 |
885 |
2.83 |
5.94 |
0.04 |
0.07 |
| 13 |
A' |
607 |
582 |
6.88 |
1.64 |
0.68 |
0.81 |
| 14 |
A' |
257 |
246 |
2.18 |
1.31 |
0.45 |
0.62 |
| 15 |
A" |
3094 |
2965 |
18.88 |
86.14 |
0.75 |
0.86 |
| 16 |
A" |
1253 |
1201 |
0.00 |
3.60 |
0.75 |
0.86 |
| 17 |
A" |
1037 |
994 |
11.71 |
1.20 |
0.75 |
0.86 |
| 18 |
A" |
1012 |
969 |
9.14 |
0.13 |
0.75 |
0.86 |
| 19 |
A" |
966 |
926 |
46.32 |
2.43 |
0.75 |
0.86 |
| 20 |
A" |
560 |
536 |
11.88 |
1.34 |
0.75 |
0.86 |
| 21 |
A" |
176 |
169 |
3.22 |
0.87 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 16036.6 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 15367.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.942 |
-0.205 |
0.000 |
| C2 |
0.000 |
0.949 |
0.000 |
| C3 |
1.328 |
0.856 |
0.000 |
| F4 |
-0.270 |
-1.421 |
0.000 |
| H5 |
1.952 |
1.748 |
0.000 |
| H6 |
1.826 |
-0.112 |
0.000 |
| H7 |
-0.487 |
1.928 |
0.000 |
| H8 |
-1.589 |
-0.186 |
0.891 |
| H9 |
-1.589 |
-0.186 |
-0.891 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4904 | 2.5063 | 1.3894 | 3.4918 | 2.7700 | 2.1812 | 1.1009 | 1.1009 |
C2 | 1.4904 | | 1.3315 | 2.3861 | 2.1092 | 2.1122 | 1.0932 | 2.1462 | 2.1462 | C3 | 2.5063 | 1.3315 | | 2.7823 | 1.0883 | 1.0885 | 2.1084 | 3.2229 | 3.2229 | F4 | 1.3894 | 2.3861 | 2.7823 | | 3.8706 | 2.4717 | 3.3563 | 2.0146 | 2.0146 | H5 | 3.4918 | 2.1092 | 1.0883 | 3.8706 | | 1.8638 | 2.4463 | 4.1316 | 4.1316 | H6 | 2.7700 | 2.1122 | 1.0885 | 2.4717 | 1.8638 | | 3.0844 | 3.5299 | 3.5299 | H7 | 2.1812 | 1.0932 | 2.1084 | 3.3563 | 2.4463 | 3.0844 | | 2.5445 | 2.5445 | H8 | 1.1009 | 2.1462 | 3.2229 | 2.0146 | 4.1316 | 3.5299 | 2.5445 | | 1.7819 | H9 | 1.1009 | 2.1462 | 3.2229 | 2.0146 | 4.1316 | 3.5299 | 2.5445 | 1.7819 | |
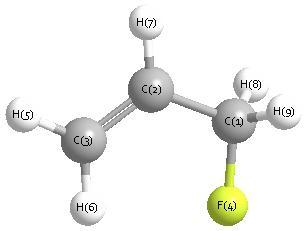 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.196 |
|
C1 |
C2 |
H7 |
114.307 |
| C2 |
C1 |
F4 |
111.855 |
|
C2 |
C1 |
H8 |
110.944 |
| C2 |
C1 |
H9 |
110.944 |
|
C2 |
C3 |
H5 |
120.970 |
| C2 |
C3 |
H6 |
121.244 |
|
C3 |
C2 |
H7 |
120.497 |
| F4 |
C1 |
H8 |
107.428 |
|
F4 |
C1 |
H9 |
107.428 |
| H5 |
C3 |
H6 |
117.786 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.877 |
|
|
|
| 2 |
C |
0.681 |
|
|
|
| 3 |
C |
0.360 |
|
|
|
| 4 |
F |
-0.452 |
|
|
|
| 5 |
H |
-0.365 |
|
|
|
| 6 |
H |
-0.390 |
|
|
|
| 7 |
H |
-0.481 |
|
|
|
| 8 |
H |
-0.115 |
|
|
|
| 9 |
H |
-0.115 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.915 |
1.535 |
0.000 |
1.787 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.225 |
-0.309 |
0.000 |
| y |
-0.309 |
-25.568 |
0.000 |
| z |
0.000 |
0.000 |
-25.351 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.234 |
-0.309 |
0.000 |
| y |
-0.309 |
-2.280 |
0.000 |
| z |
0.000 |
0.000 |
-1.954 |
|
| Polar |
|---|
| 3z2-r2 | -3.907 |
|---|
| x2-y2 | 4.343 |
|---|
| xy | -0.309 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.543 |
0.354 |
0.000 |
| y |
0.354 |
5.701 |
0.000 |
| z |
0.000 |
0.000 |
4.735 |
<r2> (average value of r
2) Å
2
| <r2> |
80.700 |
| (<r2>)1/2 |
8.983 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -217.110321 |
| Energy at 298.15K | |
| HF Energy | -217.110321 |
| Nuclear repulsion energy | 113.945957 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3264 |
3128 |
9.76 |
61.83 |
0.60 |
0.75 |
| 2 |
A |
3193 |
3060 |
6.02 |
112.63 |
0.22 |
0.36 |
| 3 |
A |
3165 |
3033 |
8.02 |
84.45 |
0.09 |
0.17 |
| 4 |
A |
3129 |
2998 |
17.39 |
66.11 |
0.71 |
0.83 |
| 5 |
A |
3065 |
2937 |
30.12 |
140.77 |
0.07 |
0.14 |
| 6 |
A |
1733 |
1661 |
0.48 |
45.83 |
0.07 |
0.13 |
| 7 |
A |
1481 |
1419 |
5.90 |
3.46 |
0.58 |
0.73 |
| 8 |
A |
1450 |
1389 |
16.64 |
9.98 |
0.53 |
0.69 |
| 9 |
A |
1368 |
1311 |
11.92 |
4.61 |
0.21 |
0.35 |
| 10 |
A |
1312 |
1257 |
0.09 |
14.17 |
0.31 |
0.47 |
| 11 |
A |
1254 |
1201 |
4.46 |
6.83 |
0.68 |
0.81 |
| 12 |
A |
1181 |
1132 |
2.84 |
1.13 |
0.40 |
0.57 |
| 13 |
A |
1043 |
1000 |
115.97 |
5.38 |
0.60 |
0.75 |
| 14 |
A |
1024 |
981 |
36.71 |
2.52 |
0.67 |
0.80 |
| 15 |
A |
988 |
947 |
3.79 |
2.46 |
0.14 |
0.25 |
| 16 |
A |
979 |
938 |
53.04 |
2.21 |
0.41 |
0.58 |
| 17 |
A |
926 |
888 |
2.25 |
3.08 |
0.13 |
0.23 |
| 18 |
A |
658 |
631 |
7.95 |
1.57 |
0.29 |
0.44 |
| 19 |
A |
433 |
415 |
2.43 |
3.55 |
0.32 |
0.48 |
| 20 |
A |
331 |
317 |
7.61 |
1.23 |
0.73 |
0.85 |
| 21 |
A |
115 |
110 |
0.99 |
2.67 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 16045.0 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 15376.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.605 |
0.429 |
0.326 |
| C2 |
0.644 |
-0.378 |
0.250 |
| C3 |
1.800 |
0.093 |
-0.219 |
| F4 |
-1.642 |
-0.225 |
-0.352 |
| H5 |
2.702 |
-0.517 |
-0.228 |
| H6 |
1.887 |
1.109 |
-0.604 |
| H7 |
0.576 |
-1.401 |
0.625 |
| H8 |
-0.942 |
0.553 |
1.365 |
| H9 |
-0.478 |
1.418 |
-0.135 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4897 | 2.4893 | 1.4011 | 3.4848 | 2.7459 | 2.1993 | 1.0997 | 1.0978 |
C2 | 1.4897 | | 1.3328 | 2.3696 | 2.1172 | 2.1178 | 1.0923 | 2.1512 | 2.1523 | C3 | 2.4893 | 1.3328 | | 3.4600 | 1.0890 | 1.0909 | 2.1075 | 3.2002 | 2.6373 | F4 | 1.4011 | 2.3696 | 3.4600 | | 4.3564 | 3.7821 | 2.6941 | 2.0109 | 2.0251 | H5 | 3.4848 | 2.1172 | 1.0890 | 4.3564 | | 1.8581 | 2.4553 | 4.1190 | 3.7244 | H6 | 2.7459 | 2.1178 | 1.0909 | 3.7821 | 1.8581 | | 3.0875 | 3.4920 | 2.4314 | H7 | 2.1993 | 1.0923 | 2.1075 | 2.6941 | 2.4553 | 3.0875 | | 2.5831 | 3.1042 | H8 | 1.0997 | 2.1512 | 3.2002 | 2.0109 | 4.1190 | 3.4920 | 2.5831 | | 1.7929 | H9 | 1.0978 | 2.1523 | 2.6373 | 2.0251 | 3.7244 | 2.4314 | 3.1042 | 1.7929 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.664 |
|
C1 |
C2 |
H7 |
115.961 |
| C2 |
C1 |
F4 |
110.072 |
|
C2 |
C1 |
H8 |
111.467 |
| C2 |
C1 |
H9 |
111.676 |
|
C2 |
C3 |
H5 |
121.579 |
| C2 |
C3 |
H6 |
121.480 |
|
C3 |
C2 |
H7 |
120.364 |
| F4 |
C1 |
H8 |
106.422 |
|
F4 |
C1 |
H9 |
107.649 |
| H5 |
C3 |
H6 |
116.939 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.762 |
|
|
|
| 2 |
C |
0.918 |
|
|
|
| 3 |
C |
0.413 |
|
|
|
| 4 |
F |
-0.465 |
|
|
|
| 5 |
H |
-0.349 |
|
|
|
| 6 |
H |
-0.391 |
|
|
|
| 7 |
H |
-0.460 |
|
|
|
| 8 |
H |
-0.150 |
|
|
|
| 9 |
H |
-0.278 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.568 |
0.761 |
0.887 |
1.956 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.874 |
-1.218 |
-2.093 |
| y |
-1.218 |
-22.579 |
-1.072 |
| z |
-2.093 |
-1.072 |
-24.962 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.103 |
-1.218 |
-2.093 |
| y |
-1.218 |
2.839 |
-1.072 |
| z |
-2.093 |
-1.072 |
-0.735 |
|
| Polar |
|---|
| 3z2-r2 | -1.470 |
|---|
| x2-y2 | -3.295 |
|---|
| xy | -1.218 |
|---|
| xz | -2.093 |
|---|
| yz | -1.072 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.549 |
0.252 |
-0.618 |
| y |
0.252 |
5.487 |
-0.209 |
| z |
-0.618 |
-0.209 |
5.068 |
<r2> (average value of r
2) Å
2
| <r2> |
89.362 |
| (<r2>)1/2 |
9.453 |
