return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH2CHCH2F (Allyl Fluoride)

using model chemistry: MP4/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS cis 1A'
Energy calculated at MP4/aug-cc-pVDZ
 hartrees
Energy at 0K-216.636779
Energy at 298.15K 
HF Energy-215.946228
Nuclear repulsion energy114.933991
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A' 3275 3200        
2 A' 3188 3114        
3 A' 3167 3093        
4 A' 3044 2973        
5 A' 1720 1680        
6 A' 1555 1520        
7 A' 1474 1440        
8 A' 1447 1413        
9 A' 1329 1298        
10 A' 1147 1121        
11 A' 1025 1001        
12 A' 930 909        
13 A' 607 593        
14 A' 281 274        
15 A" 3088 3016        
16 A" 1269 1240        
17 A" 1071 1046        
18 A" 1013 990        
19 A" 920 899        
20 A" 559 546        
21 A" 165 161        

Unscaled Zero Point Vibrational Energy (zpe) 16136.0 cm-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 15763.3 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4/aug-cc-pVDZ
ABC
0.55677 0.19667 0.14954

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4/aug-cc-pVDZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.962 -0.195 0.000
C2 0.000 0.966 0.000
C3 1.347 0.844 0.000
F4 -0.271 -1.430 0.000
H5 1.989 1.731 0.000
H6 1.826 -0.141 0.000
H7 -0.474 1.958 0.000
H8 -1.605 -0.183 0.900
H9 -1.605 -0.183 -0.900

Atom - Atom Distances (Å)
  C1 C2 C3 F4 H5 H6 H7 H8 H9
C11.50722.53211.41533.52392.78862.20741.10621.1062
C21.50721.35272.41122.13132.13501.10002.16892.1689
C32.53211.35272.79161.09501.09492.13513.25263.2526
F41.41532.41122.79163.88662.46243.39442.03592.0359
H53.52392.13131.09503.88661.87902.47364.17004.1700
H62.78862.13501.09492.46241.87903.11383.54743.5474
H72.20741.10002.13513.39442.47363.11382.58282.5828
H81.10622.16893.25262.03594.17003.54742.58281.7999
H91.10622.16893.25262.03594.17003.54742.58281.7999

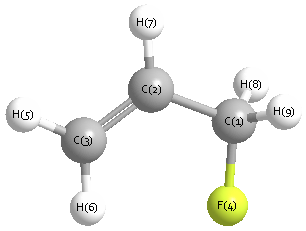
picture of Allyl Fluoride state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C3 124.504 C1 C2 H7 114.804
C2 C1 F4 111.145 C2 C1 H8 111.253
C2 C1 H9 111.253 C2 C3 H5 120.722
C2 C3 H6 121.088 C3 C2 H7 120.692
F4 C1 H8 107.053 F4 C1 H9 107.053
H5 C3 H6 118.190
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability