Jump to
S1C2
Energy calculated at CCD/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -216.608612 |
| Energy at 298.15K | |
| HF Energy | -215.947610 |
| Nuclear repulsion energy | 115.414429 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3277 |
3160 |
5.41 |
|
|
|
| 2 |
A' |
3189 |
3075 |
6.93 |
|
|
|
| 3 |
A' |
3173 |
3060 |
7.78 |
|
|
|
| 4 |
A' |
3061 |
2952 |
33.34 |
|
|
|
| 5 |
A' |
1731 |
1669 |
2.99 |
|
|
|
| 6 |
A' |
1498 |
1445 |
1.73 |
|
|
|
| 7 |
A' |
1457 |
1406 |
7.67 |
|
|
|
| 8 |
A' |
1422 |
1371 |
11.97 |
|
|
|
| 9 |
A' |
1315 |
1268 |
0.37 |
|
|
|
| 10 |
A' |
1142 |
1101 |
56.43 |
|
|
|
| 11 |
A' |
1013 |
977 |
31.96 |
|
|
|
| 12 |
A' |
922 |
889 |
3.77 |
|
|
|
| 13 |
A' |
604 |
582 |
7.93 |
|
|
|
| 14 |
A' |
265 |
256 |
2.70 |
|
|
|
| 15 |
A" |
3108 |
2998 |
20.90 |
|
|
|
| 16 |
A" |
1266 |
1221 |
0.06 |
|
|
|
| 17 |
A" |
1046 |
1009 |
13.17 |
|
|
|
| 18 |
A" |
1004 |
968 |
14.53 |
|
|
|
| 19 |
A" |
959 |
925 |
35.30 |
|
|
|
| 20 |
A" |
557 |
538 |
9.69 |
|
|
|
| 21 |
A" |
167 |
161 |
3.32 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16086.5 cm
-1
Scaled (by 0.9645) Zero Point Vibrational Energy (zpe) 15515.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.955 |
-0.204 |
0.000 |
| C2 |
0.000 |
0.961 |
0.000 |
| C3 |
1.340 |
0.850 |
0.000 |
| F4 |
-0.271 |
-1.425 |
0.000 |
| H5 |
1.975 |
1.741 |
0.000 |
| H6 |
1.830 |
-0.127 |
0.000 |
| H7 |
-0.480 |
1.948 |
0.000 |
| H8 |
-1.600 |
-0.188 |
0.897 |
| H9 |
-1.600 |
-0.188 |
-0.897 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5066 | 2.5266 | 1.3998 | 3.5173 | 2.7865 | 2.2041 | 1.1044 | 1.1044 |
C2 | 1.5066 | | 1.3450 | 2.4009 | 2.1235 | 2.1288 | 1.0977 | 2.1640 | 2.1640 | C3 | 2.5266 | 1.3450 | | 2.7880 | 1.0935 | 1.0931 | 2.1256 | 3.2447 | 3.2447 | F4 | 1.3998 | 2.4009 | 2.7880 | | 3.8814 | 2.4691 | 3.3794 | 2.0253 | 2.0253 | H5 | 3.5173 | 2.1235 | 1.0935 | 3.8814 | | 1.8737 | 2.4635 | 4.1598 | 4.1598 | H6 | 2.7865 | 2.1288 | 1.0931 | 2.4691 | 1.8737 | | 3.1049 | 3.5456 | 3.5456 | H7 | 2.2041 | 1.0977 | 2.1256 | 3.3794 | 2.4635 | 3.1049 | | 2.5730 | 2.5730 | H8 | 1.1044 | 2.1640 | 3.2447 | 2.0253 | 4.1598 | 3.5456 | 2.5730 | | 1.7936 | H9 | 1.1044 | 2.1640 | 3.2447 | 2.0253 | 4.1598 | 3.5456 | 2.5730 | 1.7936 | |
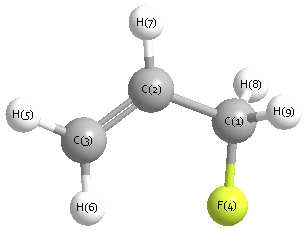 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
124.658 |
|
C1 |
C2 |
H7 |
114.721 |
| C2 |
C1 |
F4 |
111.344 |
|
C2 |
C1 |
H8 |
111.012 |
| C2 |
C1 |
H9 |
111.012 |
|
C2 |
C3 |
H5 |
120.764 |
| C2 |
C3 |
H6 |
121.299 |
|
C3 |
C2 |
H7 |
120.621 |
| F4 |
C1 |
H8 |
107.364 |
|
F4 |
C1 |
H9 |
107.364 |
| H5 |
C3 |
H6 |
117.937 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -216.608587 |
| Energy at 298.15K | |
| HF Energy | -215.947554 |
| Nuclear repulsion energy | 113.178251 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3259 |
3144 |
11.52 |
|
|
|
| 2 |
A |
3195 |
3082 |
5.61 |
|
|
|
| 3 |
A |
3162 |
3050 |
8.01 |
|
|
|
| 4 |
A |
3132 |
3021 |
18.73 |
|
|
|
| 5 |
A |
3073 |
2964 |
29.60 |
|
|
|
| 6 |
A |
1727 |
1665 |
0.24 |
|
|
|
| 7 |
A |
1503 |
1450 |
3.56 |
|
|
|
| 8 |
A |
1468 |
1416 |
15.43 |
|
|
|
| 9 |
A |
1391 |
1341 |
14.83 |
|
|
|
| 10 |
A |
1316 |
1269 |
0.07 |
|
|
|
| 11 |
A |
1267 |
1222 |
3.96 |
|
|
|
| 12 |
A |
1187 |
1145 |
4.02 |
|
|
|
| 13 |
A |
1060 |
1023 |
118.30 |
|
|
|
| 14 |
A |
1013 |
977 |
35.90 |
|
|
|
| 15 |
A |
987 |
952 |
1.78 |
|
|
|
| 16 |
A |
969 |
935 |
37.35 |
|
|
|
| 17 |
A |
929 |
896 |
2.00 |
|
|
|
| 18 |
A |
653 |
630 |
6.78 |
|
|
|
| 19 |
A |
431 |
416 |
1.88 |
|
|
|
| 20 |
A |
336 |
324 |
7.58 |
|
|
|
| 21 |
A |
113 |
109 |
1.27 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16085.9 cm
-1
Scaled (by 0.9645) Zero Point Vibrational Energy (zpe) 15514.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.612 |
0.428 |
0.337 |
| C2 |
0.649 |
-0.388 |
0.254 |
| C3 |
1.810 |
0.099 |
-0.223 |
| F4 |
-1.649 |
-0.222 |
-0.361 |
| H5 |
2.720 |
-0.508 |
-0.242 |
| H6 |
1.885 |
1.124 |
-0.603 |
| H7 |
0.587 |
-1.418 |
0.624 |
| H8 |
-0.956 |
0.540 |
1.379 |
| H9 |
-0.478 |
1.424 |
-0.117 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.5046 | 2.5077 | 1.4087 | 3.5090 | 2.7573 | 2.2203 | 1.1035 | 1.1025 |
C2 | 1.5046 | | 1.3464 | 2.3850 | 2.1325 | 2.1327 | 1.0967 | 2.1689 | 2.1663 | C3 | 2.5077 | 1.3464 | | 3.4770 | 1.0939 | 1.0957 | 2.1255 | 3.2269 | 2.6465 | F4 | 1.4087 | 2.3850 | 3.4770 | | 4.3797 | 3.7894 | 2.7203 | 2.0220 | 2.0347 | H5 | 3.5090 | 2.1325 | 1.0939 | 4.3797 | | 1.8686 | 2.4757 | 4.1521 | 3.7386 | H6 | 2.7573 | 2.1327 | 1.0957 | 3.7894 | 1.8686 | | 3.1075 | 3.5131 | 2.4314 | H7 | 2.2203 | 1.0967 | 2.1255 | 2.7203 | 2.4757 | 3.1075 | | 2.6050 | 3.1247 | H8 | 1.1035 | 2.1689 | 3.2269 | 2.0220 | 4.1521 | 3.5131 | 2.6050 | | 1.8020 | H9 | 1.1025 | 2.1663 | 2.6465 | 2.0347 | 3.7386 | 2.4314 | 3.1247 | 1.8020 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.093 |
|
C1 |
C2 |
H7 |
116.315 |
| C2 |
C1 |
F4 |
109.857 |
|
C2 |
C1 |
H8 |
111.604 |
| C2 |
C1 |
H9 |
111.456 |
|
C2 |
C3 |
H5 |
121.483 |
| C2 |
C3 |
H6 |
121.347 |
|
C3 |
C2 |
H7 |
120.584 |
| F4 |
C1 |
H8 |
106.565 |
|
F4 |
C1 |
H9 |
107.619 |
| H5 |
C3 |
H6 |
117.169 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
