Jump to
S1C2
Energy calculated at B97D3/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -217.042375 |
| Energy at 298.15K | |
| HF Energy | -217.042375 |
| Nuclear repulsion energy | 115.102198 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3201 |
3201 |
8.64 |
58.18 |
0.72 |
0.84 |
| 2 |
A' |
3108 |
3108 |
5.55 |
175.98 |
0.10 |
0.18 |
| 3 |
A' |
3091 |
3091 |
16.63 |
52.31 |
0.48 |
0.65 |
| 4 |
A' |
2960 |
2960 |
43.28 |
211.80 |
0.10 |
0.18 |
| 5 |
A' |
1673 |
1673 |
6.75 |
41.90 |
0.05 |
0.10 |
| 6 |
A' |
1433 |
1433 |
4.14 |
14.40 |
0.49 |
0.66 |
| 7 |
A' |
1409 |
1409 |
4.19 |
11.37 |
0.36 |
0.53 |
| 8 |
A' |
1355 |
1355 |
11.10 |
4.29 |
0.75 |
0.86 |
| 9 |
A' |
1283 |
1283 |
0.16 |
20.38 |
0.22 |
0.37 |
| 10 |
A' |
1084 |
1084 |
41.28 |
1.23 |
0.72 |
0.84 |
| 11 |
A' |
968 |
968 |
52.53 |
5.15 |
0.61 |
0.76 |
| 12 |
A' |
884 |
884 |
7.25 |
5.77 |
0.05 |
0.10 |
| 13 |
A' |
589 |
589 |
7.38 |
1.86 |
0.69 |
0.82 |
| 14 |
A' |
253 |
253 |
2.11 |
1.43 |
0.44 |
0.62 |
| 15 |
A" |
3000 |
3000 |
26.04 |
99.15 |
0.75 |
0.86 |
| 16 |
A" |
1214 |
1214 |
0.07 |
3.89 |
0.75 |
0.86 |
| 17 |
A" |
1003 |
1003 |
9.73 |
0.69 |
0.75 |
0.86 |
| 18 |
A" |
980 |
980 |
12.26 |
0.23 |
0.75 |
0.86 |
| 19 |
A" |
924 |
924 |
40.71 |
2.45 |
0.75 |
0.86 |
| 20 |
A" |
539 |
539 |
11.41 |
1.34 |
0.75 |
0.86 |
| 21 |
A" |
162 |
162 |
3.00 |
0.84 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 15555.8 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 15555.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.947 |
-0.206 |
0.000 |
| C2 |
0.000 |
0.955 |
0.000 |
| C3 |
1.339 |
0.876 |
0.000 |
| F4 |
-0.274 |
-1.444 |
0.000 |
| H5 |
1.954 |
1.780 |
0.000 |
| H6 |
1.853 |
-0.088 |
0.000 |
| H7 |
-0.500 |
1.934 |
0.000 |
| H8 |
-1.596 |
-0.189 |
0.896 |
| H9 |
-1.596 |
-0.189 |
-0.896 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4988 | 2.5296 | 1.4094 | 3.5164 | 2.8034 | 2.1863 | 1.1067 | 1.1067 |
C2 | 1.4988 | | 1.3410 | 2.4148 | 2.1212 | 2.1267 | 1.0988 | 2.1588 | 2.1588 | C3 | 2.5296 | 1.3410 | | 2.8258 | 1.0933 | 1.0931 | 2.1208 | 3.2486 | 3.2486 | F4 | 1.4094 | 2.4148 | 2.8258 | | 3.9191 | 2.5226 | 3.3854 | 2.0320 | 2.0320 | H5 | 3.5164 | 2.1212 | 1.0933 | 3.9191 | | 1.8708 | 2.4586 | 4.1577 | 4.1577 | H6 | 2.8034 | 2.1267 | 1.0931 | 2.5226 | 1.8708 | | 3.1022 | 3.5658 | 3.5658 | H7 | 2.1863 | 1.0988 | 2.1208 | 3.3854 | 2.4586 | 3.1022 | | 2.5517 | 2.5517 | H8 | 1.1067 | 2.1588 | 3.2486 | 2.0320 | 4.1577 | 3.5658 | 2.5517 | | 1.7926 | H9 | 1.1067 | 2.1588 | 3.2486 | 2.0320 | 4.1577 | 3.5658 | 2.5517 | 1.7926 | |
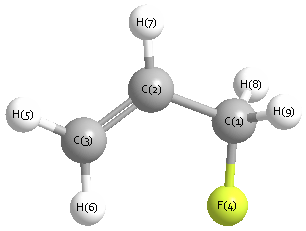 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
125.849 |
|
C1 |
C2 |
H7 |
113.740 |
| C2 |
C1 |
F4 |
112.230 |
|
C2 |
C1 |
H8 |
111.006 |
| C2 |
C1 |
H9 |
111.006 |
|
C2 |
C3 |
H5 |
120.893 |
| C2 |
C3 |
H6 |
121.447 |
|
C3 |
C2 |
H7 |
120.411 |
| F4 |
C1 |
H8 |
107.109 |
|
F4 |
C1 |
H9 |
107.109 |
| H5 |
C3 |
H6 |
117.660 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.997 |
|
|
|
| 2 |
C |
0.837 |
|
|
|
| 3 |
C |
0.532 |
|
|
|
| 4 |
F |
-0.438 |
|
|
|
| 5 |
H |
-0.459 |
|
|
|
| 6 |
H |
-0.490 |
|
|
|
| 7 |
H |
-0.589 |
|
|
|
| 8 |
H |
-0.195 |
|
|
|
| 9 |
H |
-0.195 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.898 |
1.535 |
0.000 |
1.778 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-21.538 |
-0.326 |
0.000 |
| y |
-0.326 |
-25.864 |
0.000 |
| z |
0.000 |
0.000 |
-25.456 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
4.123 |
-0.326 |
0.000 |
| y |
-0.326 |
-2.367 |
0.000 |
| z |
0.000 |
0.000 |
-1.755 |
|
| Polar |
|---|
| 3z2-r2 | -3.511 |
|---|
| x2-y2 | 4.326 |
|---|
| xy | -0.326 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.883 |
0.417 |
0.000 |
| y |
0.417 |
6.041 |
0.000 |
| z |
0.000 |
0.000 |
4.950 |
<r2> (average value of r
2) Å
2
| <r2> |
82.284 |
| (<r2>)1/2 |
9.071 |
Jump to
S1C1
Energy calculated at B97D3/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -217.042905 |
| Energy at 298.15K | |
| HF Energy | -217.042905 |
| Nuclear repulsion energy | 112.998188 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B97D3/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3182 |
3182 |
16.79 |
69.43 |
0.59 |
0.74 |
| 2 |
A |
3112 |
3112 |
10.00 |
120.66 |
0.23 |
0.37 |
| 3 |
A |
3086 |
3086 |
12.82 |
95.65 |
0.10 |
0.17 |
| 4 |
A |
3044 |
3044 |
22.82 |
74.19 |
0.71 |
0.83 |
| 5 |
A |
2982 |
2982 |
38.11 |
151.47 |
0.08 |
0.14 |
| 6 |
A |
1667 |
1667 |
0.45 |
44.49 |
0.07 |
0.13 |
| 7 |
A |
1447 |
1447 |
5.10 |
4.26 |
0.50 |
0.67 |
| 8 |
A |
1417 |
1417 |
10.43 |
11.88 |
0.48 |
0.65 |
| 9 |
A |
1330 |
1330 |
12.45 |
4.76 |
0.26 |
0.41 |
| 10 |
A |
1286 |
1286 |
0.23 |
14.41 |
0.31 |
0.48 |
| 11 |
A |
1216 |
1216 |
4.94 |
7.92 |
0.66 |
0.80 |
| 12 |
A |
1149 |
1149 |
2.30 |
0.95 |
0.39 |
0.56 |
| 13 |
A |
990 |
990 |
13.55 |
0.54 |
0.74 |
0.85 |
| 14 |
A |
960 |
960 |
52.99 |
3.25 |
0.61 |
0.75 |
| 15 |
A |
954 |
954 |
66.50 |
6.89 |
0.43 |
0.60 |
| 16 |
A |
940 |
940 |
64.55 |
1.57 |
0.53 |
0.70 |
| 17 |
A |
902 |
902 |
15.85 |
5.62 |
0.19 |
0.33 |
| 18 |
A |
638 |
638 |
7.35 |
1.68 |
0.26 |
0.41 |
| 19 |
A |
424 |
424 |
2.26 |
3.90 |
0.31 |
0.48 |
| 20 |
A |
329 |
329 |
8.26 |
1.41 |
0.74 |
0.85 |
| 21 |
A |
112 |
112 |
0.90 |
2.69 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 15583.0 cm
-1
Scaled (by 1) Zero Point Vibrational Energy (zpe) 15583.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B97D3/aug-cc-pVDZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.606 |
0.430 |
0.344 |
| C2 |
0.652 |
-0.378 |
0.264 |
| C3 |
1.807 |
0.094 |
-0.231 |
| F4 |
-1.651 |
-0.225 |
-0.369 |
| H5 |
2.716 |
-0.514 |
-0.247 |
| H6 |
1.884 |
1.111 |
-0.632 |
| H7 |
0.592 |
-1.402 |
0.654 |
| H8 |
-0.963 |
0.530 |
1.385 |
| H9 |
-0.481 |
1.430 |
-0.104 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
F4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4967 | 2.5031 | 1.4252 | 3.5036 | 2.7596 | 2.2102 | 1.1049 | 1.1031 |
C2 | 1.4967 | | 1.3427 | 2.3933 | 2.1309 | 2.1305 | 1.0972 | 2.1654 | 2.1649 | C3 | 2.5031 | 1.3427 | | 3.4757 | 1.0941 | 1.0959 | 2.1209 | 3.2368 | 2.6523 | F4 | 1.4252 | 2.3933 | 3.4757 | | 4.3786 | 3.7885 | 2.7319 | 2.0299 | 2.0447 | H5 | 3.5036 | 2.1309 | 1.0941 | 4.3786 | | 1.8666 | 2.4719 | 4.1581 | 3.7444 | H6 | 2.7596 | 2.1305 | 1.0959 | 3.7885 | 1.8666 | | 3.1046 | 3.5371 | 2.4437 | H7 | 2.2102 | 1.0972 | 2.1209 | 2.7319 | 2.4719 | 3.1046 | | 2.5853 | 3.1215 | H8 | 1.1049 | 2.1654 | 3.2368 | 2.0299 | 4.1581 | 3.5371 | 2.5853 | | 1.8053 | H9 | 1.1031 | 2.1649 | 2.6523 | 2.0447 | 3.7444 | 2.4437 | 3.1215 | 1.8053 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
123.562 |
|
C1 |
C2 |
H7 |
116.025 |
| C2 |
C1 |
F4 |
109.964 |
|
C2 |
C1 |
H8 |
111.791 |
| C2 |
C1 |
H9 |
111.864 |
|
C2 |
C3 |
H5 |
121.631 |
| C2 |
C3 |
H6 |
121.436 |
|
C3 |
C2 |
H7 |
120.407 |
| F4 |
C1 |
H8 |
106.007 |
|
F4 |
C1 |
H9 |
107.257 |
| H5 |
C3 |
H6 |
116.931 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B97D3/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.896 |
|
|
|
| 2 |
C |
1.082 |
|
|
|
| 3 |
C |
0.577 |
|
|
|
| 4 |
F |
-0.453 |
|
|
|
| 5 |
H |
-0.442 |
|
|
|
| 6 |
H |
-0.496 |
|
|
|
| 7 |
H |
-0.558 |
|
|
|
| 8 |
H |
-0.246 |
|
|
|
| 9 |
H |
-0.361 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.600 |
0.737 |
0.910 |
1.982 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.130 |
-1.218 |
-2.116 |
| y |
-1.218 |
-22.867 |
-1.033 |
| z |
-2.116 |
-1.033 |
-25.055 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.170 |
-1.218 |
-2.116 |
| y |
-1.218 |
2.726 |
-1.033 |
| z |
-2.116 |
-1.033 |
-0.557 |
|
| Polar |
|---|
| 3z2-r2 | -1.113 |
|---|
| x2-y2 | -3.264 |
|---|
| xy | -1.218 |
|---|
| xz | -2.116 |
|---|
| yz | -1.033 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.962 |
0.225 |
-0.643 |
| y |
0.225 |
5.746 |
-0.216 |
| z |
-0.643 |
-0.216 |
5.325 |
<r2> (average value of r
2) Å
2
| <r2> |
90.494 |
| (<r2>)1/2 |
9.513 |
