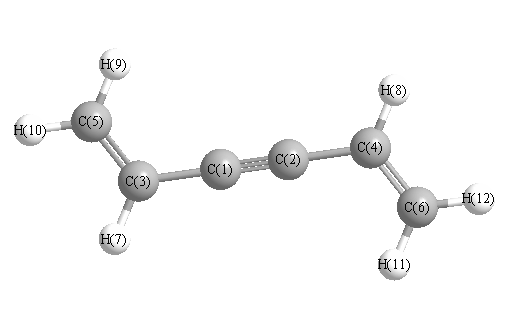
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Ag |
3286 |
3149 |
0.00 |
|
|
|
| 2 |
Ag |
3185 |
3052 |
0.00 |
|
|
|
| 3 |
Ag |
3176 |
3044 |
0.00 |
|
|
|
| 4 |
Ag |
2325 |
2228 |
0.00 |
|
|
|
| 5 |
Ag |
1664 |
1594 |
0.00 |
|
|
|
| 6 |
Ag |
1428 |
1368 |
0.00 |
|
|
|
| 7 |
Ag |
1314 |
1259 |
0.00 |
|
|
|
| 8 |
Ag |
1074 |
1029 |
0.00 |
|
|
|
| 9 |
Ag |
721 |
691 |
0.00 |
|
|
|
| 10 |
Ag |
491 |
471 |
0.00 |
|
|
|
| 11 |
Ag |
198 |
190 |
0.00 |
|
|
|
| 12 |
Au |
1003 |
962 |
33.52 |
|
|
|
| 13 |
Au |
956 |
916 |
84.81 |
|
|
|
| 14 |
Au |
686 |
657 |
11.58 |
|
|
|
| 15 |
Au |
169 |
162 |
7.40 |
|
|
|
| 16 |
Au |
47 |
46 |
0.16 |
|
|
|
| 17 |
Bg |
999 |
958 |
0.00 |
|
|
|
| 18 |
Bg |
956 |
916 |
0.00 |
|
|
|
| 19 |
Bg |
693 |
664 |
0.00 |
|
|
|
| 20 |
Bg |
301 |
289 |
0.00 |
|
|
|
| 21 |
Bu |
3286 |
3149 |
10.33 |
|
|
|
| 22 |
Bu |
3185 |
3052 |
11.52 |
|
|
|
| 23 |
Bu |
3176 |
3044 |
10.93 |
|
|
|
| 24 |
Bu |
1698 |
1627 |
40.44 |
|
|
|
| 25 |
Bu |
1451 |
1391 |
6.91 |
|
|
|
| 26 |
Bu |
1312 |
1257 |
3.92 |
|
|
|
| 27 |
Bu |
1225 |
1174 |
15.14 |
|
|
|
| 28 |
Bu |
1014 |
972 |
1.31 |
|
|
|
| 29 |
Bu |
522 |
500 |
13.18 |
|
|
|
| 30 |
Bu |
108 |
104 |
2.77 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 20824.4 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 19956.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.140 |
|
|
|
| 2 |
C |
-0.140 |
|
|
|
| 3 |
C |
1.021 |
|
|
|
| 4 |
C |
1.021 |
|
|
|
| 5 |
C |
0.473 |
|
|
|
| 6 |
C |
0.473 |
|
|
|
| 7 |
H |
-0.568 |
|
|
|
| 8 |
H |
-0.568 |
|
|
|
| 9 |
H |
-0.393 |
|
|
|
| 10 |
H |
-0.393 |
|
|
|
| 11 |
H |
-0.393 |
|
|
|
| 12 |
H |
-0.393 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
0.000 |
0.000 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.958 |
0.537 |
0.000 |
| y |
0.537 |
-27.508 |
0.000 |
| z |
0.000 |
0.000 |
-40.119 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.855 |
0.537 |
0.000 |
| y |
0.537 |
9.031 |
0.000 |
| z |
0.000 |
0.000 |
-9.886 |
|
| Polar |
|---|
| 3z2-r2 | -19.772 |
|---|
| x2-y2 | -5.450 |
|---|
| xy | 0.537 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.846 |
4.568 |
0.000 |
| y |
4.568 |
21.866 |
0.000 |
| z |
0.000 |
0.000 |
7.092 |
<r2> (average value of r
2) Å
2
| <r2> |
249.270 |
| (<r2>)1/2 |
15.788 |