return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H6 (Hexa-1,5-diene-3-yne)

using model chemistry: MP4/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2H 1Ag
Energy calculated at MP4/aug-cc-pVDZ
 hartrees
Energy at 0K-231.513438
Energy at 298.15K-231.516667
HF Energy-230.616915
Nuclear repulsion energy172.463658
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 Ag 3268 3192        
2 Ag 3178 3105        
3 Ag 3164 3091        
4 Ag 2232 2180        
5 Ag 1653 1614        
6 Ag 1457 1423        
7 Ag 1338 1307        
8 Ag 1101 1075        
9 Ag 719 702        
10 Ag 512 500        
11 Ag 217 212        
12 Au 992 970        
13 Au 899 878        
14 Au 667 652        
15 Au 177 173        
16 Au 38 37        
17 Bg 990 967        
18 Bg 900 879        
19 Bg 645 630        
20 Bg 259 253        
21 Bu 3268 3192        
22 Bu 3178 3104        
23 Bu 3164 3091        
24 Bu 1677 1639        
25 Bu 1477 1443        
26 Bu 1335 1305        
27 Bu 1230 1201        
28 Bu 1023 1000        
29 Bu 525 513        
30 Bu 110 107        

Unscaled Zero Point Vibrational Energy (zpe) 20696.2 cm-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 20218.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4/aug-cc-pVDZ
ABC
0.83928 0.04419 0.04198

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4/aug-cc-pVDZ

Point Group is C2h

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.622 0.000
C2 0.000 -0.622 0.000
C3 0.000 2.063 0.000
C4 0.000 -2.063 0.000
C5 1.133 2.819 0.000
C6 -1.133 -2.819 0.000
H7 -0.982 2.555 0.000
H8 0.982 -2.555 0.000
H9 2.127 2.357 0.000
H10 1.071 3.913 0.000
H11 -2.127 -2.357 0.000
H12 -1.071 -3.913 0.000

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12
C11.24351.44122.68472.47263.62292.16823.32482.74483.46113.66014.6594
C21.24352.68471.44123.62292.47263.32482.16823.66014.65942.74483.4611
C31.44122.68474.12591.36245.01211.09844.72092.14682.13774.90516.0711
C42.68471.44124.12595.01211.36244.72091.09844.90516.07112.14682.1377
C52.47263.62291.36245.01216.07712.13185.37621.09581.09526.11737.0840
C63.62292.47265.01211.36246.07715.37622.13186.11737.08401.09581.0952
H72.16823.32481.09844.72092.13185.37625.47403.11512.46205.04336.4681
H83.32482.16824.72091.09845.37622.13185.47405.04336.46813.11512.4620
H92.74483.66012.14684.90511.09586.11733.11515.04331.88006.34937.0384
H103.46114.65942.13776.07111.09527.08402.46206.46811.88007.03848.1137
H113.66012.74484.90512.14686.11731.09585.04333.11516.34937.03841.8800
H124.65943.46116.07112.13777.08401.09526.46812.46207.03848.11371.8800

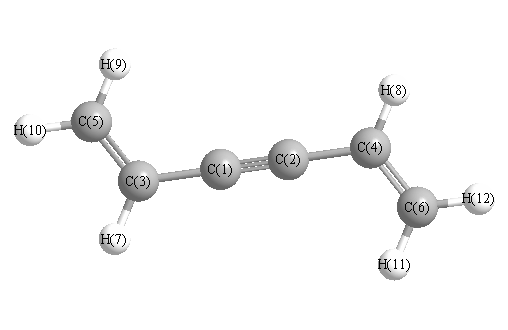
picture of Hexa-1,5-diene-3-yne state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 180.000 C1 C3 C5 123.726
C1 C3 H7 116.592 C2 C1 C3 180.000
C2 C4 C6 123.726 C2 C4 H8 116.592
C3 C5 H9 121.321 C3 C5 H10 120.486
C4 C6 H11 121.321 C4 C6 H12 120.486
C5 C3 H7 119.682 C6 C4 H8 119.682
H9 C5 H10 118.193 H11 C6 H12 118.193
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability