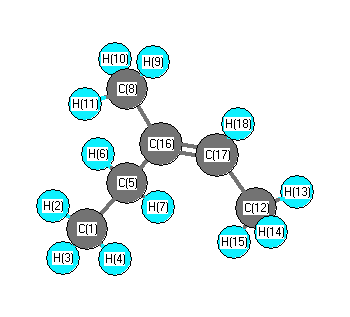
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3162 |
3031 |
52.14 |
|
|
|
| 2 |
A |
3147 |
3015 |
15.11 |
|
|
|
| 3 |
A |
3144 |
3013 |
9.81 |
|
|
|
| 4 |
A |
3142 |
3011 |
34.69 |
|
|
|
| 5 |
A |
3137 |
3007 |
36.69 |
|
|
|
| 6 |
A |
3108 |
2978 |
8.89 |
|
|
|
| 7 |
A |
3098 |
2968 |
7.22 |
|
|
|
| 8 |
A |
3096 |
2967 |
35.08 |
|
|
|
| 9 |
A |
3056 |
2928 |
30.98 |
|
|
|
| 10 |
A |
3053 |
2926 |
32.02 |
|
|
|
| 11 |
A |
3041 |
2915 |
42.00 |
|
|
|
| 12 |
A |
3036 |
2910 |
33.16 |
|
|
|
| 13 |
A |
1761 |
1688 |
1.10 |
|
|
|
| 14 |
A |
1491 |
1429 |
9.76 |
|
|
|
| 15 |
A |
1478 |
1416 |
7.96 |
|
|
|
| 16 |
A |
1477 |
1415 |
4.09 |
|
|
|
| 17 |
A |
1475 |
1413 |
5.02 |
|
|
|
| 18 |
A |
1459 |
1398 |
1.05 |
|
|
|
| 19 |
A |
1458 |
1397 |
11.62 |
|
|
|
| 20 |
A |
1455 |
1394 |
1.39 |
|
|
|
| 21 |
A |
1404 |
1345 |
2.18 |
|
|
|
| 22 |
A |
1396 |
1338 |
5.77 |
|
|
|
| 23 |
A |
1387 |
1330 |
5.14 |
|
|
|
| 24 |
A |
1376 |
1319 |
0.62 |
|
|
|
| 25 |
A |
1324 |
1269 |
2.99 |
|
|
|
| 26 |
A |
1289 |
1236 |
1.66 |
|
|
|
| 27 |
A |
1234 |
1182 |
1.31 |
|
|
|
| 28 |
A |
1141 |
1093 |
5.51 |
|
|
|
| 29 |
A |
1097 |
1051 |
3.12 |
|
|
|
| 30 |
A |
1079 |
1034 |
1.72 |
|
|
|
| 31 |
A |
1050 |
1006 |
1.61 |
|
|
|
| 32 |
A |
1042 |
999 |
1.76 |
|
|
|
| 33 |
A |
1016 |
974 |
2.92 |
|
|
|
| 34 |
A |
1009 |
967 |
6.83 |
|
|
|
| 35 |
A |
927 |
889 |
6.03 |
|
|
|
| 36 |
A |
830 |
796 |
14.09 |
|
|
|
| 37 |
A |
787 |
754 |
1.39 |
|
|
|
| 38 |
A |
756 |
725 |
1.00 |
|
|
|
| 39 |
A |
571 |
548 |
3.51 |
|
|
|
| 40 |
A |
524 |
502 |
2.60 |
|
|
|
| 41 |
A |
393 |
377 |
0.45 |
|
|
|
| 42 |
A |
302 |
289 |
0.34 |
|
|
|
| 43 |
A |
270 |
259 |
1.35 |
|
|
|
| 44 |
A |
236 |
226 |
1.31 |
|
|
|
| 45 |
A |
212 |
204 |
0.03 |
|
|
|
| 46 |
A |
179 |
172 |
1.01 |
|
|
|
| 47 |
A |
124 |
119 |
0.23 |
|
|
|
| 48 |
A |
50 |
48 |
0.03 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 36139.7 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 34632.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.459 |
|
|
|
| 2 |
H |
-0.172 |
|
|
|
| 3 |
H |
-0.157 |
|
|
|
| 4 |
H |
-0.128 |
|
|
|
| 5 |
C |
0.477 |
|
|
|
| 6 |
H |
-0.279 |
|
|
|
| 7 |
H |
-0.187 |
|
|
|
| 8 |
C |
0.097 |
|
|
|
| 9 |
H |
-0.262 |
|
|
|
| 10 |
H |
0.019 |
|
|
|
| 11 |
H |
-0.057 |
|
|
|
| 12 |
C |
0.088 |
|
|
|
| 13 |
H |
-0.002 |
|
|
|
| 14 |
H |
0.043 |
|
|
|
| 15 |
H |
-0.335 |
|
|
|
| 16 |
C |
0.738 |
|
|
|
| 17 |
C |
0.417 |
|
|
|
| 18 |
H |
-0.762 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.139 |
-0.165 |
0.055 |
0.223 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.994 |
0.315 |
0.720 |
| y |
0.315 |
-39.634 |
-0.620 |
| z |
0.720 |
-0.620 |
-41.034 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
1.340 |
0.315 |
0.720 |
| y |
0.315 |
0.380 |
-0.620 |
| z |
0.720 |
-0.620 |
-1.720 |
|
| Polar |
|---|
| 3z2-r2 | -3.439 |
|---|
| x2-y2 | 0.640 |
|---|
| xy | 0.315 |
|---|
| xz | 0.720 |
|---|
| yz | -0.620 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.879 |
0.379 |
0.418 |
| y |
0.379 |
11.401 |
-0.087 |
| z |
0.418 |
-0.087 |
9.378 |
<r2> (average value of r
2) Å
2
| <r2> |
209.339 |
| (<r2>)1/2 |
14.469 |