return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C6H10 (3-Hexyne)

using model chemistry: MP4/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 1A1
Energy calculated at MP4/aug-cc-pVDZ
 hartrees
Energy at 0K-233.958778
Energy at 298.15K 
HF Energy-232.997994
Nuclear repulsion energy204.872289
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP4/6-31G*
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 A 3154 3081        
2 A 3145 3072        
3 A 3083 3011        
4 A 3063 2993        
5 A 3044 2974        
6 A 2280 2227        
7 A 1550 1514        
8 A 1540 1504        
9 A 1531 1495        
10 A 1454 1420        
11 A 1384 1352        
12 A 1310 1280        
13 A 1116 1090        
14 A 1111 1085        
15 A 1017 994        
16 A 793 775        
17 A 697 681        
18 A 425 415        
19 A 322 315        
20 A 200 195        
21 A 88 86        
22 A 20i 20i        
23 B 3154 3081        
24 B 3144 3072        
25 B 3082 3011        
26 B 3063 2992        
27 B 3044 2974        
28 B 1550 1514        
29 B 1541 1505        
30 B 1529 1494        
31 B 1453 1419        
32 B 1385 1353        
33 B 1314 1284        
34 B 1181 1154        
35 B 1136 1110        
36 B 1102 1076        
37 B 948 926        
38 B 801 783        
39 B 494 483        
40 B 296 289        
41 B 171 167        
42 B 151 148        

Unscaled Zero Point Vibrational Energy (zpe) 31411.2 cm-1
Scaled (by 0.9769) Zero Point Vibrational Energy (zpe) 30685.6 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at MP4/aug-cc-pVDZ
ABC
0.30365 0.04613 0.04131

See section I.F.4 to change rotational constant units
Geometric Data calculated at MP4/aug-cc-pVDZ

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 -0.001 0.619 -0.586
C2 0.001 -0.619 -0.586
C3 -0.001 2.101 -0.557
C4 0.001 -2.101 -0.557
C5 -0.001 2.655 0.886
C6 0.001 -2.655 0.886
H7 0.888 2.473 -1.101
H8 -0.889 2.473 -1.101
H9 -0.888 -2.473 -1.101
H10 0.889 -2.473 -1.101
H11 -0.001 3.760 0.875
H12 -0.895 2.309 1.433
H13 0.894 2.309 1.433
H14 0.001 -3.760 0.875
H15 0.895 -2.309 1.433
H16 -0.894 -2.309 1.433

Atom - Atom Distances (Å)
  C1 C2 C3 C4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14 H15 H16
C11.23801.48272.72052.51233.58962.11982.11983.25803.25883.46402.78102.78104.61613.66793.6672
C21.23802.72051.48273.58962.51233.25803.25882.11982.11984.61613.66793.66723.46402.78102.7810
C31.48272.72054.20281.54504.97031.10651.10654.69164.69222.19082.19152.19156.03364.92084.9203
C42.72051.48274.20284.97031.54504.69164.69221.10651.10656.03364.92084.92032.19082.19152.1915
C52.51233.58961.54504.97035.30992.18432.18435.57095.57131.10501.10421.10426.41485.07385.0733
C63.58962.51234.97031.54505.30995.57095.57132.18432.18436.41485.07385.07331.10501.10421.1042
H72.11983.25801.10654.69162.18435.57091.77675.25564.94672.51993.10312.54006.59895.41255.6981
H82.11983.25881.10654.69222.18435.57131.77674.94675.25652.51992.54003.10316.59935.69905.4125
H93.25802.11984.69161.10655.57092.18435.25564.94671.77676.59895.41255.69812.51993.10312.5400
H103.25882.11984.69221.10655.57132.18434.94675.25651.77676.59935.69905.41252.51992.54003.1031
H113.46404.61612.19086.03361.10506.41482.51992.51996.59896.59931.79351.79357.51986.16006.1596
H122.78103.66792.19154.92081.10425.07383.10312.54005.41255.69901.79351.78876.16004.95294.6181
H132.78103.66722.19154.92031.10425.07332.54003.10315.69815.41251.79351.78876.15964.61814.9518
H144.61613.46406.03362.19086.41481.10506.59896.59932.51992.51997.51986.16006.15961.79351.7935
H153.66792.78104.92082.19155.07381.10425.41255.69903.10312.54006.16004.95294.61811.79351.7887
H163.66722.78104.92032.19155.07331.10425.69815.41252.54003.10316.15964.61814.95181.79351.7887

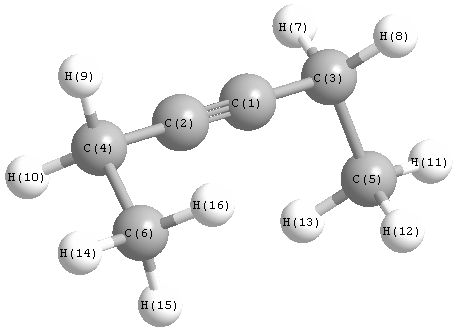
picture of 3-Hexyne state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 C4 178.862 C1 C3 C5 112.131
C1 C3 H7 109.045 C1 C3 H8 109.045
C2 C1 C3 178.862 C2 C4 C6 112.131
C2 C4 H9 109.045 C2 C4 H10 109.045
C3 C5 H11 110.426 C3 C5 H12 110.526
C3 C5 H13 110.526 C4 C6 H14 110.426
C4 C6 H15 110.526 C4 C6 H16 110.526
C5 C3 H7 109.829 C5 C3 H8 109.829
C6 C4 H9 109.829 C6 C4 H10 109.829
H7 C3 H8 106.812 H9 C4 H10 106.812
H11 C5 H12 108.554 H11 C5 H13 108.554
H12 C5 H13 108.184 H14 C6 H15 108.554
H14 C6 H16 108.554 H15 C6 H15 0.000
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability