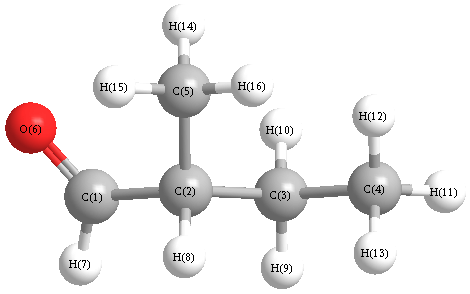
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3163 |
3031 |
16.39 |
|
|
|
| 2 |
A |
3152 |
3020 |
22.44 |
|
|
|
| 3 |
A |
3145 |
3014 |
30.69 |
|
|
|
| 4 |
A |
3137 |
3006 |
34.86 |
|
|
|
| 5 |
A |
3089 |
2960 |
18.24 |
|
|
|
| 6 |
A |
3071 |
2943 |
15.31 |
|
|
|
| 7 |
A |
3057 |
2930 |
25.72 |
|
|
|
| 8 |
A |
3047 |
2920 |
28.45 |
|
|
|
| 9 |
A |
3005 |
2879 |
18.01 |
|
|
|
| 10 |
A |
2897 |
2777 |
112.67 |
|
|
|
| 11 |
A |
1828 |
1752 |
203.93 |
|
|
|
| 12 |
A |
1487 |
1425 |
3.67 |
|
|
|
| 13 |
A |
1483 |
1421 |
11.96 |
|
|
|
| 14 |
A |
1477 |
1415 |
5.49 |
|
|
|
| 15 |
A |
1471 |
1409 |
8.77 |
|
|
|
| 16 |
A |
1465 |
1404 |
1.00 |
|
|
|
| 17 |
A |
1420 |
1361 |
8.74 |
|
|
|
| 18 |
A |
1401 |
1342 |
3.84 |
|
|
|
| 19 |
A |
1394 |
1336 |
3.71 |
|
|
|
| 20 |
A |
1364 |
1308 |
3.11 |
|
|
|
| 21 |
A |
1353 |
1297 |
2.07 |
|
|
|
| 22 |
A |
1300 |
1246 |
2.51 |
|
|
|
| 23 |
A |
1260 |
1208 |
0.65 |
|
|
|
| 24 |
A |
1190 |
1140 |
2.02 |
|
|
|
| 25 |
A |
1172 |
1123 |
2.86 |
|
|
|
| 26 |
A |
1119 |
1072 |
1.55 |
|
|
|
| 27 |
A |
1057 |
1013 |
0.36 |
|
|
|
| 28 |
A |
1019 |
977 |
13.60 |
|
|
|
| 29 |
A |
970 |
930 |
7.17 |
|
|
|
| 30 |
A |
939 |
900 |
9.07 |
|
|
|
| 31 |
A |
916 |
878 |
13.45 |
|
|
|
| 32 |
A |
798 |
765 |
6.22 |
|
|
|
| 33 |
A |
775 |
743 |
2.60 |
|
|
|
| 34 |
A |
659 |
632 |
7.64 |
|
|
|
| 35 |
A |
404 |
387 |
0.85 |
|
|
|
| 36 |
A |
389 |
372 |
0.82 |
|
|
|
| 37 |
A |
281 |
269 |
5.41 |
|
|
|
| 38 |
A |
265 |
254 |
0.48 |
|
|
|
| 39 |
A |
224 |
214 |
2.49 |
|
|
|
| 40 |
A |
189 |
181 |
0.43 |
|
|
|
| 41 |
A |
90 |
86 |
3.45 |
|
|
|
| 42 |
A |
73 |
70 |
3.86 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 30995.7 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 29703.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.997 |
|
|
|
| 2 |
C |
-0.393 |
|
|
|
| 3 |
C |
0.759 |
|
|
|
| 4 |
C |
0.394 |
|
|
|
| 5 |
C |
0.644 |
|
|
|
| 6 |
O |
-0.555 |
|
|
|
| 7 |
H |
-0.229 |
|
|
|
| 8 |
H |
-0.248 |
|
|
|
| 9 |
H |
-0.361 |
|
|
|
| 10 |
H |
-0.232 |
|
|
|
| 11 |
H |
-0.132 |
|
|
|
| 12 |
H |
-0.150 |
|
|
|
| 13 |
H |
-0.117 |
|
|
|
| 14 |
H |
-0.080 |
|
|
|
| 15 |
H |
-0.139 |
|
|
|
| 16 |
H |
-0.158 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
2.798 |
-0.465 |
0.860 |
2.964 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-46.822 |
-0.088 |
-1.979 |
| y |
-0.088 |
-37.305 |
-0.316 |
| z |
-1.979 |
-0.316 |
-37.583 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-9.378 |
-0.088 |
-1.979 |
| y |
-0.088 |
4.897 |
-0.316 |
| z |
-1.979 |
-0.316 |
4.481 |
|
| Polar |
|---|
| 3z2-r2 | 8.961 |
|---|
| x2-y2 | -9.517 |
|---|
| xy | -0.088 |
|---|
| xz | -1.979 |
|---|
| yz | -0.316 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.367 |
-0.176 |
0.281 |
| y |
-0.176 |
9.741 |
-0.237 |
| z |
0.281 |
-0.237 |
8.303 |
<r2> (average value of r
2) Å
2
| <r2> |
203.725 |
| (<r2>)1/2 |
14.273 |