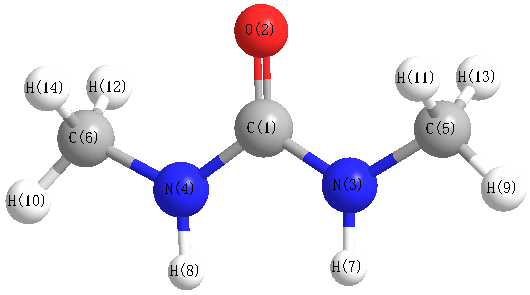
Vibrational Frequencies calculated at HF/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3851 |
3507 |
39.19 |
|
|
|
| 2 |
A |
3298 |
3003 |
19.33 |
|
|
|
| 3 |
A |
3249 |
2959 |
20.75 |
|
|
|
| 4 |
A |
3178 |
2894 |
5.96 |
|
|
|
| 5 |
A |
1903 |
1733 |
364.55 |
|
|
|
| 6 |
A |
1646 |
1499 |
32.64 |
|
|
|
| 7 |
A |
1601 |
1458 |
0.17 |
|
|
|
| 8 |
A |
1588 |
1446 |
0.71 |
|
|
|
| 9 |
A |
1559 |
1420 |
4.33 |
|
|
|
| 10 |
A |
1299 |
1183 |
0.16 |
|
|
|
| 11 |
A |
1262 |
1149 |
1.89 |
|
|
|
| 12 |
A |
1228 |
1118 |
6.47 |
|
|
|
| 13 |
A |
993 |
904 |
1.57 |
|
|
|
| 14 |
A |
562 |
512 |
63.60 |
|
|
|
| 15 |
A |
443 |
403 |
56.53 |
|
|
|
| 16 |
A |
237 |
216 |
5.57 |
|
|
|
| 17 |
A |
209 |
190 |
0.18 |
|
|
|
| 18 |
A |
131 |
120 |
0.48 |
|
|
|
| 19 |
B |
3847 |
3503 |
9.93 |
|
|
|
| 20 |
B |
3298 |
3003 |
13.26 |
|
|
|
| 21 |
B |
3249 |
2959 |
63.98 |
|
|
|
| 22 |
B |
3177 |
2893 |
121.66 |
|
|
|
| 23 |
B |
1706 |
1553 |
469.64 |
|
|
|
| 24 |
B |
1609 |
1465 |
0.16 |
|
|
|
| 25 |
B |
1587 |
1445 |
36.44 |
|
|
|
| 26 |
B |
1564 |
1424 |
5.31 |
|
|
|
| 27 |
B |
1370 |
1248 |
256.40 |
|
|
|
| 28 |
B |
1252 |
1140 |
17.72 |
|
|
|
| 29 |
B |
1233 |
1123 |
10.30 |
|
|
|
| 30 |
B |
1105 |
1006 |
4.39 |
|
|
|
| 31 |
B |
862 |
785 |
40.62 |
|
|
|
| 32 |
B |
785 |
715 |
22.60 |
|
|
|
| 33 |
B |
520 |
473 |
106.92 |
|
|
|
| 34 |
B |
341 |
311 |
43.73 |
|
|
|
| 35 |
B |
150 |
136 |
0.61 |
|
|
|
| 36 |
B |
128 |
117 |
9.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 28009.7 cm
-1
Scaled (by 0.9106) Zero Point Vibrational Energy (zpe) 25505.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at HF/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.992 |
|
|
|
| 2 |
O |
-0.692 |
|
|
|
| 3 |
N |
-0.425 |
|
|
|
| 4 |
N |
-0.425 |
|
|
|
| 5 |
C |
0.794 |
|
|
|
| 6 |
C |
0.794 |
|
|
|
| 7 |
H |
-0.141 |
|
|
|
| 8 |
H |
-0.141 |
|
|
|
| 9 |
H |
-0.203 |
|
|
|
| 10 |
H |
-0.203 |
|
|
|
| 11 |
H |
-0.107 |
|
|
|
| 12 |
H |
-0.107 |
|
|
|
| 13 |
H |
-0.067 |
|
|
|
| 14 |
H |
-0.067 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.806 |
3.806 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.828 |
-2.060 |
0.000 |
| y |
-2.060 |
-30.549 |
0.000 |
| z |
0.000 |
0.000 |
-38.201 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.453 |
-2.060 |
0.000 |
| y |
-2.060 |
7.466 |
0.000 |
| z |
0.000 |
0.000 |
-4.013 |
|
| Polar |
|---|
| 3z2-r2 | -8.026 |
|---|
| x2-y2 | -7.279 |
|---|
| xy | -2.060 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.385 |
-0.344 |
0.000 |
| y |
-0.344 |
9.787 |
0.000 |
| z |
0.000 |
0.000 |
7.933 |
<r2> (average value of r
2) Å
2
| <r2> |
190.994 |
| (<r2>)1/2 |
13.820 |