return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for NH(CH3)CONH(CH3) (Urea, N,N'-dimethyl-)

using model chemistry: B2PLYP=FULLultrafine/aug-cc-pVDZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2 1A
Energy calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
 hartrees
Energy at 0K-303.633117
Energy at 298.15K 
HF Energy-303.302941
Nuclear repulsion energy247.392281
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A 3621 3484 23.36      
2 A 3178 3058 11.68      
3 A 3134 3016 8.34      
4 A 3050 2935 5.86      
5 A 1739 1673 258.38      
6 A 1521 1464 18.32      
7 A 1484 1428 0.71      
8 A 1472 1416 2.63      
9 A 1439 1385 7.73      
10 A 1202 1157 0.13      
11 A 1177 1133 2.07      
12 A 1138 1095 6.31      
13 A 921 886 1.56      
14 A 528 508 53.02      
15 A 409 394 53.03      
16 A 221 213 4.73      
17 A 186 179 0.12      
18 A 106 102 0.38      
19 B 3618 3481 8.34      
20 B 3178 3058 5.36      
21 B 3134 3016 44.12      
22 B 3049 2934 110.69      
23 B 1565 1506 369.95      
24 B 1499 1443 1.82      
25 B 1475 1420 52.77      
26 B 1443 1389 7.12      
27 B 1267 1219 211.76      
28 B 1155 1111 24.18      
29 B 1141 1098 4.70      
30 B 1047 1008 1.45      
31 B 761 733 15.06      
32 B 725 697 22.07      
33 B 474 456 104.64      
34 B 315 303 42.72      
35 B 123 118 7.81      
36 B 120 115 2.44      

Unscaled Zero Point Vibrational Energy (zpe) 26307.8 cm-1
Scaled (by 0.9623) Zero Point Vibrational Energy (zpe) 25316.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at B2PLYP=FULLultrafine/aug-cc-pVDZ
ABC
0.32547 0.07198 0.06049

See section I.F.4 to change rotational constant units
Geometric Data calculated at B2PLYP=FULLultrafine/aug-cc-pVDZ

Point Group is C2

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 0.088
O2 0.000 0.000 1.318
N3 0.000 1.164 -0.665
N4 0.000 -1.164 -0.665
C5 -0.231 2.431 0.020
C6 0.231 -2.431 0.020
H7 -0.392 1.085 -1.594
H8 0.392 -1.085 -1.594
H9 -0.037 3.249 -0.683
H10 0.037 -3.249 -0.683
H11 0.464 2.511 0.861
H12 -0.464 -2.511 0.861
H13 -1.255 2.521 0.412
H14 1.255 -2.521 0.412

Atom - Atom Distances (Å)
  C1 O2 N3 N4 C5 C6 H7 H8 H9 H10 H11 H12 H13 H14
C11.23051.38651.38652.44312.44312.03912.03913.33913.33912.66842.66842.83432.8343
O21.23052.29972.29972.76572.76573.13213.13213.81593.81592.59432.59432.95802.9580
N31.38652.29972.32891.45853.66761.01122.46492.08464.41332.08774.00692.13854.0390
N41.38652.29972.32893.66761.45852.46491.01124.41332.08464.00692.08774.03902.1385
C52.44312.76571.45853.66764.88432.10793.91861.09565.72951.09395.01901.10035.1847
C62.44312.76573.66761.45854.88433.91862.10795.72951.09565.01901.09395.18471.1003
H72.03913.13211.01122.46492.10793.91862.30682.37414.44892.96554.35492.61324.4422
H82.03913.13212.46491.01123.91862.10792.30684.44892.37414.35492.96554.44222.6132
H93.33913.81592.08464.41331.09565.72952.37414.44896.49771.78355.97871.79246.0127
H103.33913.81594.41332.08465.72951.09564.44892.37416.49775.97871.78356.01271.7924
H112.66842.59432.08774.00691.09395.01902.96554.35491.78355.97875.10751.77695.1134
H122.66842.59434.00692.08775.01901.09394.35492.96555.97871.78355.10755.11341.7769
H132.83432.95802.13854.03901.10035.18472.61324.44221.79246.01271.77695.11345.6314
H142.83432.95804.03902.13855.18471.10034.44222.61326.01271.79245.11341.77695.6314

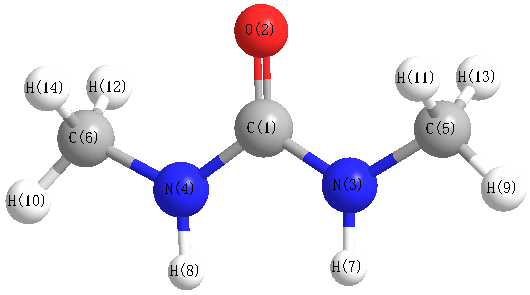
picture of Urea, N,N'-dimethyl- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 N3 C5 118.328 C1 N3 H7 115.631
C1 N4 C6 118.328 C1 N4 H8 115.631
O2 C1 N3 122.876 O2 C1 N4 122.876
N3 C1 N4 114.248 N3 C5 H9 108.574
N3 C5 H11 108.921 N3 C5 H13 112.636
N4 C6 H10 108.574 N4 C6 H12 108.921
N4 C6 H14 112.636 C5 N3 H7 116.007
C6 N4 H8 116.007 H9 C5 H11 109.089
H9 C5 H13 109.415 H10 C6 H12 109.089
H10 C6 H14 109.415 H11 C5 H13 108.153
H12 C6 H14 108.153
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability