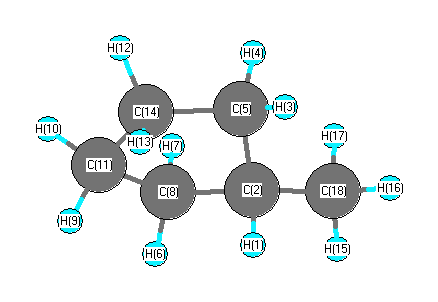
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3136 |
3005 |
93.92 |
|
|
|
| 2 |
A |
3129 |
2998 |
31.41 |
|
|
|
| 3 |
A |
3121 |
2991 |
34.76 |
|
|
|
| 4 |
A |
3118 |
2988 |
13.89 |
|
|
|
| 5 |
A |
3113 |
2983 |
4.81 |
|
|
|
| 6 |
A |
3105 |
2975 |
44.17 |
|
|
|
| 7 |
A |
3084 |
2955 |
51.36 |
|
|
|
| 8 |
A |
3070 |
2942 |
22.71 |
|
|
|
| 9 |
A |
3052 |
2925 |
76.34 |
|
|
|
| 10 |
A |
3044 |
2917 |
24.62 |
|
|
|
| 11 |
A |
3042 |
2915 |
18.89 |
|
|
|
| 12 |
A |
3024 |
2898 |
8.17 |
|
|
|
| 13 |
A |
1511 |
1448 |
1.85 |
|
|
|
| 14 |
A |
1486 |
1424 |
9.91 |
|
|
|
| 15 |
A |
1485 |
1423 |
3.16 |
|
|
|
| 16 |
A |
1479 |
1417 |
3.25 |
|
|
|
| 17 |
A |
1475 |
1414 |
4.22 |
|
|
|
| 18 |
A |
1471 |
1410 |
0.09 |
|
|
|
| 19 |
A |
1402 |
1343 |
5.30 |
|
|
|
| 20 |
A |
1385 |
1327 |
0.80 |
|
|
|
| 21 |
A |
1343 |
1287 |
0.96 |
|
|
|
| 22 |
A |
1331 |
1275 |
0.05 |
|
|
|
| 23 |
A |
1324 |
1269 |
0.15 |
|
|
|
| 24 |
A |
1311 |
1256 |
0.93 |
|
|
|
| 25 |
A |
1287 |
1233 |
0.03 |
|
|
|
| 26 |
A |
1258 |
1205 |
0.31 |
|
|
|
| 27 |
A |
1245 |
1193 |
0.49 |
|
|
|
| 28 |
A |
1216 |
1165 |
0.28 |
|
|
|
| 29 |
A |
1201 |
1151 |
0.31 |
|
|
|
| 30 |
A |
1170 |
1121 |
0.71 |
|
|
|
| 31 |
A |
1119 |
1072 |
0.00 |
|
|
|
| 32 |
A |
1068 |
1024 |
0.13 |
|
|
|
| 33 |
A |
1022 |
980 |
0.87 |
|
|
|
| 34 |
A |
1006 |
964 |
2.74 |
|
|
|
| 35 |
A |
974 |
933 |
0.73 |
|
|
|
| 36 |
A |
932 |
893 |
0.10 |
|
|
|
| 37 |
A |
925 |
887 |
0.17 |
|
|
|
| 38 |
A |
910 |
872 |
0.91 |
|
|
|
| 39 |
A |
864 |
828 |
1.08 |
|
|
|
| 40 |
A |
818 |
784 |
0.18 |
|
|
|
| 41 |
A |
760 |
729 |
0.60 |
|
|
|
| 42 |
A |
632 |
605 |
0.22 |
|
|
|
| 43 |
A |
535 |
512 |
0.46 |
|
|
|
| 44 |
A |
425 |
408 |
0.17 |
|
|
|
| 45 |
A |
305 |
292 |
0.03 |
|
|
|
| 46 |
A |
228 |
218 |
0.00 |
|
|
|
| 47 |
A |
178 |
170 |
0.00 |
|
|
|
| 48 |
A |
34 |
33 |
0.02 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 37074.3 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 35528.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
H |
-0.430 |
|
|
|
| 2 |
C |
-0.565 |
|
|
|
| 3 |
H |
-0.319 |
|
|
|
| 4 |
H |
-0.286 |
|
|
|
| 5 |
C |
0.836 |
|
|
|
| 6 |
H |
-0.319 |
|
|
|
| 7 |
H |
-0.287 |
|
|
|
| 8 |
C |
0.834 |
|
|
|
| 9 |
H |
-0.271 |
|
|
|
| 10 |
H |
-0.239 |
|
|
|
| 11 |
C |
0.548 |
|
|
|
| 12 |
H |
-0.240 |
|
|
|
| 13 |
H |
-0.274 |
|
|
|
| 14 |
C |
0.546 |
|
|
|
| 15 |
H |
-0.100 |
|
|
|
| 16 |
H |
-0.101 |
|
|
|
| 17 |
H |
-0.099 |
|
|
|
| 18 |
C |
0.764 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.009 |
-0.001 |
-0.101 |
0.101 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.092 |
-0.000 |
-0.299 |
| y |
-0.000 |
-40.520 |
-0.001 |
| z |
-0.299 |
-0.001 |
-39.251 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.207 |
-0.000 |
-0.299 |
| y |
-0.000 |
-0.349 |
-0.001 |
| z |
-0.299 |
-0.001 |
1.555 |
|
| Polar |
|---|
| 3z2-r2 | 3.111 |
|---|
| x2-y2 | -0.572 |
|---|
| xy | -0.000 |
|---|
| xz | -0.299 |
|---|
| yz | -0.001 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
11.897 |
0.000 |
0.063 |
| y |
0.000 |
10.713 |
0.000 |
| z |
0.063 |
0.000 |
9.419 |
<r2> (average value of r
2) Å
2
| <r2> |
178.839 |
| (<r2>)1/2 |
13.373 |