Jump to
S1C2
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -343.309564 |
| Energy at 298.15K | -343.314461 |
| HF Energy | -343.309564 |
| Nuclear repulsion energy | 271.784681 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3311 |
3173 |
0.01 |
162.28 |
0.11 |
0.20 |
| 2 |
A' |
3291 |
3154 |
0.25 |
48.16 |
0.36 |
0.53 |
| 3 |
A' |
3280 |
3143 |
1.00 |
82.31 |
0.70 |
0.83 |
| 4 |
A' |
2946 |
2823 |
89.67 |
142.44 |
0.30 |
0.47 |
| 5 |
A' |
1791 |
1716 |
398.08 |
177.26 |
0.32 |
0.49 |
| 6 |
A' |
1625 |
1558 |
12.28 |
8.57 |
0.47 |
0.64 |
| 7 |
A' |
1529 |
1465 |
48.46 |
130.93 |
0.23 |
0.37 |
| 8 |
A' |
1446 |
1386 |
39.30 |
39.09 |
0.46 |
0.63 |
| 9 |
A' |
1390 |
1332 |
0.05 |
21.47 |
0.11 |
0.20 |
| 10 |
A' |
1325 |
1269 |
43.69 |
8.77 |
0.44 |
0.61 |
| 11 |
A' |
1248 |
1196 |
1.46 |
11.81 |
0.29 |
0.45 |
| 12 |
A' |
1205 |
1155 |
7.23 |
4.02 |
0.09 |
0.16 |
| 13 |
A' |
1118 |
1071 |
11.69 |
14.72 |
0.19 |
0.31 |
| 14 |
A' |
1041 |
998 |
45.36 |
4.67 |
0.41 |
0.58 |
| 15 |
A' |
958 |
918 |
17.14 |
7.74 |
0.08 |
0.14 |
| 16 |
A' |
900 |
863 |
11.70 |
6.18 |
0.74 |
0.85 |
| 17 |
A' |
770 |
738 |
63.47 |
1.36 |
0.64 |
0.78 |
| 18 |
A' |
504 |
483 |
1.10 |
8.57 |
0.21 |
0.35 |
| 19 |
A' |
203 |
194 |
6.19 |
0.95 |
0.67 |
0.80 |
| 20 |
A" |
1010 |
968 |
0.14 |
1.11 |
0.75 |
0.86 |
| 21 |
A" |
893 |
856 |
2.28 |
0.74 |
0.75 |
0.86 |
| 22 |
A" |
846 |
811 |
2.72 |
1.02 |
0.75 |
0.86 |
| 23 |
A" |
777 |
745 |
68.91 |
1.27 |
0.75 |
0.86 |
| 24 |
A" |
660 |
632 |
0.24 |
0.80 |
0.75 |
0.86 |
| 25 |
A" |
610 |
585 |
8.67 |
0.16 |
0.75 |
0.86 |
| 26 |
A" |
299 |
287 |
13.95 |
0.35 |
0.75 |
0.86 |
| 27 |
A" |
136 |
130 |
1.49 |
1.17 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 17555.4 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 16823.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.518 |
-0.894 |
0.000 |
| C2 |
-0.521 |
-1.748 |
0.000 |
| C3 |
-1.713 |
-1.077 |
0.000 |
| C4 |
0.000 |
0.364 |
0.000 |
| C5 |
-1.373 |
0.303 |
0.000 |
| C6 |
0.891 |
1.512 |
0.000 |
| O7 |
2.103 |
1.464 |
0.000 |
| H8 |
-0.270 |
-2.801 |
0.000 |
| H9 |
-2.701 |
-1.520 |
0.000 |
| H10 |
-2.046 |
1.153 |
0.000 |
| H11 |
0.346 |
2.483 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3445 | 2.2377 | 1.3601 | 2.2370 | 2.4343 | 2.8406 | 2.0639 | 3.2793 | 3.2803 | 3.3805 |
C2 | 1.3445 | | 1.3673 | 2.1750 | 2.2202 | 3.5520 | 4.1466 | 1.0833 | 2.1922 | 3.2770 | 4.3180 | C3 | 2.2377 | 1.3673 | | 2.2382 | 1.4211 | 3.6715 | 4.5836 | 2.2483 | 1.0836 | 2.2548 | 4.1118 | C4 | 1.3601 | 2.1750 | 2.2382 | | 1.3739 | 1.4529 | 2.3727 | 3.1770 | 3.2936 | 2.1927 | 2.1465 | C5 | 2.2370 | 2.2202 | 1.4211 | 1.3739 | | 2.5661 | 3.6638 | 3.2943 | 2.2560 | 1.0845 | 2.7756 | C6 | 2.4343 | 3.5520 | 3.6715 | 1.4529 | 2.5661 | | 1.2125 | 4.4667 | 4.7008 | 2.9586 | 1.1133 | O7 | 2.8406 | 4.1466 | 4.5836 | 2.3727 | 3.6638 | 1.2125 | | 4.8805 | 5.6552 | 4.1600 | 2.0308 | H8 | 2.0639 | 1.0833 | 2.2483 | 3.1770 | 3.2943 | 4.4667 | 4.8805 | | 2.7482 | 4.3349 | 5.3197 | H9 | 3.2793 | 2.1922 | 1.0836 | 3.2936 | 2.2560 | 4.7008 | 5.6552 | 2.7482 | | 2.7525 | 5.0307 | H10 | 3.2803 | 3.2770 | 2.2548 | 2.1927 | 1.0845 | 2.9586 | 4.1600 | 4.3349 | 2.7525 | | 2.7363 | H11 | 3.3805 | 4.3180 | 4.1118 | 2.1465 | 2.7756 | 1.1133 | 2.0308 | 5.3197 | 5.0307 | 2.7363 | |
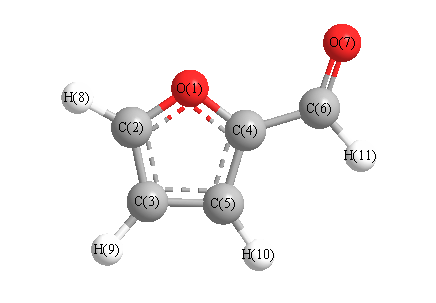 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
111.204 |
|
O1 |
C2 |
H8 |
116.032 |
| O1 |
C4 |
C5 |
109.814 |
|
O1 |
C4 |
C6 |
119.809 |
| C2 |
O1 |
C4 |
107.059 |
|
C2 |
C3 |
C5 |
105.526 |
| C2 |
C3 |
H9 |
126.484 |
|
C3 |
C2 |
H8 |
132.764 |
| C3 |
C5 |
C4 |
106.398 |
|
C3 |
C5 |
H10 |
127.783 |
| C4 |
C5 |
H10 |
125.820 |
|
C4 |
C6 |
O7 |
125.546 |
| C4 |
C6 |
H11 |
112.862 |
|
C5 |
C3 |
H9 |
127.990 |
| C5 |
C4 |
C6 |
130.377 |
|
O7 |
C6 |
H11 |
121.592 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.586 |
|
|
|
| 2 |
C |
0.148 |
|
|
|
| 3 |
C |
0.776 |
|
|
|
| 4 |
C |
0.427 |
|
|
|
| 5 |
C |
0.649 |
|
|
|
| 6 |
C |
0.666 |
|
|
|
| 7 |
O |
-0.577 |
|
|
|
| 8 |
H |
-0.412 |
|
|
|
| 9 |
H |
-0.399 |
|
|
|
| 10 |
H |
-0.491 |
|
|
|
| 11 |
H |
-0.199 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-4.194 |
-1.251 |
0.000 |
4.377 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-44.368 |
-3.257 |
0.000 |
| y |
-3.257 |
-33.958 |
0.000 |
| z |
0.000 |
0.000 |
-41.342 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-6.718 |
-3.257 |
0.000 |
| y |
-3.257 |
8.897 |
0.000 |
| z |
0.000 |
0.000 |
-2.179 |
|
| Polar |
|---|
| 3z2-r2 | -4.358 |
|---|
| x2-y2 | -10.410 |
|---|
| xy | -3.257 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
12.045 |
2.200 |
0.000 |
| y |
2.200 |
12.079 |
0.000 |
| z |
0.000 |
0.000 |
5.941 |
<r2> (average value of r
2) Å
2
| <r2> |
190.910 |
| (<r2>)1/2 |
13.817 |
Jump to
S1C1
Energy calculated at mPW1PW91/aug-cc-pVDZ
| | hartrees |
|---|
| Energy at 0K | -343.310266 |
| Energy at 298.15K | -343.315127 |
| HF Energy | -343.310266 |
| Nuclear repulsion energy | 270.983455 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at mPW1PW91/aug-cc-pVDZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3311 |
3173 |
0.07 |
172.47 |
0.11 |
0.20 |
| 2 |
A' |
3299 |
3162 |
1.22 |
27.48 |
0.41 |
0.58 |
| 3 |
A' |
3283 |
3146 |
0.63 |
84.05 |
0.73 |
0.84 |
| 4 |
A' |
2963 |
2840 |
82.84 |
151.58 |
0.30 |
0.46 |
| 5 |
A' |
1784 |
1709 |
324.28 |
137.25 |
0.35 |
0.52 |
| 6 |
A' |
1635 |
1567 |
72.69 |
27.26 |
0.37 |
0.54 |
| 7 |
A' |
1520 |
1457 |
114.81 |
218.74 |
0.24 |
0.39 |
| 8 |
A' |
1451 |
1391 |
1.46 |
9.56 |
0.75 |
0.86 |
| 9 |
A' |
1401 |
1343 |
13.59 |
26.57 |
0.36 |
0.53 |
| 10 |
A' |
1273 |
1220 |
25.26 |
5.67 |
0.23 |
0.37 |
| 11 |
A' |
1247 |
1195 |
0.23 |
4.91 |
0.57 |
0.73 |
| 12 |
A' |
1193 |
1143 |
13.79 |
6.19 |
0.12 |
0.21 |
| 13 |
A' |
1125 |
1078 |
27.47 |
18.33 |
0.17 |
0.29 |
| 14 |
A' |
1035 |
992 |
42.07 |
3.24 |
0.36 |
0.53 |
| 15 |
A' |
970 |
930 |
6.82 |
8.83 |
0.12 |
0.21 |
| 16 |
A' |
900 |
863 |
7.58 |
5.02 |
0.73 |
0.85 |
| 17 |
A' |
758 |
727 |
73.86 |
3.07 |
0.53 |
0.69 |
| 18 |
A' |
500 |
479 |
1.08 |
6.26 |
0.24 |
0.39 |
| 19 |
A' |
205 |
196 |
7.12 |
0.36 |
0.27 |
0.42 |
| 20 |
A" |
1018 |
975 |
0.12 |
1.79 |
0.75 |
0.86 |
| 21 |
A" |
899 |
862 |
1.94 |
0.86 |
0.75 |
0.86 |
| 22 |
A" |
848 |
813 |
4.04 |
0.57 |
0.75 |
0.86 |
| 23 |
A" |
776 |
744 |
65.47 |
1.68 |
0.75 |
0.86 |
| 24 |
A" |
647 |
620 |
0.59 |
0.40 |
0.75 |
0.86 |
| 25 |
A" |
611 |
586 |
10.91 |
0.20 |
0.75 |
0.86 |
| 26 |
A" |
250 |
239 |
14.16 |
1.59 |
0.75 |
0.86 |
| 27 |
A" |
158 |
151 |
3.03 |
0.88 |
0.75 |
0.86 |
Unscaled Zero Point Vibrational Energy (zpe) 17529.6 cm
-1
Scaled (by 0.9583) Zero Point Vibrational Energy (zpe) 16798.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at mPW1PW91/aug-cc-pVDZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
1.241 |
-0.288 |
0.000 |
| C2 |
1.058 |
-1.620 |
0.000 |
| C3 |
-0.275 |
-1.931 |
0.000 |
| C4 |
0.000 |
0.287 |
0.000 |
| C5 |
-0.964 |
-0.689 |
0.000 |
| C6 |
-0.052 |
1.739 |
0.000 |
| O7 |
-1.086 |
2.376 |
0.000 |
| H8 |
1.955 |
-2.226 |
0.000 |
| H9 |
-0.701 |
-2.927 |
0.000 |
| H10 |
-2.033 |
-0.513 |
0.000 |
| H11 |
0.942 |
2.238 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
C5 |
C6 |
O7 |
H8 |
H9 |
H10 |
H11 |
| O1 | | 1.3441 | 2.2348 | 1.3674 | 2.2409 | 2.4047 | 3.5374 | 2.0653 | 3.2762 | 3.2815 | 2.5434 |
C2 | 1.3441 | | 1.3682 | 2.1803 | 2.2256 | 3.5378 | 4.5348 | 1.0832 | 2.1912 | 3.2831 | 3.8592 | C3 | 2.2348 | 1.3682 | | 2.2344 | 1.4201 | 3.6768 | 4.3827 | 2.2496 | 1.0838 | 2.2590 | 4.3423 | C4 | 1.3674 | 2.1803 | 2.2344 | | 1.3718 | 1.4536 | 2.3550 | 3.1838 | 3.2895 | 2.1846 | 2.1664 | C5 | 2.2409 | 2.2256 | 1.4201 | 1.3718 | | 2.5941 | 3.0679 | 3.2992 | 2.2535 | 1.0836 | 3.4928 | C6 | 2.4047 | 3.5378 | 3.6768 | 1.4536 | 2.5941 | | 1.2143 | 4.4445 | 4.7115 | 2.9992 | 1.1121 | O7 | 3.5374 | 4.5348 | 4.3827 | 2.3550 | 3.0679 | 1.2143 | | 5.5164 | 5.3175 | 3.0402 | 2.0329 | H8 | 2.0653 | 1.0832 | 2.2496 | 3.1838 | 3.2992 | 4.4445 | 5.5164 | | 2.7472 | 4.3410 | 4.5771 | H9 | 3.2762 | 2.1912 | 1.0838 | 3.2895 | 2.2535 | 4.7115 | 5.3175 | 2.7472 | | 2.7579 | 5.4199 | H10 | 3.2815 | 3.2831 | 2.2590 | 2.1846 | 1.0836 | 2.9992 | 3.0402 | 4.3410 | 2.7579 | | 4.0516 | H11 | 2.5434 | 3.8592 | 4.3423 | 2.1664 | 3.4928 | 1.1121 | 2.0329 | 4.5771 | 5.4199 | 4.0516 | |
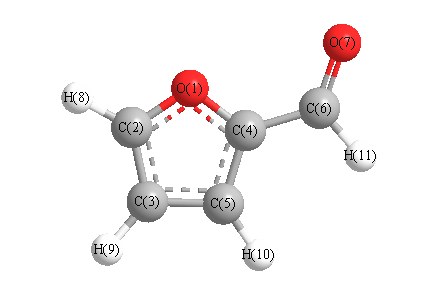 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C2 |
C3 |
110.958 |
|
O1 |
C2 |
H8 |
116.197 |
| O1 |
C4 |
C5 |
109.786 |
|
O1 |
C4 |
C6 |
116.923 |
| C2 |
O1 |
C4 |
107.040 |
|
C2 |
C3 |
C5 |
105.906 |
| C2 |
C3 |
H9 |
126.281 |
|
C3 |
C2 |
H8 |
132.846 |
| C3 |
C5 |
C4 |
106.310 |
|
C3 |
C5 |
H10 |
128.417 |
| C4 |
C5 |
H10 |
125.273 |
|
C4 |
C6 |
O7 |
123.694 |
| C4 |
C6 |
H11 |
114.560 |
|
C5 |
C3 |
H9 |
127.813 |
| C5 |
C4 |
C6 |
133.292 |
|
O7 |
C6 |
H11 |
121.747 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at mPW1PW91/aug-cc-pVDZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
O |
-0.552 |
|
|
|
| 2 |
C |
0.123 |
|
|
|
| 3 |
C |
0.839 |
|
|
|
| 4 |
C |
0.376 |
|
|
|
| 5 |
C |
0.614 |
|
|
|
| 6 |
C |
0.735 |
|
|
|
| 7 |
O |
-0.575 |
|
|
|
| 8 |
H |
-0.413 |
|
|
|
| 9 |
H |
-0.395 |
|
|
|
| 10 |
H |
-0.465 |
|
|
|
| 11 |
H |
-0.287 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.544 |
-3.401 |
0.000 |
3.735 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-38.686 |
4.333 |
0.000 |
| y |
4.333 |
-40.542 |
0.000 |
| z |
0.000 |
0.000 |
-41.297 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.234 |
4.333 |
0.000 |
| y |
4.333 |
-0.550 |
0.000 |
| z |
0.000 |
0.000 |
-1.683 |
|
| Polar |
|---|
| 3z2-r2 | -3.367 |
|---|
| x2-y2 | 1.856 |
|---|
| xy | 4.333 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
9.962 |
-1.249 |
0.000 |
| y |
-1.249 |
14.062 |
0.000 |
| z |
0.000 |
0.000 |
5.894 |
<r2> (average value of r
2) Å
2
| <r2> |
193.171 |
| (<r2>)1/2 |
13.899 |
