Jump to
S1C2
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -191.569587 |
| Energy at 298.15K | |
| HF Energy | -190.836508 |
| Nuclear repulsion energy | 103.281971 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.151 |
-0.746 |
0.000 |
| C2 |
0.000 |
0.723 |
0.000 |
| C3 |
1.212 |
1.279 |
0.000 |
| O4 |
-1.213 |
-1.317 |
0.000 |
| H5 |
0.797 |
-1.314 |
0.000 |
| H6 |
-0.910 |
1.308 |
0.000 |
| H7 |
1.351 |
2.350 |
0.000 |
| H8 |
2.103 |
0.662 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4768 | 2.4406 | 1.2060 | 1.1045 | 2.1902 | 3.4416 | 2.6577 |
C2 | 1.4768 | | 1.3331 | 2.3732 | 2.1867 | 1.0819 | 2.1152 | 2.1037 | C3 | 2.4406 | 1.3331 | | 3.5520 | 2.6252 | 2.1217 | 1.0806 | 1.0834 | O4 | 1.2060 | 2.3732 | 3.5520 | | 2.0102 | 2.6424 | 4.4748 | 3.8619 | H5 | 1.1045 | 2.1867 | 2.6252 | 2.0102 | | 3.1283 | 3.7055 | 2.3686 | H6 | 2.1902 | 1.0819 | 2.1217 | 2.6424 | 3.1283 | | 2.4897 | 3.0811 | H7 | 3.4416 | 2.1152 | 1.0806 | 4.4748 | 3.7055 | 2.4897 | | 1.8475 | H8 | 2.6577 | 2.1037 | 1.0834 | 3.8619 | 2.3686 | 3.0811 | 1.8475 | |
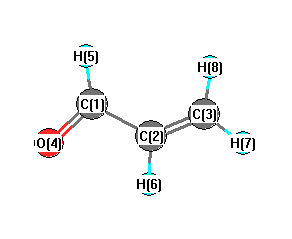 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
120.505 |
|
C1 |
C2 |
H6 |
116.891 |
| C2 |
C1 |
O4 |
124.097 |
|
C2 |
C1 |
H5 |
115.041 |
| C2 |
C3 |
H7 |
122.064 |
|
C2 |
C3 |
H8 |
120.695 |
| C3 |
C2 |
H6 |
122.605 |
|
O4 |
C1 |
H5 |
120.862 |
| H7 |
C3 |
H8 |
117.241 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -191.566058 |
| Energy at 298.15K | |
| HF Energy | -190.832483 |
| Nuclear repulsion energy | 104.809238 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3274 |
3133 |
1.42 |
|
|
|
| 2 |
A' |
3205 |
3067 |
8.09 |
|
|
|
| 3 |
A' |
3176 |
3039 |
3.38 |
|
|
|
| 4 |
A' |
2988 |
2860 |
101.27 |
|
|
|
| 5 |
A' |
1832 |
1753 |
113.79 |
|
|
|
| 6 |
A' |
1714 |
1640 |
29.32 |
|
|
|
| 7 |
A' |
1458 |
1396 |
37.65 |
|
|
|
| 8 |
A' |
1449 |
1387 |
0.34 |
|
|
|
| 9 |
A' |
1327 |
1270 |
1.41 |
|
|
|
| 10 |
A' |
1080 |
1034 |
3.83 |
|
|
|
| 11 |
A' |
946 |
905 |
55.33 |
|
|
|
| 12 |
A' |
686 |
657 |
15.20 |
|
|
|
| 13 |
A' |
288 |
275 |
6.79 |
|
|
|
| 14 |
A" |
1043 |
998 |
0.86 |
|
|
|
| 15 |
A" |
1026 |
982 |
39.78 |
|
|
|
| 16 |
A" |
1009 |
965 |
8.92 |
|
|
|
| 17 |
A" |
564 |
540 |
10.36 |
|
|
|
| 18 |
A" |
115 |
110 |
6.61 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13589.6 cm
-1
Scaled (by 0.957) Zero Point Vibrational Energy (zpe) 13005.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.889 |
-0.297 |
0.000 |
| C2 |
0.000 |
0.896 |
0.000 |
| C3 |
1.327 |
0.778 |
0.000 |
| O4 |
-0.492 |
-1.436 |
0.000 |
| H5 |
-1.971 |
-0.087 |
0.000 |
| H6 |
-0.481 |
1.866 |
0.000 |
| H7 |
1.971 |
1.646 |
0.000 |
| H8 |
1.786 |
-0.202 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
O4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.4877 | 2.4628 | 1.2062 | 1.1022 | 2.2011 | 3.4573 | 2.6764 |
C2 | 1.4877 | | 1.3323 | 2.3832 | 2.2026 | 1.0826 | 2.1088 | 2.0963 | C3 | 2.4628 | 1.3323 | | 2.8653 | 3.4096 | 2.1100 | 1.0807 | 1.0819 | O4 | 1.2062 | 2.3832 | 2.8653 | | 2.0015 | 3.3019 | 3.9450 | 2.5906 | H5 | 1.1022 | 2.2026 | 3.4096 | 2.0015 | | 2.4568 | 4.3062 | 3.7585 | H6 | 2.2011 | 1.0826 | 2.1100 | 3.3019 | 2.4568 | | 2.4617 | 3.0682 | H7 | 3.4573 | 2.1088 | 1.0807 | 3.9450 | 4.3062 | 2.4617 | | 1.8568 | H8 | 2.6764 | 2.0963 | 1.0819 | 2.5906 | 3.7585 | 3.0682 | 1.8568 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
121.608 |
|
C1 |
C2 |
H6 |
116.944 |
| C2 |
C1 |
O4 |
124.095 |
|
C2 |
C1 |
H5 |
115.729 |
| C2 |
C3 |
H7 |
121.500 |
|
C2 |
C3 |
H8 |
120.174 |
| C3 |
C2 |
H6 |
121.448 |
|
O4 |
C1 |
H5 |
120.176 |
| H7 |
C3 |
H8 |
118.326 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
