Jump to
S1C2
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -3110.900637 |
| Energy at 298.15K | |
| HF Energy | -3110.080508 |
| Nuclear repulsion energy | 290.191965 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.641 |
0.000 |
| C2 |
1.223 |
-0.245 |
0.000 |
| Br3 |
-1.587 |
-0.469 |
0.000 |
| Cl4 |
2.692 |
0.780 |
0.000 |
| H5 |
-0.044 |
1.262 |
0.888 |
| H6 |
-0.044 |
1.262 |
-0.888 |
| H7 |
1.262 |
-0.869 |
0.887 |
| H8 |
1.262 |
-0.869 |
-0.887 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5107 | 1.9366 | 2.6954 | 1.0846 | 1.0846 | 2.1587 | 2.1587 |
C2 | 1.5107 | | 2.8191 | 1.7911 | 2.1604 | 2.1604 | 1.0855 | 1.0855 | Br3 | 1.9366 | 2.8191 | | 4.4573 | 2.4827 | 2.4827 | 3.0099 | 3.0099 | Cl4 | 2.6954 | 1.7911 | 4.4573 | | 2.9168 | 2.9168 | 2.3569 | 2.3569 | H5 | 1.0846 | 2.1604 | 2.4827 | 2.9168 | | 1.7769 | 2.4993 | 3.0659 | H6 | 1.0846 | 2.1604 | 2.4827 | 2.9168 | 1.7769 | | 3.0659 | 2.4993 | H7 | 2.1587 | 1.0855 | 3.0099 | 2.3569 | 2.4993 | 3.0659 | | 1.7746 | H8 | 2.1587 | 1.0855 | 3.0099 | 2.3569 | 3.0659 | 2.4993 | 1.7746 | |
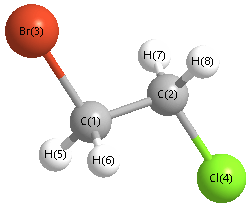 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
109.145 |
|
C1 |
C2 |
H7 |
111.448 |
| C1 |
C2 |
H8 |
111.448 |
|
C2 |
C1 |
Br3 |
109.099 |
| C2 |
C1 |
Cl4 |
38.884 |
|
C2 |
C1 |
H6 |
111.644 |
| Br3 |
C1 |
H5 |
107.122 |
|
Br3 |
C1 |
H6 |
107.122 |
| Cl4 |
C2 |
H7 |
107.489 |
|
Cl4 |
C2 |
H8 |
107.489 |
| H5 |
C1 |
H6 |
109.997 |
|
H7 |
C2 |
H8 |
109.658 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -3110.898073 |
| Energy at 298.15K | |
| HF Energy | -3110.076972 |
| Nuclear repulsion energy | 306.667485 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.002 |
1.098 |
-0.386 |
| C2 |
1.273 |
0.899 |
0.405 |
| Br3 |
-1.342 |
-0.225 |
0.036 |
| Cl4 |
2.151 |
-0.574 |
-0.087 |
| H5 |
-0.434 |
2.064 |
-0.144 |
| H6 |
0.184 |
1.033 |
-1.454 |
| H7 |
1.944 |
1.738 |
0.227 |
| H8 |
1.070 |
0.816 |
1.468 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Br3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5105 | 1.9325 | 2.7390 | 1.0869 | 1.0853 | 2.1348 | 2.1585 |
C2 | 1.5105 | | 2.8706 | 1.7841 | 2.1380 | 2.1589 | 1.0882 | 1.0860 | Br3 | 1.9325 | 2.8706 | | 3.5123 | 2.4687 | 2.4767 | 3.8323 | 2.9920 | Cl4 | 2.7390 | 1.7841 | 3.5123 | | 3.6933 | 2.8845 | 2.3424 | 2.3499 | H5 | 1.0869 | 2.1380 | 2.4687 | 3.6933 | | 1.7777 | 2.4285 | 2.5329 | H6 | 1.0853 | 2.1589 | 2.4767 | 2.8845 | 1.7777 | | 2.5340 | 3.0616 | H7 | 2.1348 | 1.0882 | 3.8323 | 2.3424 | 2.4285 | 2.5340 | | 1.7758 | H8 | 2.1585 | 1.0860 | 2.9920 | 2.3499 | 2.5329 | 3.0616 | 1.7758 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.207 |
|
C1 |
C2 |
H7 |
109.388 |
| C1 |
C2 |
H8 |
111.417 |
|
C2 |
C1 |
Br3 |
112.389 |
| C2 |
C1 |
Cl4 |
37.090 |
|
C2 |
C1 |
H6 |
111.490 |
| Br3 |
C1 |
H5 |
106.290 |
|
Br3 |
C1 |
H6 |
106.930 |
| Cl4 |
C2 |
H7 |
106.761 |
|
Cl4 |
C2 |
H8 |
107.410 |
| H5 |
C1 |
H6 |
109.853 |
|
H7 |
C2 |
H8 |
109.519 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
