Jump to
S1C2
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
Geometric Data calculated at CCD/aug-cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -997.920400 |
| Energy at 298.15K | -997.925291 |
| HF Energy | -997.134196 |
| Nuclear repulsion energy | 202.796709 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3171 |
3035 |
0.70 |
|
|
|
| 2 |
A |
3116 |
2982 |
20.62 |
|
|
|
| 3 |
A |
1506 |
1441 |
0.37 |
|
|
|
| 4 |
A |
1372 |
1313 |
15.10 |
|
|
|
| 5 |
A |
1256 |
1202 |
0.23 |
|
|
|
| 6 |
A |
1078 |
1032 |
1.19 |
|
|
|
| 7 |
A |
983 |
940 |
9.33 |
|
|
|
| 8 |
A |
695 |
665 |
14.90 |
|
|
|
| 9 |
A |
264 |
253 |
0.83 |
|
|
|
| 10 |
A |
117 |
112 |
0.82 |
|
|
|
| 11 |
B |
3183 |
3046 |
2.74 |
|
|
|
| 12 |
B |
3109 |
2975 |
3.31 |
|
|
|
| 13 |
B |
1504 |
1439 |
11.08 |
|
|
|
| 14 |
B |
1351 |
1293 |
35.32 |
|
|
|
| 15 |
B |
1190 |
1139 |
0.41 |
|
|
|
| 16 |
B |
924 |
884 |
14.91 |
|
|
|
| 17 |
B |
722 |
691 |
20.21 |
|
|
|
| 18 |
B |
418 |
400 |
6.94 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12979.3 cm
-1
Scaled (by 0.957) Zero Point Vibrational Energy (zpe) 12421.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.297 |
0.695 |
0.894 |
| C2 |
-0.297 |
-0.695 |
0.894 |
| Cl3 |
-0.297 |
1.679 |
-0.470 |
| Cl4 |
0.297 |
-1.679 |
-0.470 |
| H5 |
0.008 |
1.212 |
1.807 |
| H6 |
1.380 |
0.659 |
0.822 |
| H7 |
-0.008 |
-1.212 |
1.807 |
| H8 |
-1.380 |
-0.659 |
0.822 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
Cl4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5114 | 1.7847 | 2.7386 | 1.0879 | 1.0861 | 2.1359 | 2.1569 |
C2 | 1.5114 | | 2.7386 | 1.7847 | 2.1359 | 2.1569 | 1.0879 | 1.0861 | Cl3 | 1.7847 | 2.7386 | | 3.4110 | 2.3448 | 2.3501 | 3.6919 | 2.8832 | Cl4 | 2.7386 | 1.7847 | 3.4110 | | 3.6919 | 2.8832 | 2.3448 | 2.3501 | H5 | 1.0879 | 2.1359 | 2.3448 | 3.6919 | | 1.7770 | 2.4240 | 2.5299 | H6 | 1.0861 | 2.1569 | 2.3501 | 2.8832 | 1.7770 | | 2.5299 | 3.0591 | H7 | 2.1359 | 1.0879 | 3.6919 | 2.3448 | 2.4240 | 2.5299 | | 1.7770 | H8 | 2.1569 | 1.0861 | 2.8832 | 2.3501 | 2.5299 | 3.0591 | 1.7770 | |
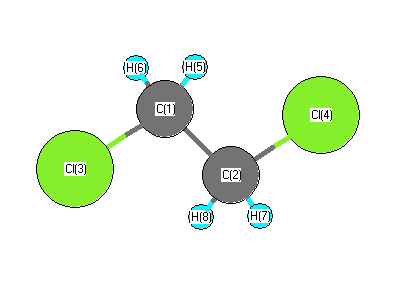 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
Cl4 |
112.113 |
|
C1 |
C2 |
H7 |
109.435 |
| C1 |
C2 |
H8 |
111.221 |
|
C2 |
C1 |
Cl3 |
112.113 |
| C2 |
C1 |
H5 |
109.435 |
|
C2 |
C1 |
H6 |
111.221 |
| Cl3 |
C1 |
H5 |
106.912 |
|
Cl3 |
C1 |
H6 |
107.385 |
| Cl4 |
C2 |
H7 |
106.912 |
|
Cl4 |
C2 |
H8 |
107.385 |
| H5 |
C1 |
H6 |
109.652 |
|
H7 |
C2 |
H8 |
109.652 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
