Jump to
S1C2
Energy calculated at B1B95/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -209.123825 |
| Energy at 298.15K | -209.129881 |
| HF Energy | -209.123825 |
| Nuclear repulsion energy | 118.116839 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3875 |
3714 |
97.80 |
|
|
|
| 2 |
A' |
3162 |
3031 |
10.30 |
|
|
|
| 3 |
A' |
3115 |
2985 |
13.34 |
|
|
|
| 4 |
A' |
3054 |
2927 |
16.98 |
|
|
|
| 5 |
A' |
1764 |
1691 |
0.95 |
|
|
|
| 6 |
A' |
1481 |
1419 |
11.57 |
|
|
|
| 7 |
A' |
1446 |
1386 |
6.94 |
|
|
|
| 8 |
A' |
1396 |
1338 |
21.98 |
|
|
|
| 9 |
A' |
1292 |
1238 |
51.96 |
|
|
|
| 10 |
A' |
1157 |
1109 |
3.64 |
|
|
|
| 11 |
A' |
1036 |
993 |
159.87 |
|
|
|
| 12 |
A' |
917 |
879 |
10.13 |
|
|
|
| 13 |
A' |
570 |
546 |
13.50 |
|
|
|
| 14 |
A' |
323 |
310 |
3.25 |
|
|
|
| 15 |
A" |
3109 |
2980 |
11.62 |
|
|
|
| 16 |
A" |
1476 |
1414 |
8.78 |
|
|
|
| 17 |
A" |
1081 |
1036 |
1.39 |
|
|
|
| 18 |
A" |
918 |
880 |
8.77 |
|
|
|
| 19 |
A" |
415 |
398 |
128.04 |
|
|
|
| 20 |
A" |
292 |
280 |
0.24 |
|
|
|
| 21 |
A" |
203 |
194 |
0.16 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16040.3 cm
-1
Scaled (by 0.9585) Zero Point Vibrational Energy (zpe) 15374.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.291 |
1.296 |
0.000 |
| C2 |
0.000 |
0.562 |
0.000 |
| N3 |
0.014 |
-0.700 |
0.000 |
| O4 |
1.300 |
-1.211 |
0.000 |
| H5 |
1.163 |
-2.158 |
0.000 |
| H6 |
-2.122 |
0.598 |
0.000 |
| H7 |
-1.363 |
1.937 |
0.878 |
| H8 |
-1.363 |
1.937 |
-0.878 |
| H9 |
0.934 |
1.121 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4848 | 2.3840 | 3.6047 | 4.2366 | 1.0854 | 1.0893 | 1.0893 | 2.2314 |
C2 | 1.4848 | | 1.2617 | 2.1981 | 2.9581 | 2.1221 | 2.1257 | 2.1257 | 1.0883 | N3 | 2.3840 | 1.2617 | | 1.3841 | 1.8567 | 2.4987 | 3.1012 | 3.1012 | 2.0399 | O4 | 3.6047 | 2.1981 | 1.3841 | | 0.9574 | 3.8703 | 4.2153 | 4.2153 | 2.3600 | H5 | 4.2366 | 2.9581 | 1.8567 | 0.9574 | | 4.2876 | 4.8907 | 4.8907 | 3.2869 | H6 | 1.0854 | 2.1221 | 2.4987 | 3.8703 | 4.2876 | | 1.7721 | 1.7721 | 3.1001 | H7 | 1.0893 | 2.1257 | 3.1012 | 4.2153 | 4.8907 | 1.7721 | | 1.7550 | 2.5906 | H8 | 1.0893 | 2.1257 | 3.1012 | 4.2153 | 4.8907 | 1.7721 | 1.7550 | | 2.5906 | H9 | 2.2314 | 1.0883 | 2.0399 | 2.3600 | 3.2869 | 3.1001 | 2.5906 | 2.5906 | |
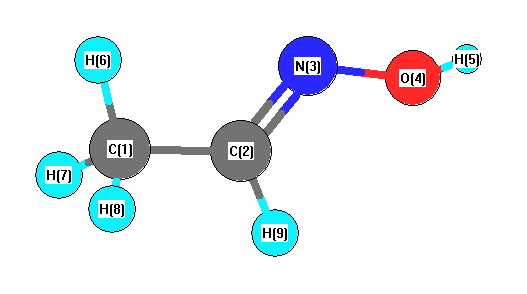 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
120.241 |
|
C1 |
C2 |
H9 |
119.478 |
| C2 |
C1 |
H6 |
110.351 |
|
C2 |
C1 |
H7 |
110.400 |
| C2 |
C1 |
H8 |
110.400 |
|
C2 |
N3 |
O4 |
112.280 |
| N3 |
C2 |
H9 |
120.281 |
|
N3 |
O4 |
H5 |
103.420 |
| H6 |
C1 |
H7 |
109.146 |
|
H6 |
C1 |
H8 |
109.146 |
| H7 |
C1 |
H8 |
107.333 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/aug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.770 |
|
|
|
| 2 |
C |
-0.247 |
|
|
|
| 3 |
N |
-0.312 |
|
|
|
| 4 |
O |
-0.307 |
|
|
|
| 5 |
H |
0.174 |
|
|
|
| 6 |
H |
0.335 |
|
|
|
| 7 |
H |
0.280 |
|
|
|
| 8 |
H |
0.280 |
|
|
|
| 9 |
H |
0.567 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.392 |
0.695 |
0.000 |
0.798 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.453 |
-2.046 |
0.000 |
| y |
-2.046 |
-18.980 |
0.000 |
| z |
0.000 |
0.000 |
-25.239 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.344 |
-2.046 |
0.000 |
| y |
-2.046 |
6.367 |
0.000 |
| z |
0.000 |
0.000 |
-3.023 |
|
| Polar |
|---|
| 3z2-r2 | -6.046 |
|---|
| x2-y2 | -6.474 |
|---|
| xy | -2.046 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.385 |
-1.410 |
0.000 |
| y |
-1.410 |
7.643 |
0.000 |
| z |
0.000 |
0.000 |
4.461 |
<r2> (average value of r
2) Å
2
| <r2> |
90.829 |
| (<r2>)1/2 |
9.530 |
Jump to
S1C1
Energy calculated at B1B95/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -209.123265 |
| Energy at 298.15K | -209.129211 |
| HF Energy | -209.123265 |
| Nuclear repulsion energy | 120.663956 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B1B95/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3880 |
3719 |
99.04 |
|
|
|
| 2 |
A' |
3193 |
3061 |
6.19 |
|
|
|
| 3 |
A' |
3172 |
3040 |
4.83 |
|
|
|
| 4 |
A' |
3062 |
2934 |
11.79 |
|
|
|
| 5 |
A' |
1768 |
1695 |
9.56 |
|
|
|
| 6 |
A' |
1474 |
1413 |
16.51 |
|
|
|
| 7 |
A' |
1400 |
1341 |
22.73 |
|
|
|
| 8 |
A' |
1380 |
1323 |
28.56 |
|
|
|
| 9 |
A' |
1344 |
1288 |
39.65 |
|
|
|
| 10 |
A' |
1144 |
1097 |
8.89 |
|
|
|
| 11 |
A' |
959 |
920 |
150.67 |
|
|
|
| 12 |
A' |
927 |
889 |
1.55 |
|
|
|
| 13 |
A' |
677 |
649 |
12.05 |
|
|
|
| 14 |
A' |
302 |
290 |
1.50 |
|
|
|
| 15 |
A" |
3114 |
2985 |
9.50 |
|
|
|
| 16 |
A" |
1483 |
1422 |
9.12 |
|
|
|
| 17 |
A" |
1064 |
1020 |
0.67 |
|
|
|
| 18 |
A" |
873 |
837 |
13.61 |
|
|
|
| 19 |
A" |
510 |
489 |
37.90 |
|
|
|
| 20 |
A" |
406 |
389 |
77.82 |
|
|
|
| 21 |
A" |
18 |
18 |
0.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16075.5 cm
-1
Scaled (by 0.9585) Zero Point Vibrational Energy (zpe) 15408.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B1B95/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-1.438 |
0.483 |
0.000 |
| C2 |
0.000 |
0.855 |
0.000 |
| N3 |
1.006 |
0.087 |
0.000 |
| O4 |
0.655 |
-1.258 |
0.000 |
| H5 |
1.506 |
-1.696 |
0.000 |
| H6 |
-1.571 |
-0.592 |
0.000 |
| H7 |
-1.931 |
0.904 |
0.875 |
| H8 |
-1.931 |
0.904 |
-0.875 |
| H9 |
0.270 |
1.904 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
O4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.4853 | 2.4759 | 2.7219 | 3.6627 | 1.0829 | 1.0896 | 1.0896 | 2.2222 |
C2 | 1.4853 | | 1.2661 | 2.2125 | 2.9631 | 2.1363 | 2.1208 | 2.1208 | 1.0831 | N3 | 2.4759 | 1.2661 | | 1.3903 | 1.8521 | 2.6655 | 3.1722 | 3.1722 | 1.9606 | O4 | 2.7219 | 2.2125 | 1.3903 | | 0.9578 | 2.3234 | 3.4826 | 3.4826 | 3.1855 | H5 | 3.6627 | 2.9631 | 1.8521 | 0.9578 | | 3.2697 | 4.3984 | 4.3984 | 3.8067 | H6 | 1.0829 | 2.1363 | 2.6655 | 2.3234 | 3.2697 | | 1.7704 | 1.7704 | 3.1019 | H7 | 1.0896 | 2.1208 | 3.1722 | 3.4826 | 4.3984 | 1.7704 | | 1.7503 | 2.5714 | H8 | 1.0896 | 2.1208 | 3.1722 | 3.4826 | 4.3984 | 1.7704 | 1.7503 | | 2.5714 | H9 | 2.2222 | 1.0831 | 1.9606 | 3.1855 | 3.8067 | 3.1019 | 2.5714 | 2.5714 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
N3 |
128.106 |
|
C1 |
C2 |
H9 |
118.983 |
| C2 |
C1 |
H6 |
111.612 |
|
C2 |
C1 |
H7 |
109.956 |
| C2 |
C1 |
H8 |
109.956 |
|
C2 |
N3 |
O4 |
112.711 |
| N3 |
C2 |
H9 |
112.911 |
|
N3 |
O4 |
H5 |
102.577 |
| H6 |
C1 |
H7 |
109.160 |
|
H6 |
C1 |
H8 |
109.160 |
| H7 |
C1 |
H8 |
106.875 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/aug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.775 |
|
|
|
| 2 |
C |
-0.306 |
|
|
|
| 3 |
N |
-0.341 |
|
|
|
| 4 |
O |
-0.263 |
|
|
|
| 5 |
H |
0.151 |
|
|
|
| 6 |
H |
0.370 |
|
|
|
| 7 |
H |
0.280 |
|
|
|
| 8 |
H |
0.280 |
|
|
|
| 9 |
H |
0.604 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-0.821 |
0.239 |
0.000 |
0.855 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-23.216 |
-3.849 |
0.000 |
| y |
-3.849 |
-21.364 |
0.000 |
| z |
0.000 |
0.000 |
-25.263 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.097 |
-3.849 |
0.000 |
| y |
-3.849 |
2.875 |
0.000 |
| z |
0.000 |
0.000 |
-2.972 |
|
| Polar |
|---|
| 3z2-r2 | -5.945 |
|---|
| x2-y2 | -1.852 |
|---|
| xy | -3.849 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.793 |
-1.083 |
0.000 |
| y |
-1.083 |
6.768 |
0.000 |
| z |
0.000 |
0.000 |
4.459 |
<r2> (average value of r
2) Å
2
| <r2> |
78.255 |
| (<r2>)1/2 |
8.846 |
