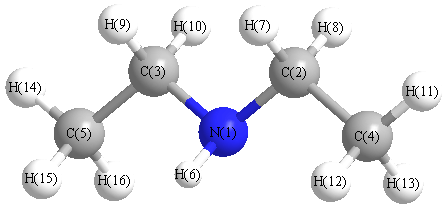
All results from a given calculation for NH(C2H5)2 (diethylamine)
using model chemistry: CCD/aug-cc-pVTZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
CS |
1A' |
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -213.385171 |
| Energy at 298.15K | |
| HF Energy | -212.396266 |
| Nuclear repulsion energy | 188.823081 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.017 |
-0.288 |
0.000 |
| C2 |
0.017 |
0.517 |
1.214 |
| C3 |
0.017 |
0.517 |
-1.214 |
| C4 |
0.017 |
-0.368 |
2.451 |
| C5 |
0.017 |
-0.368 |
-2.451 |
| H6 |
-0.798 |
-0.890 |
0.000 |
| H7 |
-0.835 |
1.212 |
1.251 |
| H8 |
0.921 |
1.131 |
1.201 |
| H9 |
-0.835 |
1.212 |
-1.251 |
| H10 |
0.921 |
1.131 |
-1.201 |
| H11 |
0.034 |
0.235 |
3.359 |
| H12 |
-0.878 |
-0.993 |
2.481 |
| H13 |
0.888 |
-1.023 |
2.448 |
| H14 |
0.034 |
0.235 |
-3.359 |
| H15 |
-0.878 |
-0.993 |
-2.481 |
| H16 |
0.888 |
-1.023 |
-2.448 |
Atom - Atom Distances (Å)
| |
N1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
| N1 | | 1.4567 | 1.4567 | 2.4519 | 2.4519 | 1.0131 | 2.1316 | 2.0675 | 2.1316 | 2.0675 | 3.3995 | 2.7302 | 2.7003 | 3.3995 | 2.7302 | 2.7003 |
C2 | 1.4567 | | 2.4281 | 1.5206 | 3.7700 | 2.0290 | 1.1009 | 1.0929 | 2.6996 | 2.6512 | 2.1635 | 2.1650 | 2.1571 | 4.5818 | 4.0909 | 4.0671 | C3 | 1.4567 | 2.4281 | | 3.7700 | 1.5206 | 2.0290 | 2.6996 | 2.6512 | 1.1009 | 1.0929 | 4.5818 | 4.0909 | 4.0671 | 2.1635 | 2.1650 | 2.1571 | C4 | 2.4519 | 1.5206 | 3.7700 | | 4.9012 | 2.6347 | 2.1596 | 2.1506 | 4.1144 | 4.0500 | 1.0905 | 1.0923 | 1.0896 | 5.8409 | 5.0510 | 5.0183 | C5 | 2.4519 | 3.7700 | 1.5206 | 4.9012 | | 2.6347 | 4.1144 | 4.0500 | 2.1596 | 2.1506 | 5.8409 | 5.0510 | 5.0183 | 1.0905 | 1.0923 | 1.0896 | H6 | 1.0131 | 2.0290 | 2.0290 | 2.6347 | 2.6347 | | 2.4467 | 2.9124 | 2.4467 | 2.9124 | 3.6385 | 2.4844 | 2.9752 | 3.6385 | 2.4844 | 2.9752 | H7 | 2.1316 | 1.1009 | 2.6996 | 2.1596 | 4.1144 | 2.4467 | | 1.7589 | 2.5023 | 3.0177 | 2.4807 | 2.5259 | 3.0660 | 4.7921 | 4.3355 | 4.6532 | H8 | 2.0675 | 1.0929 | 2.6512 | 2.1506 | 4.0500 | 2.9124 | 1.7589 | | 3.0177 | 2.4030 | 2.4993 | 3.0638 | 2.4893 | 4.7318 | 4.6163 | 4.2381 | H9 | 2.1316 | 2.6996 | 1.1009 | 4.1144 | 2.1596 | 2.4467 | 2.5023 | 3.0177 | | 1.7589 | 4.7921 | 4.3355 | 4.6532 | 2.4807 | 2.5259 | 3.0660 | H10 | 2.0675 | 2.6512 | 1.0929 | 4.0500 | 2.1506 | 2.9124 | 3.0177 | 2.4030 | 1.7589 | | 4.7318 | 4.6163 | 4.2381 | 2.4993 | 3.0638 | 2.4893 | H11 | 3.3995 | 2.1635 | 4.5818 | 1.0905 | 5.8409 | 3.6385 | 2.4807 | 2.4993 | 4.7921 | 4.7318 | | 1.7635 | 1.7729 | 6.7181 | 6.0370 | 6.0029 | H12 | 2.7302 | 2.1650 | 4.0909 | 1.0923 | 5.0510 | 2.4844 | 2.5259 | 3.0638 | 4.3355 | 4.6163 | 1.7635 | | 1.7663 | 6.0370 | 4.9619 | 5.2358 | H13 | 2.7003 | 2.1571 | 4.0671 | 1.0896 | 5.0183 | 2.9752 | 3.0660 | 2.4893 | 4.6532 | 4.2381 | 1.7729 | 1.7663 | | 6.0029 | 5.2358 | 4.8960 | H14 | 3.3995 | 4.5818 | 2.1635 | 5.8409 | 1.0905 | 3.6385 | 4.7921 | 4.7318 | 2.4807 | 2.4993 | 6.7181 | 6.0370 | 6.0029 | | 1.7635 | 1.7729 | H15 | 2.7302 | 4.0909 | 2.1650 | 5.0510 | 1.0923 | 2.4844 | 4.3355 | 4.6163 | 2.5259 | 3.0638 | 6.0370 | 4.9619 | 5.2358 | 1.7635 | | 1.7663 | H16 | 2.7003 | 4.0671 | 2.1571 | 5.0183 | 1.0896 | 2.9752 | 4.6532 | 4.2381 | 3.0660 | 2.4893 | 6.0029 | 5.2358 | 4.8960 | 1.7729 | 1.7663 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
C2 |
C4 |
110.862 |
|
N1 |
C2 |
H7 |
112.163 |
| N1 |
C2 |
H8 |
107.514 |
|
N1 |
C3 |
C5 |
110.862 |
| N1 |
C3 |
H9 |
112.163 |
|
N1 |
C3 |
H10 |
107.514 |
| C2 |
N1 |
C3 |
112.910 |
|
C2 |
N1 |
H6 |
109.160 |
| C2 |
C4 |
H11 |
110.831 |
|
C2 |
C4 |
H12 |
110.840 |
| C2 |
C4 |
H13 |
110.366 |
|
C3 |
N1 |
H6 |
109.160 |
| C3 |
C5 |
H14 |
110.831 |
|
C3 |
C5 |
H15 |
110.840 |
| C3 |
C5 |
H16 |
110.366 |
|
C4 |
C2 |
H7 |
109.898 |
| C4 |
C2 |
H8 |
109.661 |
|
C5 |
C3 |
H8 |
151.129 |
| C5 |
C3 |
H10 |
109.661 |
|
H7 |
C2 |
H8 |
106.604 |
| H9 |
C3 |
H10 |
106.604 |
|
H11 |
C4 |
H12 |
107.792 |
| H11 |
C4 |
H13 |
108.822 |
|
H12 |
C4 |
H13 |
108.098 |
| H14 |
C5 |
H15 |
107.792 |
|
H14 |
C5 |
H16 |
108.822 |
| H15 |
C5 |
H16 |
108.098 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
