return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CCO (Dicarbon monoxide)

using model chemistry: QCISD(T)/aug-cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C*V 3Σ
2 1 yes C*V 1Π

State 1 (3Σ)

Jump to S2C1
Energy calculated at QCISD(T)/aug-cc-pVTZ
 hartrees
Energy at 0K-151.027923
Energy at 298.15K-151.026289
HF Energy-150.519077
Nuclear repulsion energy45.686073
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 Σ 2006 1940        
2 Σ 1067 1031        
3 Π 376 364        
3 Π 376 363        

Unscaled Zero Point Vibrational Energy (zpe) 1912.5 cm-1
Scaled (by 0.9668) Zero Point Vibrational Energy (zpe) 1849.0 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)/aug-cc-pVTZ
B
0.38133

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)/aug-cc-pVTZ

Point Group is C∞v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 -1.427
C2 0.000 0.000 -0.055
O3 0.000 0.000 1.111

Atom - Atom Distances (Å)
  C1 C2 O3
C11.37172.5378
C21.37171.1661
O32.53781.1661

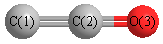
picture of Dicarbon monoxide state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 180.000
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability

State 2 (1Π)

Jump to S1C1
Energy calculated at QCISD(T)/aug-cc-pVTZ
 hartrees
Energy at 0K-150.997262
Energy at 298.15K-150.995632
HF Energy-150.457612
Nuclear repulsion energy45.473350
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
4/u)
Dep P Dep U
1 Σ 1961 1896        
2 Σ 1079 1043        
3 Π 459 444        
3 Π 308 298        

Unscaled Zero Point Vibrational Energy (zpe) 1903.3 cm-1
Scaled (by 0.9668) Zero Point Vibrational Energy (zpe) 1840.1 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at QCISD(T)/aug-cc-pVTZ
B
0.37871

See section I.F.4 to change rotational constant units
Geometric Data calculated at QCISD(T)/aug-cc-pVTZ

Point Group is C∞v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
C1 0.000 0.000 -1.429
C2 0.000 0.000 -0.060
O3 0.000 0.000 1.117

Atom - Atom Distances (Å)
  C1 C2 O3
C11.36902.5463
C21.36901.1773
O32.54631.1773

picture of Dicarbon monoxide state 2 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
C1 C2 O3 180.000
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability