Jump to
S1C2
S1C3
Energy calculated at B3PW91/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -209.205502 |
| Energy at 298.15K | -209.211575 |
| HF Energy | -209.205502 |
| Nuclear repulsion energy | 119.609661 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3613 |
3487 |
28.03 |
|
|
|
| 2 |
A' |
3122 |
3013 |
12.19 |
|
|
|
| 3 |
A' |
3028 |
2922 |
52.73 |
|
|
|
| 4 |
A' |
2931 |
2829 |
80.16 |
|
|
|
| 5 |
A' |
1793 |
1731 |
533.21 |
|
|
|
| 6 |
A' |
1528 |
1475 |
19.87 |
|
|
|
| 7 |
A' |
1480 |
1429 |
5.80 |
|
|
|
| 8 |
A' |
1459 |
1408 |
5.81 |
|
|
|
| 9 |
A' |
1388 |
1339 |
10.63 |
|
|
|
| 10 |
A' |
1307 |
1262 |
109.99 |
|
|
|
| 11 |
A' |
1166 |
1125 |
25.31 |
|
|
|
| 12 |
A' |
1013 |
977 |
41.70 |
|
|
|
| 13 |
A' |
614 |
592 |
13.42 |
|
|
|
| 14 |
A' |
341 |
329 |
7.90 |
|
|
|
| 15 |
A" |
3083 |
2975 |
22.57 |
|
|
|
| 16 |
A" |
1478 |
1427 |
6.50 |
|
|
|
| 17 |
A" |
1147 |
1107 |
0.34 |
|
|
|
| 18 |
A" |
1035 |
999 |
2.02 |
|
|
|
| 19 |
A" |
618 |
596 |
109.12 |
|
|
|
| 20 |
A" |
204 |
197 |
0.35 |
|
|
|
| 21 |
A" |
109 |
105 |
0.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16228.4 cm
-1
Scaled (by 0.9651) Zero Point Vibrational Energy (zpe) 15662.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.284 |
-0.753 |
0.000 |
| O2 |
1.399 |
-1.226 |
0.000 |
| N3 |
0.000 |
0.569 |
0.000 |
| C4 |
-1.330 |
1.125 |
0.000 |
| H5 |
-0.630 |
-1.378 |
0.000 |
| H6 |
0.795 |
1.189 |
0.000 |
| H7 |
-2.053 |
0.309 |
0.000 |
| H8 |
-1.511 |
1.736 |
0.888 |
| H9 |
-1.511 |
1.736 |
-0.888 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2115 | 1.3524 | 2.4765 | 1.1070 | 2.0086 | 2.5664 | 3.1944 | 3.1944 |
O2 | 1.2115 | | 2.2762 | 3.6026 | 2.0347 | 2.4901 | 3.7776 | 4.2463 | 4.2463 | N3 | 1.3524 | 2.2762 | | 1.4418 | 2.0466 | 1.0082 | 2.0691 | 2.1053 | 2.1053 | C4 | 2.4765 | 3.6026 | 1.4418 | | 2.5992 | 2.1261 | 1.0901 | 1.0925 | 1.0925 | H5 | 1.1070 | 2.0347 | 2.0466 | 2.5992 | | 2.9363 | 2.2066 | 3.3557 | 3.3557 | H6 | 2.0086 | 2.4901 | 1.0082 | 2.1261 | 2.9363 | | 2.9806 | 2.5305 | 2.5305 | H7 | 2.5664 | 3.7776 | 2.0691 | 1.0901 | 2.2066 | 2.9806 | | 1.7658 | 1.7658 | H8 | 3.1944 | 4.2463 | 2.1053 | 1.0925 | 3.3557 | 2.5305 | 1.7658 | | 1.7751 | H9 | 3.1944 | 4.2463 | 2.1053 | 1.0925 | 3.3557 | 2.5305 | 1.7658 | 1.7751 | |
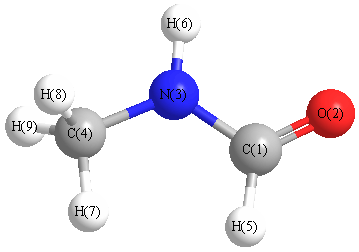 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
124.791 |
|
C1 |
N3 |
H6 |
115.851 |
| O2 |
C1 |
N3 |
125.106 |
|
O2 |
C1 |
H5 |
122.640 |
| N3 |
C1 |
H5 |
112.255 |
|
N3 |
C4 |
H7 |
108.825 |
| N3 |
C4 |
H8 |
111.605 |
|
N3 |
C4 |
H9 |
111.605 |
| C4 |
N3 |
H6 |
119.357 |
|
H7 |
C4 |
H8 |
108.005 |
| H7 |
C4 |
H9 |
108.005 |
|
H8 |
C4 |
H9 |
108.667 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/aug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.060 |
|
|
|
| 2 |
O |
-0.613 |
|
|
|
| 3 |
N |
0.007 |
|
|
|
| 4 |
C |
-0.611 |
|
|
|
| 5 |
H |
0.434 |
|
|
|
| 6 |
H |
0.063 |
|
|
|
| 7 |
H |
0.292 |
|
|
|
| 8 |
H |
0.244 |
|
|
|
| 9 |
H |
0.244 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.158 |
2.864 |
0.000 |
4.263 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.920 |
4.221 |
0.000 |
| y |
4.221 |
-24.489 |
0.000 |
| z |
0.000 |
0.000 |
-24.790 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-2.280 |
4.221 |
0.000 |
| y |
4.221 |
1.366 |
0.000 |
| z |
0.000 |
0.000 |
0.914 |
|
| Polar |
|---|
| 3z2-r2 | 1.828 |
|---|
| x2-y2 | -2.431 |
|---|
| xy | 4.221 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.778 |
-1.189 |
0.000 |
| y |
-1.189 |
6.676 |
0.000 |
| z |
0.000 |
0.000 |
4.351 |
<r2> (average value of r
2) Å
2
| <r2> |
88.740 |
| (<r2>)1/2 |
9.420 |
Jump to
S1C1
S1C3
Energy calculated at B3PW91/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -209.205502 |
| Energy at 298.15K | -209.211575 |
| HF Energy | -209.205502 |
| Nuclear repulsion energy | 119.609661 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/aug-cc-pVTZ
Geometric Data calculated at B3PW91/aug-cc-pVTZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at B3PW91/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -209.206477 |
| Energy at 298.15K | |
| HF Energy | -209.206477 |
| Nuclear repulsion energy | 121.413267 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/aug-cc-pVTZ
Geometric Data calculated at B3PW91/aug-cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.862 |
0.434 |
0.003 |
| O2 |
1.373 |
-0.669 |
0.000 |
| N3 |
-0.469 |
0.653 |
-0.019 |
| C4 |
-1.422 |
-0.438 |
0.004 |
| H5 |
1.454 |
1.364 |
0.017 |
| H6 |
-0.794 |
1.601 |
0.056 |
| H7 |
-2.415 |
-0.047 |
-0.210 |
| H8 |
-1.154 |
-1.174 |
-0.752 |
| H9 |
-1.434 |
-0.936 |
0.976 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
| C1 | | 1.2157 | 1.3494 | 2.4453 | 1.1024 | 2.0272 | 3.3191 | 2.6873 | 2.8448 |
O2 | 1.2157 | | 2.2682 | 2.8056 | 2.0349 | 3.1395 | 3.8451 | 2.6851 | 2.9838 | N3 | 1.3494 | 2.2682 | | 1.4493 | 2.0499 | 1.0044 | 2.0769 | 2.0850 | 2.1088 | C4 | 2.4453 | 2.8056 | 1.4493 | | 3.3938 | 2.1337 | 1.0882 | 1.0890 | 1.0917 | H5 | 1.1024 | 2.0349 | 2.0499 | 3.3938 | | 2.2608 | 4.1239 | 3.7194 | 3.8138 | H6 | 2.0272 | 3.1395 | 1.0044 | 2.1337 | 2.2608 | | 2.3261 | 2.9124 | 2.7731 | H7 | 3.3191 | 3.8451 | 2.0769 | 1.0882 | 4.1239 | 2.3261 | | 1.7759 | 1.7781 | H8 | 2.6873 | 2.6851 | 2.0850 | 1.0890 | 3.7194 | 2.9124 | 1.7759 | | 1.7659 | H9 | 2.8448 | 2.9838 | 2.1088 | 1.0917 | 3.8138 | 2.7731 | 1.7781 | 1.7659 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
C4 |
121.742 |
|
C1 |
N3 |
H6 |
118.171 |
| O2 |
C1 |
N3 |
124.237 |
|
O2 |
C1 |
H5 |
122.689 |
| N3 |
C1 |
H5 |
113.073 |
|
N3 |
C4 |
H7 |
109.036 |
| N3 |
C4 |
H8 |
109.630 |
|
N3 |
C4 |
H9 |
111.395 |
| C4 |
N3 |
H6 |
119.713 |
|
H7 |
C4 |
H8 |
109.308 |
| H7 |
C4 |
H9 |
109.299 |
|
H8 |
C4 |
H9 |
108.144 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/aug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.082 |
|
|
|
| 2 |
O |
-0.607 |
|
|
|
| 3 |
N |
-0.005 |
|
|
|
| 4 |
C |
-0.553 |
|
|
|
| 5 |
H |
0.416 |
|
|
|
| 6 |
H |
0.064 |
|
|
|
| 7 |
H |
0.274 |
|
|
|
| 8 |
H |
0.269 |
|
|
|
| 9 |
H |
0.225 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-3.101 |
2.695 |
-0.071 |
4.108 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.524 |
2.745 |
0.084 |
| y |
2.745 |
-22.592 |
-0.063 |
| z |
0.084 |
-0.063 |
-24.920 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-1.768 |
2.745 |
0.084 |
| y |
2.745 |
2.630 |
-0.063 |
| z |
0.084 |
-0.063 |
-0.862 |
|
| Polar |
|---|
| 3z2-r2 | -1.725 |
|---|
| x2-y2 | -2.932 |
|---|
| xy | 2.745 |
|---|
| xz | 0.084 |
|---|
| yz | -0.063 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
6.778 |
-1.189 |
0.000 |
| y |
-1.189 |
6.676 |
0.000 |
| z |
0.000 |
0.000 |
4.351 |
<r2> (average value of r
2) Å
2
| <r2> |
78.189 |
| (<r2>)1/2 |
8.842 |
