Jump to
S1C2
Vibrational Frequencies calculated at MP2=FULL/aug-cc-pVTZ
Geometric Data calculated at MP2=FULL/aug-cc-pVTZ
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at MP2=FULL/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -166.321479 |
| Energy at 298.15K | -166.322859 |
| HF Energy | -165.725626 |
| Nuclear repulsion energy | 48.966374 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at MP2=FULL/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
4032 |
3836 |
23.30 |
|
|
|
| 2 |
A |
746 |
709 |
1.16 |
|
|
|
| 3 |
A |
508 |
483 |
98.23 |
|
|
|
| 4 |
A |
324 |
308 |
21.57 |
|
|
|
| 5 |
A |
190 |
181 |
142.60 |
|
|
|
| 6 |
B |
4029 |
3833 |
227.87 |
|
|
|
| 7 |
B |
1532 |
1457 |
422.54 |
|
|
|
| 8 |
B |
496 |
472 |
325.43 |
|
|
|
| 9 |
B |
312 |
297 |
68.71 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 6083.6 cm
-1
Scaled (by 0.9514) Zero Point Vibrational Energy (zpe) 5788.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at MP2=FULL/aug-cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| Be1 |
0.000 |
0.000 |
0.015 |
| O2 |
0.000 |
1.425 |
-0.052 |
| O3 |
0.000 |
-1.425 |
-0.052 |
| H4 |
0.525 |
2.082 |
0.390 |
| H5 |
-0.525 |
-2.082 |
0.390 |
Atom - Atom Distances (Å)
| |
Be1 |
O2 |
O3 |
H4 |
H5 |
| Be1 | | 1.4267 | 1.4267 | 2.1797 | 2.1797 |
O2 | 1.4267 | | 2.8503 | 0.9501 | 3.5737 | O3 | 1.4267 | 2.8503 | | 3.5737 | 0.9501 | H4 | 2.1797 | 0.9501 | 3.5737 | | 4.2942 | H5 | 2.1797 | 3.5737 | 0.9501 | 4.2942 | |
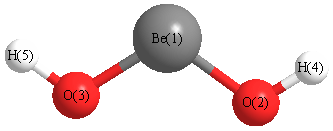 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| Be1 |
O2 |
H4 |
131.964 |
|
Be1 |
O3 |
H5 |
131.964 |
| O2 |
Be1 |
O3 |
174.602 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
