Jump to
S1C2
Energy calculated at QCISD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -1194.400686 |
| Energy at 298.15K | -1194.403059 |
| HF Energy | -1193.814038 |
| Nuclear repulsion energy | 195.269382 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
2693 |
2592 |
0.33 |
|
|
|
| 2 |
A |
898 |
864 |
0.03 |
|
|
|
| 3 |
A |
501 |
482 |
0.20 |
|
|
|
| 4 |
A |
304 |
292 |
17.18 |
|
|
|
| 5 |
A |
207 |
200 |
0.01 |
|
|
|
| 6 |
B |
2692 |
2591 |
0.46 |
|
|
|
| 7 |
B |
886 |
853 |
9.17 |
|
|
|
| 8 |
B |
506 |
487 |
17.02 |
|
|
|
| 9 |
B |
327 |
314 |
12.60 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4506.8 cm
-1
Scaled (by 0.9624) Zero Point Vibrational Energy (zpe) 4337.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/aug-cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.850 |
| S2 |
0.000 |
1.661 |
-0.390 |
| S3 |
0.000 |
-1.661 |
-0.390 |
| H4 |
-1.329 |
1.757 |
-0.562 |
| H5 |
1.329 |
-1.757 |
-0.562 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
H4 |
H5 |
| S1 | | 2.0731 | 2.0731 | 2.6168 | 2.6168 |
S2 | 2.0731 | | 3.3230 | 1.3434 | 3.6721 | S3 | 2.0731 | 3.3230 | | 3.6721 | 1.3434 | H4 | 2.6168 | 1.3434 | 3.6721 | | 4.4065 | H5 | 2.6168 | 3.6721 | 1.3434 | 4.4065 | |
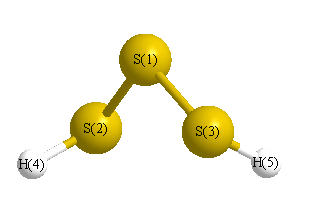 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
H4 |
97.689 |
|
S1 |
S3 |
H5 |
97.689 |
| S2 |
S1 |
S3 |
106.540 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -1194.400606 |
| Energy at 298.15K | -1194.402996 |
| HF Energy | -1193.813869 |
| Nuclear repulsion energy | 195.322843 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
2688 |
2587 |
1.72 |
|
|
|
| 2 |
A' |
900 |
866 |
3.70 |
|
|
|
| 3 |
A' |
501 |
482 |
0.26 |
|
|
|
| 4 |
A' |
328 |
315 |
13.18 |
|
|
|
| 5 |
A' |
208 |
200 |
0.04 |
|
|
|
| 6 |
A" |
2691 |
2589 |
0.15 |
|
|
|
| 7 |
A" |
888 |
855 |
6.04 |
|
|
|
| 8 |
A" |
507 |
488 |
17.94 |
|
|
|
| 9 |
A" |
314 |
302 |
8.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4512.2 cm
-1
Scaled (by 0.9624) Zero Point Vibrational Energy (zpe) 4342.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
-0.053 |
0.848 |
0.000 |
| S2 |
-0.053 |
-0.392 |
1.660 |
| S3 |
-0.053 |
-0.392 |
-1.660 |
| H4 |
1.278 |
-0.512 |
1.793 |
| H5 |
1.278 |
-0.512 |
-1.793 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
H4 |
H5 |
| S1 | | 2.0726 | 2.0726 | 2.6154 | 2.6154 |
S2 | 2.0726 | | 3.3204 | 1.3437 | 3.7030 | S3 | 2.0726 | 3.3204 | | 3.7030 | 1.3437 | H4 | 2.6154 | 1.3437 | 3.7030 | | 3.5859 | H5 | 2.6154 | 3.7030 | 1.3437 | 3.5859 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
H4 |
97.628 |
|
S1 |
S3 |
H5 |
97.628 |
| S2 |
S1 |
S3 |
106.457 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
