Jump to
S1C2
Energy calculated at B2PLYP/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -1195.462409 |
| Energy at 298.15K | |
| HF Energy | -1195.281102 |
| Nuclear repulsion energy | 194.671785 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
2672 |
2567 |
0.35 |
|
|
|
| 2 |
A |
881 |
847 |
0.04 |
|
|
|
| 3 |
A |
488 |
469 |
0.15 |
|
|
|
| 4 |
A |
314 |
301 |
17.53 |
|
|
|
| 5 |
A |
204 |
196 |
0.00 |
|
|
|
| 6 |
B |
2671 |
2566 |
0.27 |
|
|
|
| 7 |
B |
868 |
834 |
8.97 |
|
|
|
| 8 |
B |
476 |
457 |
26.05 |
|
|
|
| 9 |
B |
335 |
322 |
12.43 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4453.2 cm
-1
Scaled (by 0.961) Zero Point Vibrational Energy (zpe) 4279.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/aug-cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
0.000 |
0.000 |
0.841 |
| S2 |
0.000 |
1.675 |
-0.385 |
| S3 |
0.000 |
-1.675 |
-0.385 |
| H4 |
-1.330 |
1.781 |
-0.560 |
| H5 |
1.330 |
-1.781 |
-0.560 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
H4 |
H5 |
| S1 | | 2.0759 | 2.0759 | 2.6270 | 2.6270 |
S2 | 2.0759 | | 3.3501 | 1.3454 | 3.7070 | S3 | 2.0759 | 3.3501 | | 3.7070 | 1.3454 | H4 | 2.6270 | 1.3454 | 3.7070 | | 4.4451 | H5 | 2.6270 | 3.7070 | 1.3454 | 4.4451 | |
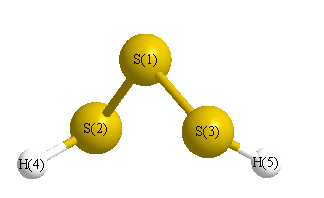 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
H4 |
98.043 |
|
S1 |
S3 |
H5 |
98.043 |
| S2 |
S1 |
S3 |
107.584 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/aug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.026 |
|
|
|
| 2 |
S |
-0.135 |
|
|
|
| 3 |
S |
-0.135 |
|
|
|
| 4 |
H |
0.148 |
|
|
|
| 5 |
H |
0.148 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-0.522 |
0.522 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.688 |
-3.482 |
0.000 |
| y |
-3.482 |
-39.692 |
0.000 |
| z |
0.000 |
0.000 |
-41.697 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.006 |
-3.482 |
0.000 |
| y |
-3.482 |
0.001 |
0.000 |
| z |
0.000 |
0.000 |
-3.007 |
|
| Polar |
|---|
| 3z2-r2 | -6.014 |
|---|
| x2-y2 | 2.004 |
|---|
| xy | -3.482 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.788 |
-0.634 |
0.000 |
| y |
-0.634 |
13.841 |
0.000 |
| z |
0.000 |
0.000 |
8.845 |
<r2> (average value of r
2) Å
2
| <r2> |
141.151 |
| (<r2>)1/2 |
11.881 |
Jump to
S1C1
Energy calculated at B2PLYP/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -1195.462327 |
| Energy at 298.15K | |
| HF Energy | -1195.280986 |
| Nuclear repulsion energy | 194.744445 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B2PLYP/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
204 |
196 |
0.03 |
|
|
|
| 2 |
A' |
336 |
323 |
13.24 |
|
|
|
| 3 |
A' |
488 |
469 |
0.22 |
|
|
|
| 4 |
A' |
883 |
848 |
2.80 |
|
|
|
| 5 |
A' |
2666 |
2562 |
1.43 |
|
|
|
| 6 |
A" |
325 |
312 |
7.01 |
|
|
|
| 7 |
A" |
477 |
458 |
27.92 |
|
|
|
| 8 |
A" |
871 |
837 |
6.59 |
|
|
|
| 9 |
A" |
2669 |
2565 |
0.14 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4459.1 cm
-1
Scaled (by 0.961) Zero Point Vibrational Energy (zpe) 4285.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B2PLYP/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| S1 |
-0.053 |
0.839 |
0.000 |
| S2 |
-0.053 |
-0.388 |
1.673 |
| S3 |
-0.053 |
-0.388 |
-1.673 |
| H4 |
1.279 |
-0.507 |
1.819 |
| H5 |
1.279 |
-0.507 |
-1.819 |
Atom - Atom Distances (Å)
| |
S1 |
S2 |
S3 |
H4 |
H5 |
| S1 | | 2.0752 | 2.0752 | 2.6258 | 2.6258 |
S2 | 2.0752 | | 3.3468 | 1.3457 | 3.7396 | S3 | 2.0752 | 3.3468 | | 3.7396 | 1.3457 | H4 | 2.6258 | 1.3457 | 3.7396 | | 3.6375 | H5 | 2.6258 | 3.7396 | 1.3457 | 3.6375 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| S1 |
S2 |
H4 |
98.002 |
|
S1 |
S3 |
H5 |
98.002 |
| S2 |
S1 |
S3 |
107.490 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/aug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
S |
-0.025 |
|
|
|
| 2 |
S |
-0.131 |
|
|
|
| 3 |
S |
-0.131 |
|
|
|
| 4 |
H |
0.144 |
|
|
|
| 5 |
H |
0.144 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.574 |
-0.452 |
0.000 |
1.638 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-37.867 |
-1.294 |
0.008 |
| y |
-1.294 |
-41.819 |
0.000 |
| z |
0.008 |
0.000 |
-39.505 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
2.795 |
-1.294 |
0.008 |
| y |
-1.294 |
-3.133 |
0.000 |
| z |
0.008 |
0.000 |
0.337 |
|
| Polar |
|---|
| 3z2-r2 | 0.675 |
|---|
| x2-y2 | 3.952 |
|---|
| xy | -1.294 |
|---|
| xz | 0.008 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.791 |
-0.233 |
-0.016 |
| y |
-0.233 |
8.831 |
0.005 |
| z |
-0.016 |
0.005 |
13.868 |
<r2> (average value of r
2) Å
2
| <r2> |
141.049 |
| (<r2>)1/2 |
11.876 |
