Jump to
S1C2
Energy calculated at CCSD(T)/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -454.199245 |
| Energy at 298.15K | -454.202985 |
| HF Energy | -453.746450 |
| Nuclear repulsion energy | 57.443149 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3510 |
3405 |
|
|
|
|
| 2 |
A' |
2672 |
2592 |
|
|
|
|
| 3 |
A' |
1628 |
1579 |
|
|
|
|
| 4 |
A' |
1040 |
1008 |
|
|
|
|
| 5 |
A' |
891 |
864 |
|
|
|
|
| 6 |
A' |
666 |
646 |
|
|
|
|
| 7 |
A" |
3603 |
3495 |
|
|
|
|
| 8 |
A" |
1136 |
1102 |
|
|
|
|
| 9 |
A" |
408 |
396 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7777.4 cm
-1
Scaled (by 0.9699) Zero Point Vibrational Energy (zpe) 7543.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD(T)/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.014 |
1.116 |
0.000 |
| S2 |
0.014 |
-0.621 |
0.000 |
| H3 |
-1.322 |
-0.773 |
0.000 |
| H4 |
0.505 |
1.446 |
0.822 |
| H5 |
0.505 |
1.446 |
-0.822 |
Atom - Atom Distances (Å)
| |
N1 |
S2 |
H3 |
H4 |
H5 |
| N1 | | 1.7371 | 2.3132 | 1.0130 | 1.0130 |
S2 | 1.7371 | | 1.3440 | 2.2777 | 2.2777 | H3 | 2.3132 | 1.3440 | | 2.9892 | 2.9892 | H4 | 1.0130 | 2.2777 | 2.9892 | | 1.6439 | H5 | 1.0130 | 2.2777 | 2.9892 | 1.6439 | |
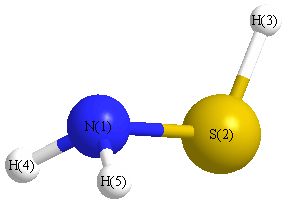 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
S2 |
H3 |
96.482 |
|
S2 |
N1 |
H4 |
108.975 |
| S2 |
N1 |
H5 |
108.975 |
|
H4 |
N1 |
H5 |
108.472 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCSD(T)/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -454.198709 |
| Energy at 298.15K | -454.202510 |
| HF Energy | -453.746712 |
| Nuclear repulsion energy | 57.628177 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCSD(T)/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3513 |
3407 |
|
|
|
|
| 2 |
A' |
2606 |
2527 |
|
|
|
|
| 3 |
A' |
1617 |
1568 |
|
|
|
|
| 4 |
A' |
1020 |
989 |
|
|
|
|
| 5 |
A' |
883 |
856 |
|
|
|
|
| 6 |
A' |
644 |
625 |
|
|
|
|
| 7 |
A" |
3615 |
3506 |
|
|
|
|
| 8 |
A" |
1119 |
1086 |
|
|
|
|
| 9 |
A" |
501 |
486 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7758.5 cm
-1
Scaled (by 0.9699) Zero Point Vibrational Energy (zpe) 7525.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCSD(T)/aug-cc-pVTZ
Point Group is Cs
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
