Jump to
S1C2
Energy calculated at B3LYPultrafine/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -454.791770 |
| Energy at 298.15K | -454.795498 |
| HF Energy | -454.791770 |
| Nuclear repulsion energy | 57.462196 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYPultrafine/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3510 |
3395 |
5.79 |
|
|
|
| 2 |
A' |
2623 |
2538 |
10.61 |
|
|
|
| 3 |
A' |
1622 |
1570 |
12.21 |
|
|
|
| 4 |
A' |
1031 |
998 |
7.10 |
|
|
|
| 5 |
A' |
870 |
842 |
40.18 |
|
|
|
| 6 |
A' |
632 |
612 |
73.20 |
|
|
|
| 7 |
A" |
3598 |
3481 |
25.10 |
|
|
|
| 8 |
A" |
1124 |
1088 |
0.80 |
|
|
|
| 9 |
A" |
423 |
409 |
42.95 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7717.2 cm
-1
Scaled (by 0.9675) Zero Point Vibrational Energy (zpe) 7466.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYPultrafine/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.015 |
1.113 |
0.000 |
| S2 |
0.015 |
-0.620 |
0.000 |
| H3 |
-1.323 |
-0.788 |
0.000 |
| H4 |
0.487 |
1.459 |
0.825 |
| H5 |
0.487 |
1.459 |
-0.825 |
Atom - Atom Distances (Å)
| |
N1 |
S2 |
H3 |
H4 |
H5 |
| N1 | | 1.7337 | 2.3255 | 1.0111 | 1.0111 |
S2 | 1.7337 | | 1.3489 | 2.2863 | 2.2863 | H3 | 2.3255 | 1.3489 | | 3.0012 | 3.0012 | H4 | 1.0111 | 2.2863 | 3.0012 | | 1.6497 | H5 | 1.0111 | 2.2863 | 3.0012 | 1.6497 | |
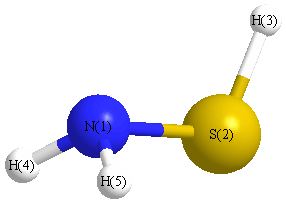 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
S2 |
H3 |
97.154 |
|
S2 |
N1 |
H4 |
110.004 |
| S2 |
N1 |
H5 |
110.004 |
|
H4 |
N1 |
H5 |
109.339 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYPultrafine/aug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.376 |
|
|
|
| 2 |
S |
0.001 |
|
|
|
| 3 |
H |
0.123 |
|
|
|
| 4 |
H |
0.126 |
|
|
|
| 5 |
H |
0.126 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.520 |
0.891 |
0.000 |
1.032 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-19.565 |
3.425 |
0.000 |
| y |
3.425 |
-19.416 |
0.000 |
| z |
0.000 |
0.000 |
-20.576 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
0.431 |
3.425 |
0.000 |
| y |
3.425 |
0.654 |
0.000 |
| z |
0.000 |
0.000 |
-1.086 |
|
| Polar |
|---|
| 3z2-r2 | -2.171 |
|---|
| x2-y2 | -0.149 |
|---|
| xy | 3.425 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.662 |
0.278 |
0.000 |
| y |
0.278 |
5.872 |
0.000 |
| z |
0.000 |
0.000 |
4.594 |
<r2> (average value of r
2) Å
2
| <r2> |
35.705 |
| (<r2>)1/2 |
5.975 |
Jump to
S1C1
Energy calculated at B3LYPultrafine/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -454.791407 |
| Energy at 298.15K | -454.795187 |
| HF Energy | -454.791407 |
| Nuclear repulsion energy | 57.706639 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3LYPultrafine/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3512 |
3398 |
2.36 |
|
|
|
| 2 |
A' |
2548 |
2465 |
35.14 |
|
|
|
| 3 |
A' |
1610 |
1558 |
12.29 |
|
|
|
| 4 |
A' |
1008 |
976 |
20.14 |
|
|
|
| 5 |
A' |
864 |
836 |
23.70 |
|
|
|
| 6 |
A' |
606 |
586 |
115.12 |
|
|
|
| 7 |
A" |
3610 |
3493 |
24.96 |
|
|
|
| 8 |
A" |
1105 |
1070 |
1.27 |
|
|
|
| 9 |
A" |
514 |
498 |
2.36 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 7689.1 cm
-1
Scaled (by 0.9675) Zero Point Vibrational Energy (zpe) 7439.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3LYPultrafine/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| N1 |
0.083 |
1.103 |
0.000 |
| S2 |
0.083 |
-0.613 |
0.000 |
| H3 |
-1.242 |
-0.911 |
0.000 |
| H4 |
-0.334 |
1.496 |
0.832 |
| H5 |
-0.334 |
1.496 |
-0.832 |
Atom - Atom Distances (Å)
| |
N1 |
S2 |
H3 |
H4 |
H5 |
| N1 | | 1.7157 | 2.4109 | 1.0101 | 1.0101 |
S2 | 1.7157 | | 1.3580 | 2.3051 | 2.3051 | H3 | 2.4109 | 1.3580 | | 2.7042 | 2.7042 | H4 | 1.0101 | 2.3051 | 2.7042 | | 1.6633 | H5 | 1.0101 | 2.3051 | 2.7042 | 1.6633 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| N1 |
S2 |
H3 |
102.704 |
|
S2 |
N1 |
H4 |
112.921 |
| S2 |
N1 |
H5 |
112.921 |
|
H4 |
N1 |
H5 |
110.846 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYPultrafine/aug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
N |
-0.382 |
|
|
|
| 2 |
S |
0.048 |
|
|
|
| 3 |
H |
0.096 |
|
|
|
| 4 |
H |
0.119 |
|
|
|
| 5 |
H |
0.119 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.872 |
1.005 |
0.000 |
2.125 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-20.391 |
-1.181 |
0.000 |
| y |
-1.181 |
-18.716 |
0.000 |
| z |
0.000 |
0.000 |
-20.507 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-0.779 |
-1.181 |
0.000 |
| y |
-1.181 |
1.733 |
0.000 |
| z |
0.000 |
0.000 |
-0.954 |
|
| Polar |
|---|
| 3z2-r2 | -1.908 |
|---|
| x2-y2 | -1.675 |
|---|
| xy | -1.181 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
4.685 |
0.247 |
0.000 |
| y |
0.247 |
5.810 |
0.000 |
| z |
0.000 |
0.000 |
4.593 |
<r2> (average value of r
2) Å
2
| <r2> |
35.549 |
| (<r2>)1/2 |
5.962 |
