Jump to
S1C2
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -117.009121 |
| Energy at 298.15K | -117.013451 |
| HF Energy | -116.512425 |
| Nuclear repulsion energy | 65.243597 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3285 |
3144 |
9.89 |
|
|
|
| 2 |
A1 |
3190 |
3053 |
0.45 |
|
|
|
| 3 |
A1 |
3181 |
3045 |
10.73 |
|
|
|
| 4 |
A1 |
1550 |
1484 |
2.74 |
|
|
|
| 5 |
A1 |
1303 |
1247 |
1.31 |
|
|
|
| 6 |
A1 |
1060 |
1015 |
0.00 |
|
|
|
| 7 |
A1 |
430 |
411 |
0.25 |
|
|
|
| 8 |
A2 |
826 |
790 |
0.00 |
|
|
|
| 9 |
A2 |
574 |
549 |
0.00 |
|
|
|
| 10 |
B1 |
1043 |
998 |
23.81 |
|
|
|
| 11 |
B1 |
848 |
812 |
75.04 |
|
|
|
| 12 |
B1 |
548 |
525 |
16.73 |
|
|
|
| 13 |
B2 |
3282 |
3141 |
3.14 |
|
|
|
| 14 |
B2 |
3178 |
3042 |
7.19 |
|
|
|
| 15 |
B2 |
1518 |
1453 |
0.60 |
|
|
|
| 16 |
B2 |
1431 |
1370 |
6.16 |
|
|
|
| 17 |
B2 |
1146 |
1097 |
0.20 |
|
|
|
| 18 |
B2 |
942 |
902 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14668.2 cm
-1
Scaled (by 0.957) Zero Point Vibrational Energy (zpe) 14037.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.445 |
| H2 |
0.000 |
0.000 |
1.529 |
| C3 |
0.000 |
1.219 |
-0.197 |
| C4 |
0.000 |
-1.219 |
-0.197 |
| H5 |
0.000 |
2.146 |
0.356 |
| H6 |
0.000 |
-2.146 |
0.356 |
| H7 |
0.000 |
1.276 |
-1.276 |
| H8 |
0.000 |
-1.276 |
-1.276 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.0839 | 1.3775 | 1.3775 | 2.1473 | 2.1473 | 2.1430 | 2.1430 |
H2 | 1.0839 | | 2.1127 | 2.1127 | 2.4451 | 2.4451 | 3.0821 | 3.0821 | C3 | 1.3775 | 2.1127 | | 2.4377 | 1.0791 | 3.4095 | 1.0813 | 2.7187 | C4 | 1.3775 | 2.1127 | 2.4377 | | 3.4095 | 1.0791 | 2.7187 | 1.0813 | H5 | 2.1473 | 2.4451 | 1.0791 | 3.4095 | | 4.2910 | 1.8497 | 3.7913 | H6 | 2.1473 | 2.4451 | 3.4095 | 1.0791 | 4.2910 | | 3.7913 | 1.8497 | H7 | 2.1430 | 3.0821 | 1.0813 | 2.7187 | 1.8497 | 3.7913 | | 2.5524 | H8 | 2.1430 | 3.0821 | 2.7187 | 1.0813 | 3.7913 | 1.8497 | 2.5524 | |
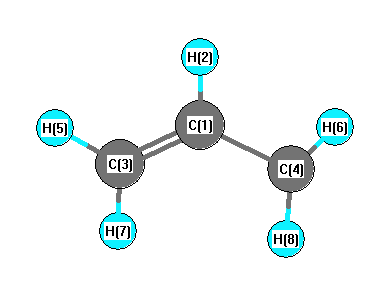 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
H5 |
121.408 |
|
C1 |
C3 |
H7 |
120.806 |
| C1 |
C4 |
H6 |
121.408 |
|
C1 |
C4 |
H8 |
120.806 |
| H2 |
C1 |
C3 |
117.767 |
|
H2 |
C1 |
C4 |
117.767 |
| C3 |
C1 |
C4 |
124.465 |
|
H5 |
C3 |
H7 |
117.786 |
| H6 |
C4 |
H8 |
117.786 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -117.009121 |
| Energy at 298.15K | -117.013452 |
| HF Energy | -116.512421 |
| Nuclear repulsion energy | 65.246602 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3285 |
3144 |
9.92 |
|
|
|
| 2 |
A' |
3282 |
3141 |
3.14 |
|
|
|
| 3 |
A' |
3190 |
3053 |
0.58 |
|
|
|
| 4 |
A' |
3181 |
3045 |
10.56 |
|
|
|
| 5 |
A' |
3178 |
3041 |
7.21 |
|
|
|
| 6 |
A' |
1550 |
1484 |
2.75 |
|
|
|
| 7 |
A' |
1518 |
1453 |
0.59 |
|
|
|
| 8 |
A' |
1431 |
1370 |
6.22 |
|
|
|
| 9 |
A' |
1304 |
1247 |
1.32 |
|
|
|
| 10 |
A' |
1146 |
1097 |
0.19 |
|
|
|
| 11 |
A' |
1061 |
1015 |
0.00 |
|
|
|
| 12 |
A' |
943 |
902 |
0.00 |
|
|
|
| 13 |
A' |
430 |
411 |
0.25 |
|
|
|
| 14 |
A" |
1043 |
998 |
23.80 |
|
|
|
| 15 |
A" |
848 |
812 |
75.02 |
|
|
|
| 16 |
A" |
826 |
790 |
0.04 |
|
|
|
| 17 |
A" |
574 |
549 |
0.00 |
|
|
|
| 18 |
A" |
549 |
525 |
16.72 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14668.6 cm
-1
Scaled (by 0.957) Zero Point Vibrational Energy (zpe) 14037.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.445 |
0.000 |
| H2 |
0.000 |
1.529 |
0.000 |
| C3 |
-1.219 |
-0.197 |
0.000 |
| C4 |
1.219 |
-0.196 |
0.000 |
| H5 |
-2.146 |
0.355 |
0.000 |
| H6 |
2.145 |
0.357 |
0.000 |
| H7 |
-1.275 |
-1.277 |
0.000 |
| H8 |
1.276 |
-1.276 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
H2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.0839 | 1.3777 | 1.3772 | 2.1476 | 2.1471 | 2.1427 | 2.1428 |
H2 | 1.0839 | | 2.1132 | 2.1124 | 2.4458 | 2.4448 | 3.0821 | 3.0819 | C3 | 1.3777 | 2.1132 | | 2.4374 | 1.0792 | 3.4093 | 1.0813 | 2.7182 | C4 | 1.3772 | 2.1124 | 2.4374 | | 3.4093 | 1.0791 | 2.7176 | 1.0813 | H5 | 2.1476 | 2.4458 | 1.0792 | 3.4093 | | 4.2910 | 1.8502 | 3.7909 | H6 | 2.1471 | 2.4448 | 3.4093 | 1.0791 | 4.2910 | | 3.7902 | 1.8497 | H7 | 2.1427 | 3.0821 | 1.0813 | 2.7176 | 1.8502 | 3.7902 | | 2.5510 | H8 | 2.1428 | 3.0819 | 2.7182 | 1.0813 | 3.7909 | 1.8497 | 2.5510 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C3 |
H5 |
121.410 |
|
C1 |
C3 |
H7 |
120.766 |
| C1 |
C4 |
H6 |
121.410 |
|
C1 |
C4 |
H8 |
120.813 |
| H2 |
C1 |
C3 |
117.792 |
|
H2 |
C1 |
C4 |
117.764 |
| C3 |
C1 |
C4 |
124.444 |
|
H5 |
C3 |
H7 |
117.823 |
| H6 |
C4 |
H8 |
117.777 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
