return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for C4H4N (pyrrolide radical)

using model chemistry: CCD/aug-cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes C2V 2A2
Energy calculated at CCD/aug-cc-pVTZ
 hartrees
Energy at 0K-209.091872
Energy at 298.15K-209.096664
HF Energy-208.270260
Nuclear repulsion energy151.678681
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A1 3289 3148 0.59      
2 A1 3249 3110 0.16      
3 A1 1605 1536 9.54      
4 A1 1486 1422 28.35      
5 A1 1253 1199 5.93      
6 A1 1115 1067 38.17      
7 A1 1070 1024 0.06      
8 A1 899 860 10.93      
9 A2 929 889 0.00      
10 A2 888 850 0.00      
11 A2 520 498 0.00      
12 B1 898 859 0.01      
13 B1 736 704 68.68      
14 B1 572 547 21.51      
15 B2 4831 4623 78114.10      
16 B2 3260 3120 75.40      
17 B2 3238 3099 177.22      
18 B2 1336 1278 132.23      
19 B2 1303 1247 53.98      
20 B2 1037 993 4.27      
21 B2 897 859 1.90      

Unscaled Zero Point Vibrational Energy (zpe) 17205.0 cm-1
Scaled (by 0.957) Zero Point Vibrational Energy (zpe) 16465.2 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/aug-cc-pVTZ
ABC
0.33613 0.29724 0.15775

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/aug-cc-pVTZ

Point Group is C2v

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
N1 0.000 0.000 1.238
C2 0.000 1.057 0.424
C3 0.000 -1.057 0.424
C4 0.000 0.680 -0.979
C5 0.000 -0.680 -0.979
H6 0.000 2.062 0.818
H7 0.000 -2.062 0.818
H8 0.000 1.352 -1.820
H9 0.000 -1.352 -1.820

Atom - Atom Distances (Å)
  N1 C2 C3 C4 C5 H6 H7 H8 H9
N11.33361.33362.31832.31832.10402.10403.34333.3433
C21.33362.11311.45212.23241.08003.14332.26343.2921
C31.33362.11312.23241.45213.14331.08003.29212.2634
C42.31831.45212.23241.36052.26683.27861.07682.1998
C52.31832.23241.45211.36053.27862.26682.19981.0768
H62.10401.08003.14332.26683.27864.12372.73244.3149
H72.10403.14331.08003.27862.26684.12374.31492.7324
H83.34332.26343.29211.07682.19982.73244.31492.7044
H93.34333.29212.26342.19981.07684.31492.73242.7044

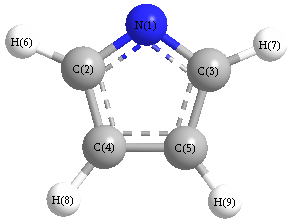
picture of pyrrolide radical state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
N1 C2 C4 112.583 N1 C2 H6 120.967
N1 C3 C5 112.583 N1 C3 H7 120.967
C2 N1 C3 104.798 C2 C4 C5 105.018
C2 C4 H8 126.373 C3 C5 C4 105.018
C3 C5 H9 126.373 C4 C2 H6 126.450
C4 C5 H9 128.609 C5 C3 H7 126.450
C5 C4 H8 128.609
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability