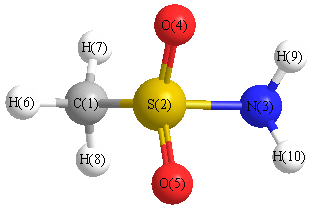
Vibrational Frequencies calculated at B3LYPultrafine/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3509 |
3395 |
35.64 |
|
|
|
| 2 |
A' |
3160 |
3058 |
0.20 |
|
|
|
| 3 |
A' |
3065 |
2965 |
0.23 |
|
|
|
| 4 |
A' |
1583 |
1532 |
34.56 |
|
|
|
| 5 |
A' |
1456 |
1409 |
5.44 |
|
|
|
| 6 |
A' |
1350 |
1306 |
10.93 |
|
|
|
| 7 |
A' |
1136 |
1099 |
167.41 |
|
|
|
| 8 |
A' |
986 |
954 |
23.18 |
|
|
|
| 9 |
A' |
864 |
836 |
105.82 |
|
|
|
| 10 |
A' |
704 |
681 |
11.27 |
|
|
|
| 11 |
A' |
651 |
630 |
212.40 |
|
|
|
| 12 |
A' |
490 |
474 |
40.45 |
|
|
|
| 13 |
A' |
468 |
453 |
9.00 |
|
|
|
| 14 |
A' |
290 |
281 |
4.61 |
|
|
|
| 15 |
A" |
3612 |
3495 |
51.19 |
|
|
|
| 16 |
A" |
3169 |
3066 |
0.36 |
|
|
|
| 17 |
A" |
1457 |
1410 |
1.47 |
|
|
|
| 18 |
A" |
1342 |
1299 |
259.49 |
|
|
|
| 19 |
A" |
1089 |
1054 |
4.24 |
|
|
|
| 20 |
A" |
972 |
940 |
0.82 |
|
|
|
| 21 |
A" |
392 |
380 |
0.03 |
|
|
|
| 22 |
A" |
327 |
317 |
2.81 |
|
|
|
| 23 |
A" |
218 |
211 |
5.07 |
|
|
|
| 24 |
A" |
185 |
179 |
28.31 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16238.1 cm
-1
Scaled (by 0.9675) Zero Point Vibrational Energy (zpe) 15710.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3LYPultrafine/aug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.700 |
|
|
|
| 2 |
S |
2.083 |
|
|
|
| 3 |
N |
-0.442 |
|
|
|
| 4 |
O |
-0.886 |
|
|
|
| 5 |
O |
-0.886 |
|
|
|
| 6 |
H |
0.216 |
|
|
|
| 7 |
H |
0.221 |
|
|
|
| 8 |
H |
0.221 |
|
|
|
| 9 |
H |
0.087 |
|
|
|
| 10 |
H |
0.087 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.810 |
3.023 |
0.000 |
3.523 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-32.701 |
5.371 |
0.000 |
| y |
5.371 |
-36.970 |
0.000 |
| z |
0.000 |
0.000 |
-41.439 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.504 |
5.371 |
0.000 |
| y |
5.371 |
0.100 |
0.000 |
| z |
0.000 |
0.000 |
-6.604 |
|
| Polar |
|---|
| 3z2-r2 | -13.208 |
|---|
| x2-y2 | 4.269 |
|---|
| xy | 5.371 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.783 |
0.258 |
0.000 |
| y |
0.258 |
7.408 |
0.000 |
| z |
0.000 |
0.000 |
7.262 |
<r2> (average value of r
2) Å
2
| <r2> |
121.367 |
| (<r2>)1/2 |
11.017 |