All results from a given calculation for CH3CHFCH3 (2-Fluoropropane)
using model chemistry: CCD/aug-cc-pVTZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
CS |
1A' |
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -218.034354 |
| Energy at 298.15K | |
| HF Energy | -217.206932 |
| Nuclear repulsion energy | 132.968170 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.279 |
0.246 |
0.000 |
| F2 |
-0.869 |
1.038 |
0.000 |
| H3 |
1.124 |
0.938 |
0.000 |
| C4 |
0.279 |
-0.584 |
1.266 |
| C5 |
0.279 |
-0.584 |
-1.266 |
| H6 |
1.189 |
-1.182 |
1.323 |
| H7 |
1.189 |
-1.182 |
-1.323 |
| H8 |
0.231 |
0.061 |
2.143 |
| H9 |
0.231 |
0.061 |
-2.143 |
| H10 |
-0.581 |
-1.254 |
1.276 |
| H11 |
-0.581 |
-1.254 |
-1.276 |
Atom - Atom Distances (Å)
| |
C1 |
F2 |
H3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
| C1 | | 1.3953 | 1.0922 | 1.5137 | 1.5137 | 2.1486 | 2.1486 | 2.1517 | 2.1517 | 2.1483 | 2.1483 |
F2 | 1.3953 | | 1.9959 | 2.3563 | 2.3563 | 3.3040 | 3.3040 | 2.5997 | 2.5997 | 2.6385 | 2.6385 | H3 | 1.0922 | 1.9959 | | 2.1521 | 2.1521 | 2.4997 | 2.4997 | 2.4820 | 2.4820 | 3.0557 | 3.0557 | C4 | 1.5137 | 2.3563 | 2.1521 | | 2.5328 | 1.0907 | 2.8090 | 1.0892 | 3.4703 | 1.0906 | 2.7662 | C5 | 1.5137 | 2.3563 | 2.1521 | 2.5328 | | 2.8090 | 1.0907 | 3.4703 | 1.0892 | 2.7662 | 1.0906 | H6 | 2.1486 | 3.3040 | 2.4997 | 1.0907 | 2.8090 | | 2.6457 | 1.7710 | 3.8049 | 1.7723 | 3.1451 | H7 | 2.1486 | 3.3040 | 2.4997 | 2.8090 | 1.0907 | 2.6457 | | 3.8049 | 1.7710 | 3.1451 | 1.7723 | H8 | 2.1517 | 2.5997 | 2.4820 | 1.0892 | 3.4703 | 1.7710 | 3.8049 | | 4.2865 | 1.7719 | 3.7519 | H9 | 2.1517 | 2.5997 | 2.4820 | 3.4703 | 1.0892 | 3.8049 | 1.7710 | 4.2865 | | 3.7519 | 1.7719 | H10 | 2.1483 | 2.6385 | 3.0557 | 1.0906 | 2.7662 | 1.7723 | 3.1451 | 1.7719 | 3.7519 | | 2.5515 | H11 | 2.1483 | 2.6385 | 3.0557 | 2.7662 | 1.0906 | 3.1451 | 1.7723 | 3.7519 | 1.7719 | 2.5515 | |
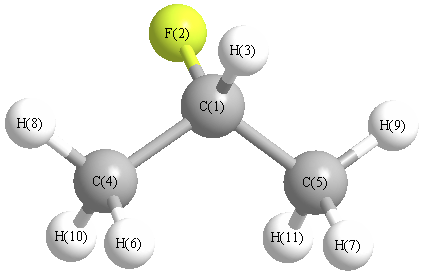 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C4 |
H8 |
110.457 |
|
C1 |
C4 |
H10 |
110.103 |
| C1 |
C5 |
H7 |
110.121 |
|
C1 |
C5 |
H9 |
110.457 |
| C1 |
C5 |
H11 |
110.103 |
|
F2 |
C1 |
H3 |
106.074 |
| F2 |
C1 |
C4 |
108.124 |
|
F2 |
C1 |
C5 |
108.124 |
| H3 |
C1 |
C4 |
110.308 |
|
H3 |
C1 |
C5 |
110.308 |
| C4 |
C1 |
C5 |
113.577 |
|
H7 |
C5 |
H9 |
108.669 |
| H7 |
C5 |
H11 |
108.684 |
|
H8 |
C4 |
H10 |
108.762 |
| H9 |
C5 |
H11 |
108.762 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
