All results from a given calculation for C4H2 (Diacetylene)
using model chemistry: QCISD(T)/aug-cc-pVTZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
D*H |
1Σg |
Energy calculated at QCISD(T)/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -153.205961 |
| Energy at 298.15K | |
| HF Energy | -152.550335 |
| Nuclear repulsion energy | 76.711775 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVTZ
Geometric Data calculated at QCISD(T)/aug-cc-pVTZ
Point Group is D∞h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.690 |
| C2 |
0.000 |
0.000 |
-0.690 |
| C3 |
0.000 |
0.000 |
1.905 |
| C4 |
0.000 |
0.000 |
-1.905 |
| H5 |
0.000 |
0.000 |
2.970 |
| H6 |
0.000 |
0.000 |
-2.970 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
H5 |
H6 |
| C1 | | 1.3790 | 1.2159 | 2.5949 | 2.2803 | 3.6593 |
C2 | 1.3790 | | 2.5949 | 1.2159 | 3.6593 | 2.2803 | C3 | 1.2159 | 2.5949 | | 3.8108 | 1.0644 | 4.8752 | C4 | 2.5949 | 1.2159 | 3.8108 | | 4.8752 | 1.0644 | H5 | 2.2803 | 3.6593 | 1.0644 | 4.8752 | | 5.9396 | H6 | 3.6593 | 2.2803 | 4.8752 | 1.0644 | 5.9396 | |
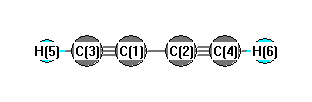 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
180.000 |
|
C1 |
C3 |
H5 |
180.000 |
| C2 |
C1 |
C3 |
180.000 |
|
C2 |
C4 |
H6 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
