Jump to
S1C2
Energy calculated at QCISD(T)/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -192.803262 |
| Energy at 298.15K | |
| HF Energy | -191.981643 |
| Nuclear repulsion energy | 127.840339 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3086 |
2983 |
|
|
|
|
| 2 |
A1 |
3040 |
2939 |
|
|
|
|
| 3 |
A1 |
1549 |
1497 |
|
|
|
|
| 4 |
A1 |
1499 |
1449 |
|
|
|
|
| 5 |
A1 |
1373 |
1327 |
|
|
|
|
| 6 |
A1 |
1045 |
1010 |
|
|
|
|
| 7 |
A1 |
925 |
894 |
|
|
|
|
| 8 |
A1 |
806 |
780 |
|
|
|
|
| 9 |
A2 |
3083 |
2981 |
|
|
|
|
| 10 |
A2 |
1236 |
1195 |
|
|
|
|
| 11 |
A2 |
1164 |
1125 |
|
|
|
|
| 12 |
A2 |
830 |
803 |
|
|
|
|
| 13 |
B1 |
3145 |
3040 |
|
|
|
|
| 14 |
B1 |
3081 |
2979 |
|
|
|
|
| 15 |
B1 |
1198 |
1159 |
|
|
|
|
| 16 |
B1 |
1145 |
1107 |
|
|
|
|
| 17 |
B1 |
758 |
733 |
|
|
|
|
| 18 |
B1 |
78i |
75i |
|
|
|
|
| 19 |
B2 |
3033 |
2933 |
|
|
|
|
| 20 |
B2 |
1520 |
1470 |
|
|
|
|
| 21 |
B2 |
1313 |
1269 |
|
|
|
|
| 22 |
B2 |
1256 |
1214 |
|
|
|
|
| 23 |
B2 |
1034 |
1000 |
|
|
|
|
| 24 |
B2 |
946 |
914 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 18993.0 cm
-1
Scaled (by 0.9668) Zero Point Vibrational Energy (zpe) 18362.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/aug-cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.074 |
| C2 |
0.000 |
0.000 |
-1.077 |
| C3 |
0.000 |
1.041 |
0.062 |
| C4 |
0.000 |
-1.041 |
0.062 |
| H5 |
0.891 |
0.000 |
-1.704 |
| H6 |
-0.891 |
0.000 |
-1.704 |
| H7 |
0.894 |
1.666 |
0.133 |
| H8 |
-0.894 |
1.666 |
0.133 |
| H9 |
-0.894 |
-1.666 |
0.133 |
| H10 |
0.894 |
-1.666 |
0.133 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1514 | 1.4518 | 1.4518 | 2.9171 | 2.9171 | 2.1122 | 2.1122 | 2.1122 | 2.1122 |
C2 | 2.1514 | | 1.5430 | 1.5430 | 1.0889 | 1.0889 | 2.2450 | 2.2450 | 2.2450 | 2.2450 | C3 | 1.4518 | 1.5430 | | 2.0815 | 2.2347 | 2.2347 | 1.0932 | 1.0932 | 2.8517 | 2.8517 | C4 | 1.4518 | 1.5430 | 2.0815 | | 2.2347 | 2.2347 | 2.8517 | 2.8517 | 1.0932 | 1.0932 | H5 | 2.9171 | 1.0889 | 2.2347 | 2.2347 | | 1.7814 | 2.4799 | 3.0551 | 3.0551 | 2.4799 | H6 | 2.9171 | 1.0889 | 2.2347 | 2.2347 | 1.7814 | | 3.0551 | 2.4799 | 2.4799 | 3.0551 | H7 | 2.1122 | 2.2450 | 1.0932 | 2.8517 | 2.4799 | 3.0551 | | 1.7871 | 3.7818 | 3.3329 | H8 | 2.1122 | 2.2450 | 1.0932 | 2.8517 | 3.0551 | 2.4799 | 1.7871 | | 3.3329 | 3.7818 | H9 | 2.1122 | 2.2450 | 2.8517 | 1.0932 | 3.0551 | 2.4799 | 3.7818 | 3.3329 | | 1.7871 | H10 | 2.1122 | 2.2450 | 2.8517 | 1.0932 | 2.4799 | 3.0551 | 3.3329 | 3.7818 | 1.7871 | |
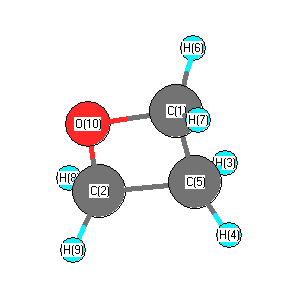 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
91.789 |
|
O1 |
C3 |
H7 |
111.413 |
| O1 |
C3 |
H8 |
111.413 |
|
O1 |
C4 |
C2 |
91.789 |
| O1 |
C4 |
H9 |
111.413 |
|
O1 |
C4 |
H10 |
111.413 |
| C2 |
C3 |
H7 |
115.721 |
|
C2 |
C3 |
H8 |
115.721 |
| C2 |
C4 |
H9 |
115.721 |
|
C2 |
C4 |
H10 |
115.721 |
| C3 |
O1 |
C4 |
91.594 |
|
C3 |
C2 |
C4 |
84.828 |
| C3 |
C2 |
H5 |
115.133 |
|
C3 |
C2 |
H6 |
115.133 |
| C4 |
C2 |
H5 |
115.133 |
|
C4 |
C2 |
H6 |
115.133 |
| H5 |
C2 |
H6 |
109.761 |
|
H7 |
C3 |
H8 |
109.652 |
| H9 |
C4 |
H10 |
109.652 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -192.803333 |
| Energy at 298.15K | |
| HF Energy | -191.981373 |
| Nuclear repulsion energy | 127.968491 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVTZ
Geometric Data calculated at QCISD(T)/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.182 |
-1.058 |
0.000 |
| C2 |
0.049 |
1.075 |
0.000 |
| C3 |
0.049 |
-0.067 |
1.036 |
| C4 |
0.049 |
-0.067 |
-1.036 |
| H5 |
0.913 |
1.738 |
0.000 |
| H6 |
-0.869 |
1.661 |
0.000 |
| H7 |
1.013 |
-0.229 |
1.529 |
| H8 |
-0.750 |
-0.065 |
1.779 |
| H9 |
1.013 |
-0.229 |
-1.529 |
| H10 |
-0.750 |
-0.065 |
-1.779 |
Atom - Atom Distances (Å)
| |
O1 |
C2 |
C3 |
C4 |
H5 |
H6 |
H7 |
H8 |
H9 |
H10 |
| O1 | | 2.1452 | 1.4523 | 1.4523 | 3.0022 | 2.8039 | 2.1104 | 2.1151 | 2.1104 | 2.1151 |
C2 | 2.1452 | | 1.5419 | 1.5419 | 1.0888 | 1.0893 | 2.2292 | 2.2599 | 2.2292 | 2.2599 | C3 | 1.4523 | 1.5419 | | 2.0725 | 2.2529 | 2.2140 | 1.0947 | 1.0918 | 2.7454 | 2.9271 | C4 | 1.4523 | 1.5419 | 2.0725 | | 2.2529 | 2.2140 | 2.7454 | 2.9271 | 1.0947 | 1.0918 | H5 | 3.0022 | 1.0888 | 2.2529 | 2.2529 | | 1.7840 | 2.4934 | 3.0308 | 2.4934 | 3.0308 | H6 | 2.8039 | 1.0893 | 2.2140 | 2.2140 | 1.7840 | | 3.0748 | 2.4822 | 3.0748 | 2.4822 | H7 | 2.1104 | 2.2292 | 1.0947 | 2.7454 | 2.4934 | 3.0748 | | 1.7887 | 3.0586 | 3.7530 | H8 | 2.1151 | 2.2599 | 1.0918 | 2.9271 | 3.0308 | 2.4822 | 1.7887 | | 3.7530 | 3.5590 | H9 | 2.1104 | 2.2292 | 2.7454 | 1.0947 | 2.4934 | 3.0748 | 3.0586 | 3.7530 | | 1.7887 | H10 | 2.1151 | 2.2599 | 2.9271 | 1.0918 | 3.0308 | 2.4822 | 3.7530 | 3.5590 | 1.7887 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
C3 |
C2 |
91.473 |
|
O1 |
C3 |
H7 |
111.129 |
| O1 |
C3 |
H8 |
111.693 |
|
O1 |
C4 |
C2 |
91.473 |
| O1 |
C4 |
H9 |
111.129 |
|
O1 |
C4 |
H10 |
111.693 |
| C2 |
C3 |
H7 |
114.374 |
|
C2 |
C3 |
H8 |
117.174 |
| C2 |
C4 |
H9 |
114.374 |
|
C2 |
C4 |
H10 |
117.174 |
| C3 |
O1 |
C4 |
91.040 |
|
C3 |
C2 |
C4 |
84.450 |
| C3 |
C2 |
H5 |
116.778 |
|
C3 |
C2 |
H6 |
113.471 |
| C4 |
C2 |
H5 |
116.778 |
|
C4 |
C2 |
H6 |
113.471 |
| H5 |
C2 |
H6 |
109.987 |
|
H7 |
C3 |
H8 |
109.780 |
| H9 |
C4 |
H10 |
109.780 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
