Jump to
S1C2
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -133.040639 |
| Energy at 298.15K | -133.044871 |
| HF Energy | -132.518410 |
| Nuclear repulsion energy | 65.264020 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3517 |
3365 |
1.63 |
|
|
|
| 2 |
A' |
3304 |
3162 |
1.66 |
|
|
|
| 3 |
A' |
3193 |
3056 |
4.14 |
|
|
|
| 4 |
A' |
3113 |
2979 |
28.34 |
|
|
|
| 5 |
A' |
1528 |
1463 |
13.92 |
|
|
|
| 6 |
A' |
1473 |
1410 |
12.65 |
|
|
|
| 7 |
A' |
1387 |
1328 |
13.20 |
|
|
|
| 8 |
A' |
1236 |
1183 |
25.69 |
|
|
|
| 9 |
A' |
1073 |
1027 |
4.35 |
|
|
|
| 10 |
A' |
1015 |
972 |
10.05 |
|
|
|
| 11 |
A' |
495 |
473 |
14.07 |
|
|
|
| 12 |
A" |
1110 |
1063 |
0.22 |
|
|
|
| 13 |
A" |
848 |
812 |
8.63 |
|
|
|
| 14 |
A" |
722 |
691 |
104.37 |
|
|
|
| 15 |
A" |
535 |
512 |
3.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 12275.1 cm
-1
Scaled (by 0.957) Zero Point Vibrational Energy (zpe) 11747.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.415 |
0.000 |
| C2 |
1.144 |
-0.387 |
0.000 |
| N3 |
-1.183 |
-0.122 |
0.000 |
| H4 |
0.142 |
1.496 |
0.000 |
| H5 |
2.132 |
0.047 |
0.000 |
| H6 |
1.037 |
-1.461 |
0.000 |
| H7 |
-1.897 |
0.603 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.3976 | 1.2990 | 1.0901 | 2.1636 | 2.1439 | 1.9059 |
C2 | 1.3976 | | 2.3426 | 2.1331 | 1.0790 | 1.0797 | 3.1982 | N3 | 1.2990 | 2.3426 | | 2.0909 | 3.3195 | 2.5930 | 1.0169 | H4 | 1.0901 | 2.1331 | 2.0909 | | 2.4614 | 3.0896 | 2.2257 | H5 | 2.1636 | 1.0790 | 3.3195 | 2.4614 | | 1.8642 | 4.0669 | H6 | 2.1439 | 1.0797 | 2.5930 | 3.0896 | 1.8642 | | 3.5872 | H7 | 1.9059 | 3.1982 | 1.0169 | 2.2257 | 4.0669 | 3.5872 | |
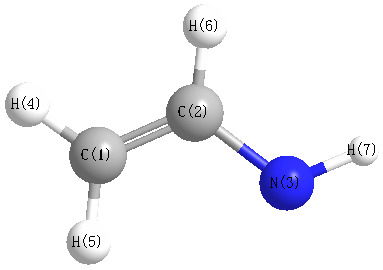 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.238 |
|
C1 |
C2 |
H6 |
119.313 |
| C1 |
N3 |
H7 |
110.171 |
|
C2 |
C1 |
N3 |
120.579 |
| C2 |
C1 |
H4 |
117.534 |
|
N3 |
C1 |
H4 |
121.887 |
| H5 |
C2 |
H6 |
119.448 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -133.039656 |
| Energy at 298.15K | |
| HF Energy | -132.517520 |
| Nuclear repulsion energy | 65.287077 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.450 |
0.000 |
| C2 |
1.118 |
-0.388 |
0.000 |
| N3 |
-1.244 |
0.071 |
0.000 |
| H4 |
0.174 |
1.523 |
0.000 |
| H5 |
2.119 |
0.016 |
0.000 |
| H6 |
0.997 |
-1.463 |
0.000 |
| H7 |
-1.289 |
-0.949 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
N3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.3975 | 1.3007 | 1.0865 | 2.1634 | 2.1571 | 1.9023 |
C2 | 1.3975 | | 2.4065 | 2.1316 | 1.0794 | 1.0813 | 2.4716 | N3 | 1.3007 | 2.4065 | | 2.0294 | 3.3639 | 2.7156 | 1.0206 | H4 | 1.0865 | 2.1316 | 2.0294 | | 2.4611 | 3.0969 | 2.8720 | H5 | 2.1634 | 1.0794 | 3.3639 | 2.4611 | | 1.8559 | 3.5420 | H6 | 2.1571 | 1.0813 | 2.7156 | 3.0969 | 1.8559 | | 2.3430 | H7 | 1.9023 | 2.4716 | 1.0206 | 2.8720 | 3.5420 | 2.3430 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
121.178 |
|
C1 |
C2 |
H6 |
120.428 |
| C1 |
N3 |
H7 |
109.474 |
|
C2 |
C1 |
N3 |
126.180 |
| C2 |
C1 |
H4 |
117.663 |
|
N3 |
C1 |
H4 |
116.157 |
| H5 |
C2 |
H6 |
118.394 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
