Jump to
S1C2
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -168.293909 |
| Energy at 298.15K | -168.294633 |
| HF Energy | -167.698501 |
| Nuclear repulsion energy | 60.687321 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3517 |
3366 |
278.75 |
|
|
|
| 2 |
Σ |
2338 |
2238 |
483.97 |
|
|
|
| 3 |
Σ |
1300 |
1244 |
188.98 |
|
|
|
| 4 |
Π |
576 |
551 |
2.38 |
|
|
|
| 4 |
Π |
576 |
551 |
2.38 |
|
|
|
| 5 |
Π |
323 |
309 |
64.60 |
|
|
|
| 5 |
Π |
323 |
309 |
64.60 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4475.8 cm
-1
Scaled (by 0.957) Zero Point Vibrational Energy (zpe) 4283.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/aug-cc-pVTZ
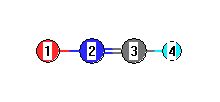
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
0.000 |
0.000 |
1.178 |
| N2 |
0.000 |
0.000 |
-0.021 |
| C3 |
0.000 |
0.000 |
-1.174 |
| H4 |
0.000 |
0.000 |
-2.234 |
Atom - Atom Distances (Å)
| |
O1 |
N2 |
C3 |
H4 |
| O1 | | 1.1999 | 2.3526 | 3.4120 |
N2 | 1.1999 | | 1.1527 | 2.2121 | C3 | 2.3526 | 1.1527 | | 1.0594 | H4 | 3.4120 | 2.2121 | 1.0594 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
N2 |
C3 |
180.000 |
|
N2 |
C3 |
H4 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -168.293909 |
| Energy at 298.15K | -168.294576 |
| HF Energy | -167.698499 |
| Nuclear repulsion energy | 60.686208 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3517 |
3366 |
278.74 |
|
|
|
| 2 |
A' |
2338 |
2238 |
483.87 |
|
|
|
| 3 |
A' |
1300 |
1244 |
189.02 |
|
|
|
| 4 |
A' |
576 |
551 |
2.38 |
|
|
|
| 5 |
A' |
323 |
309 |
64.61 |
|
|
|
| 6 |
A" |
555 |
532 |
1.17 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4304.3 cm
-1
Scaled (by 0.957) Zero Point Vibrational Energy (zpe) 4119.2 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| O1 |
-0.000 |
-1.179 |
0.000 |
| N2 |
0.000 |
0.021 |
0.000 |
| C3 |
0.000 |
1.174 |
0.000 |
| H4 |
0.001 |
2.234 |
0.000 |
Atom - Atom Distances (Å)
| |
O1 |
N2 |
C3 |
H4 |
| O1 | | 1.1999 | 2.3526 | 3.4121 |
N2 | 1.1999 | | 1.1527 | 2.2121 | C3 | 2.3526 | 1.1527 | | 1.0594 | H4 | 3.4121 | 2.2121 | 1.0594 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| O1 |
N2 |
C3 |
179.998 |
|
N2 |
C3 |
H4 |
179.974 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
