Jump to
S1C2
Energy calculated at QCISD(T)/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -151.688814 |
| Energy at 298.15K | |
| HF Energy | -151.140831 |
| Nuclear repulsion energy | 52.269744 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
Σ |
3451 |
3337 |
|
|
|
|
| 2 |
Σ |
2080 |
2011 |
|
|
|
|
| 3 |
Σ |
1282 |
1239 |
|
|
|
|
| 4 |
Π |
551 |
533 |
|
|
|
|
| 4 |
Π |
503 |
486 |
|
|
|
|
| 5 |
Π |
377 |
365 |
|
|
|
|
| 5 |
Π |
441i |
426i |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 3901.9 cm
-1
Scaled (by 0.9668) Zero Point Vibrational Energy (zpe) 3772.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/aug-cc-pVTZ
Point Group is C∞v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.019 |
| C2 |
0.000 |
0.000 |
-1.247 |
| O3 |
0.000 |
0.000 |
1.210 |
| H4 |
0.000 |
0.000 |
-2.310 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.2667 | 1.1904 | 2.3293 |
C2 | 1.2667 | | 2.4571 | 1.0626 | O3 | 1.1904 | 2.4571 | | 3.5197 | H4 | 2.3293 | 1.0626 | 3.5197 | |
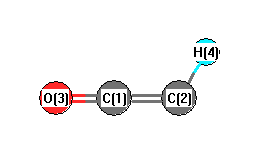 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
180.000 |
|
C2 |
C1 |
O3 |
180.000 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -151.692633 |
| Energy at 298.15K | |
| HF Energy | -151.140044 |
| Nuclear repulsion energy | 52.076959 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3312 |
3202 |
|
|
|
|
| 2 |
A' |
2049 |
1981 |
|
|
|
|
| 3 |
A' |
1210 |
1170 |
|
|
|
|
| 4 |
A' |
577 |
558 |
|
|
|
|
| 5 |
A' |
518 |
501 |
|
|
|
|
| 6 |
A" |
490 |
474 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 4078.3 cm
-1
Scaled (by 0.9668) Zero Point Vibrational Energy (zpe) 3942.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.060 |
0.000 |
| C2 |
1.122 |
-0.615 |
0.000 |
| O3 |
-1.110 |
0.452 |
0.000 |
| H4 |
2.146 |
-0.288 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
| C1 | | 1.3096 | 1.1770 | 2.1743 |
C2 | 1.3096 | | 2.4739 | 1.0751 | O3 | 1.1770 | 2.4739 | | 3.3391 | H4 | 2.1743 | 1.0751 | 3.3391 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H4 |
131.247 |
|
C2 |
C1 |
O3 |
168.401 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
