Jump to
S1C2
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -93.784019 |
| Energy at 298.15K | -93.785310 |
| HF Energy | -93.433438 |
| Nuclear repulsion energy | 28.539593 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3446 |
3298 |
12.00 |
|
|
|
| 2 |
A' |
3041 |
2910 |
134.73 |
|
|
|
| 3 |
A' |
2255 |
2158 |
309.50 |
|
|
|
| 4 |
A' |
1042 |
997 |
122.90 |
|
|
|
| 5 |
A' |
812 |
777 |
183.76 |
|
|
|
| 6 |
A" |
935 |
895 |
3.20 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5765.2 cm
-1
Scaled (by 0.957) Zero Point Vibrational Energy (zpe) 5517.3 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/aug-cc-pVTZ
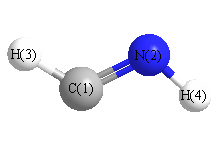
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.110 |
0.627 |
0.000 |
| N2 |
0.110 |
-0.579 |
0.000 |
| H3 |
-0.663 |
1.404 |
0.000 |
| H4 |
-0.760 |
-1.111 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2064 | 1.0957 | 1.9433 |
N2 | 1.2064 | | 2.1284 | 1.0193 | H3 | 1.0957 | 2.1284 | | 2.5163 | H4 | 1.9433 | 1.0193 | 2.5163 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
121.420 |
|
H3 |
C1 |
N2 |
135.138 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -93.791890 |
| Energy at 298.15K | -93.793223 |
| HF Energy | -93.439114 |
| Nuclear repulsion energy | 28.488238 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3544 |
3392 |
38.47 |
|
|
|
| 2 |
A' |
3075 |
2943 |
85.51 |
|
|
|
| 3 |
A' |
2253 |
2156 |
331.30 |
|
|
|
| 4 |
A' |
1192 |
1141 |
13.53 |
|
|
|
| 5 |
A' |
866 |
829 |
333.22 |
|
|
|
| 6 |
A" |
971 |
929 |
143.30 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 5950.3 cm
-1
Scaled (by 0.957) Zero Point Vibrational Energy (zpe) 5694.5 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.001 |
0.639 |
0.000 |
| N2 |
-0.001 |
-0.575 |
0.000 |
| H3 |
0.886 |
1.280 |
0.000 |
| H4 |
-0.878 |
-1.085 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
N2 |
H3 |
H4 |
| C1 | | 1.2137 | 1.0941 | 1.9340 |
N2 | 1.2137 | | 2.0559 | 1.0144 | H3 | 1.0941 | 2.0559 | | 2.9500 | H4 | 1.9340 | 1.0144 | 2.9500 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N2 |
H4 |
120.185 |
|
H3 |
C1 |
N2 |
125.880 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
