Jump to
S1C2
S1C3
Energy calculated at B3PW91/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -225.280934 |
| Energy at 298.15K | -225.287049 |
| HF Energy | -225.280934 |
| Nuclear repulsion energy | 124.036873 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3710 |
3581 |
41.36 |
|
|
|
| 2 |
A |
3597 |
3471 |
3.51 |
|
|
|
| 3 |
A |
1797 |
1735 |
454.04 |
|
|
|
| 4 |
A |
1626 |
1569 |
0.00 |
|
|
|
| 5 |
A |
1174 |
1133 |
4.06 |
|
|
|
| 6 |
A |
960 |
927 |
10.35 |
|
|
|
| 7 |
A |
529 |
511 |
49.31 |
|
|
|
| 8 |
A |
472 |
456 |
1.53 |
|
|
|
| 9 |
A |
370 |
357 |
64.66 |
|
|
|
| 10 |
B |
3709 |
3580 |
33.59 |
|
|
|
| 11 |
B |
3591 |
3466 |
48.77 |
|
|
|
| 12 |
B |
1627 |
1570 |
194.85 |
|
|
|
| 13 |
B |
1417 |
1368 |
211.81 |
|
|
|
| 14 |
B |
1041 |
1005 |
24.86 |
|
|
|
| 15 |
B |
782 |
754 |
39.66 |
|
|
|
| 16 |
B |
578 |
558 |
59.39 |
|
|
|
| 17 |
B |
533 |
514 |
120.69 |
|
|
|
| 18 |
B |
437 |
422 |
210.79 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13975.1 cm
-1
Scaled (by 0.9651) Zero Point Vibrational Energy (zpe) 13487.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/aug-cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.145 |
| O2 |
0.000 |
0.000 |
1.357 |
| N3 |
0.000 |
1.158 |
-0.610 |
| N4 |
0.000 |
-1.158 |
-0.610 |
| H5 |
0.196 |
1.988 |
-0.075 |
| H6 |
0.425 |
1.135 |
-1.522 |
| H7 |
-0.196 |
-1.988 |
-0.075 |
| H8 |
-0.425 |
-1.135 |
-1.522 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2116 | 1.3827 | 1.3827 | 2.0097 | 2.0608 | 2.0097 | 2.0608 |
O2 | 1.2116 | | 2.2823 | 2.2823 | 2.4576 | 3.1233 | 2.4576 | 3.1233 | N3 | 1.3827 | 2.2823 | | 2.3168 | 1.0064 | 1.0066 | 3.1975 | 2.5041 | N4 | 1.3827 | 2.2823 | 2.3168 | | 3.1975 | 2.5041 | 1.0064 | 1.0066 | H5 | 2.0097 | 2.4576 | 1.0064 | 3.1975 | | 1.6956 | 3.9952 | 3.4972 | H6 | 2.0608 | 3.1233 | 1.0066 | 2.5041 | 1.6956 | | 3.4972 | 2.4230 | H7 | 2.0097 | 2.4576 | 3.1975 | 1.0064 | 3.9952 | 3.4972 | | 1.6956 | H8 | 2.0608 | 3.1233 | 2.5041 | 1.0066 | 3.4972 | 2.4230 | 1.6956 | |
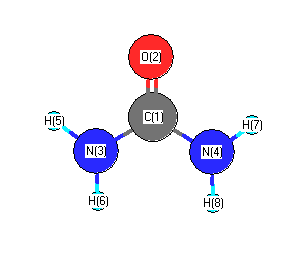 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
113.600 |
|
C1 |
N3 |
H6 |
118.357 |
| C1 |
N4 |
H7 |
113.600 |
|
C1 |
N4 |
H8 |
118.357 |
| O2 |
C1 |
N3 |
123.091 |
|
O2 |
C1 |
N4 |
123.091 |
| N3 |
C1 |
N4 |
113.819 |
|
H5 |
N3 |
H6 |
114.775 |
| H7 |
N4 |
H8 |
114.775 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/aug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.530 |
|
|
|
| 2 |
O |
-0.581 |
|
|
|
| 3 |
N |
-0.155 |
|
|
|
| 4 |
N |
-0.155 |
|
|
|
| 5 |
H |
0.108 |
|
|
|
| 6 |
H |
0.072 |
|
|
|
| 7 |
H |
0.108 |
|
|
|
| 8 |
H |
0.072 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-3.845 |
3.845 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-25.250 |
3.004 |
0.000 |
| y |
3.004 |
-17.833 |
0.000 |
| z |
0.000 |
0.000 |
-25.266 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-3.701 |
3.004 |
0.000 |
| y |
3.004 |
7.425 |
0.000 |
| z |
0.000 |
0.000 |
-3.724 |
|
| Polar |
|---|
| 3z2-r2 | -7.449 |
|---|
| x2-y2 | -7.417 |
|---|
| xy | 3.004 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.955 |
0.084 |
0.000 |
| y |
0.084 |
6.038 |
0.000 |
| z |
0.000 |
0.000 |
6.068 |
<r2> (average value of r
2) Å
2
| <r2> |
68.557 |
| (<r2>)1/2 |
8.280 |
Jump to
S1C1
S1C3
Energy calculated at B3PW91/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -225.280934 |
| Energy at 298.15K | -225.287049 |
| HF Energy | -225.280934 |
| Nuclear repulsion energy | 124.036873 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/aug-cc-pVTZ
Geometric Data calculated at B3PW91/aug-cc-pVTZ
Point Group is C2
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at B3PW91/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -225.279538 |
| Energy at 298.15K | |
| HF Energy | -225.279538 |
| Nuclear repulsion energy | 124.204291 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at B3PW91/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3770 |
3638 |
54.59 |
|
|
|
| 2 |
A1 |
3640 |
3513 |
5.81 |
|
|
|
| 3 |
A1 |
1782 |
1720 |
505.18 |
|
|
|
| 4 |
A1 |
1621 |
1564 |
0.49 |
|
|
|
| 5 |
A1 |
1142 |
1102 |
0.92 |
|
|
|
| 6 |
A1 |
974 |
940 |
10.83 |
|
|
|
| 7 |
A1 |
476 |
459 |
3.45 |
|
|
|
| 8 |
A2 |
377 |
364 |
0.00 |
|
|
|
| 9 |
A2 |
343i |
331i |
0.00 |
|
|
|
| 10 |
B1 |
778 |
750 |
4.71 |
|
|
|
| 11 |
B1 |
568 |
548 |
7.47 |
|
|
|
| 12 |
B1 |
233i |
225i |
416.64 |
|
|
|
| 13 |
B2 |
3768 |
3636 |
44.04 |
|
|
|
| 14 |
B2 |
3631 |
3504 |
68.41 |
|
|
|
| 15 |
B2 |
1616 |
1560 |
271.24 |
|
|
|
| 16 |
B2 |
1424 |
1375 |
217.38 |
|
|
|
| 17 |
B2 |
985 |
951 |
15.19 |
|
|
|
| 18 |
B2 |
564 |
544 |
13.59 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13268.8 cm
-1
Scaled (by 0.9651) Zero Point Vibrational Energy (zpe) 12805.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at B3PW91/aug-cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.139 |
| O2 |
0.000 |
0.000 |
1.356 |
| N3 |
0.000 |
1.156 |
-0.594 |
| N4 |
0.000 |
-1.156 |
-0.594 |
| H5 |
0.000 |
2.018 |
-0.085 |
| H6 |
0.000 |
1.181 |
-1.596 |
| H7 |
0.000 |
-2.018 |
-0.085 |
| H8 |
0.000 |
-1.181 |
-1.596 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2172 | 1.3688 | 1.3688 | 2.0308 | 2.0988 | 2.0308 | 2.0988 |
O2 | 1.2172 | | 2.2673 | 2.2673 | 2.4800 | 3.1795 | 2.4800 | 3.1795 | N3 | 1.3688 | 2.2673 | | 2.3115 | 1.0020 | 1.0016 | 3.2149 | 2.5427 | N4 | 1.3688 | 2.2673 | 2.3115 | | 3.2149 | 2.5427 | 1.0020 | 1.0016 | H5 | 2.0308 | 2.4800 | 1.0020 | 3.2149 | | 1.7274 | 4.0369 | 3.5387 | H6 | 2.0988 | 3.1795 | 1.0016 | 2.5427 | 1.7274 | | 3.5387 | 2.3629 | H7 | 2.0308 | 2.4800 | 3.2149 | 1.0020 | 4.0369 | 3.5387 | | 1.7274 | H8 | 2.0988 | 3.1795 | 2.5427 | 1.0016 | 3.5387 | 2.3629 | 1.7274 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
117.025 |
|
C1 |
N3 |
H6 |
123.866 |
| C1 |
N4 |
H7 |
117.025 |
|
C1 |
N4 |
H8 |
123.866 |
| O2 |
C1 |
N3 |
122.398 |
|
O2 |
C1 |
N4 |
122.398 |
| N3 |
C1 |
N4 |
115.204 |
|
H5 |
N3 |
H6 |
119.109 |
| H7 |
N4 |
H8 |
119.109 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/aug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.550 |
|
|
|
| 2 |
O |
-0.561 |
|
|
|
| 3 |
N |
-0.134 |
|
|
|
| 4 |
N |
-0.134 |
|
|
|
| 5 |
H |
0.080 |
|
|
|
| 6 |
H |
0.059 |
|
|
|
| 7 |
H |
0.080 |
|
|
|
| 8 |
H |
0.059 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.000 |
0.000 |
-4.349 |
4.349 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-26.113 |
0.000 |
0.000 |
| y |
0.000 |
-16.760 |
0.000 |
| z |
0.000 |
0.000 |
-24.441 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-5.512 |
0.000 |
0.000 |
| y |
0.000 |
8.517 |
0.000 |
| z |
0.000 |
0.000 |
-3.004 |
|
| Polar |
|---|
| 3z2-r2 | -6.009 |
|---|
| x2-y2 | -9.353 |
|---|
| xy | 0.000 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
3.945 |
0.000 |
0.000 |
| y |
0.000 |
6.093 |
0.000 |
| z |
0.000 |
0.000 |
6.089 |
<r2> (average value of r
2) Å
2
| <r2> |
68.539 |
| (<r2>)1/2 |
8.279 |
