Jump to
S1C2
S1C3
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -224.915322 |
| Energy at 298.15K | -224.921558 |
| HF Energy | -224.085233 |
| Nuclear repulsion energy | 124.175079 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3737 |
3576 |
50.02 |
|
|
|
| 2 |
A |
3625 |
3469 |
5.26 |
|
|
|
| 3 |
A |
1853 |
1773 |
482.14 |
|
|
|
| 4 |
A |
1664 |
1593 |
0.01 |
|
|
|
| 5 |
A |
1211 |
1159 |
5.36 |
|
|
|
| 6 |
A |
973 |
931 |
11.67 |
|
|
|
| 7 |
A |
591 |
566 |
82.88 |
|
|
|
| 8 |
A |
482 |
461 |
1.39 |
|
|
|
| 9 |
A |
382 |
365 |
72.61 |
|
|
|
| 10 |
B |
3736 |
3576 |
28.71 |
|
|
|
| 11 |
B |
3622 |
3466 |
57.99 |
|
|
|
| 12 |
B |
1670 |
1598 |
183.69 |
|
|
|
| 13 |
B |
1445 |
1383 |
209.61 |
|
|
|
| 14 |
B |
1082 |
1035 |
25.66 |
|
|
|
| 15 |
B |
807 |
772 |
65.83 |
|
|
|
| 16 |
B |
593 |
567 |
95.61 |
|
|
|
| 17 |
B |
553 |
529 |
189.11 |
|
|
|
| 18 |
B |
454 |
435 |
93.37 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14238.6 cm
-1
Scaled (by 0.957) Zero Point Vibrational Energy (zpe) 13626.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.145 |
| O2 |
0.000 |
0.000 |
1.354 |
| N3 |
0.000 |
1.158 |
-0.610 |
| N4 |
0.000 |
-1.158 |
-0.610 |
| H5 |
0.228 |
1.975 |
-0.071 |
| H6 |
0.448 |
1.115 |
-1.508 |
| H7 |
-0.228 |
-1.975 |
-0.071 |
| H8 |
-0.448 |
-1.115 |
-1.508 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2091 | 1.3820 | 1.3820 | 1.9998 | 2.0437 | 1.9998 | 2.0437 |
O2 | 1.2091 | | 2.2798 | 2.2798 | 2.4462 | 3.1043 | 2.4462 | 3.1043 | N3 | 1.3820 | 2.2798 | | 2.3154 | 1.0049 | 1.0046 | 3.1868 | 2.4847 | N4 | 1.3820 | 2.2798 | 2.3154 | | 3.1868 | 2.4847 | 1.0049 | 1.0046 | H5 | 1.9998 | 2.4462 | 1.0049 | 3.1868 | | 1.6889 | 3.9762 | 3.4742 | H6 | 2.0437 | 3.1043 | 1.0046 | 2.4847 | 1.6889 | | 3.4742 | 2.4035 | H7 | 1.9998 | 2.4462 | 3.1868 | 1.0049 | 3.9762 | 3.4742 | | 1.6889 | H8 | 2.0437 | 3.1043 | 2.4847 | 1.0046 | 3.4742 | 2.4035 | 1.6889 | |
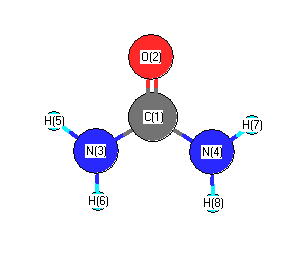 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
112.865 |
|
C1 |
N3 |
H6 |
116.927 |
| C1 |
N4 |
H7 |
112.865 |
|
C1 |
N4 |
H8 |
116.927 |
| O2 |
C1 |
N3 |
123.103 |
|
O2 |
C1 |
N4 |
123.103 |
| N3 |
C1 |
N4 |
113.794 |
|
H5 |
N3 |
H6 |
114.366 |
| H7 |
N4 |
H8 |
114.366 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C3
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -224.913725 |
| Energy at 298.15K | -224.919553 |
| HF Energy | -224.084051 |
| Nuclear repulsion energy | 124.239993 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3768 |
3606 |
54.87 |
|
|
|
| 2 |
A' |
3649 |
3492 |
9.15 |
|
|
|
| 3 |
A' |
1840 |
1761 |
513.97 |
|
|
|
| 4 |
A' |
1673 |
1601 |
11.32 |
|
|
|
| 5 |
A' |
1203 |
1151 |
5.28 |
|
|
|
| 6 |
A' |
981 |
939 |
15.84 |
|
|
|
| 7 |
A' |
797 |
763 |
25.38 |
|
|
|
| 8 |
A' |
547 |
523 |
32.46 |
|
|
|
| 9 |
A' |
489 |
468 |
23.50 |
|
|
|
| 10 |
A' |
417 |
399 |
338.86 |
|
|
|
| 11 |
A" |
3765 |
3603 |
30.77 |
|
|
|
| 12 |
A" |
3641 |
3485 |
64.26 |
|
|
|
| 13 |
A" |
1662 |
1590 |
208.03 |
|
|
|
| 14 |
A" |
1449 |
1387 |
211.76 |
|
|
|
| 15 |
A" |
1047 |
1002 |
21.21 |
|
|
|
| 16 |
A" |
585 |
560 |
25.56 |
|
|
|
| 17 |
A" |
422 |
404 |
71.93 |
|
|
|
| 18 |
A" |
192 |
184 |
34.48 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 14062.3 cm
-1
Scaled (by 0.957) Zero Point Vibrational Energy (zpe) 13457.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.010 |
0.141 |
0.000 |
| O2 |
0.038 |
1.352 |
0.000 |
| N3 |
0.038 |
-0.600 |
1.160 |
| N4 |
0.038 |
-0.600 |
-1.160 |
| H5 |
-0.168 |
-0.074 |
1.989 |
| H6 |
-0.277 |
-1.552 |
1.151 |
| H7 |
-0.168 |
-0.074 |
-1.989 |
| H8 |
-0.277 |
-1.552 |
-1.151 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2110 | 1.3768 | 1.3768 | 2.0085 | 2.0674 | 2.0085 | 2.0674 |
O2 | 1.2110 | | 2.2706 | 2.2706 | 2.4560 | 3.1394 | 2.4560 | 3.1394 | N3 | 1.3768 | 2.2706 | | 2.3197 | 1.0031 | 1.0023 | 3.1991 | 2.5193 | N4 | 1.3768 | 2.2706 | 2.3197 | | 3.1991 | 2.5193 | 1.0031 | 1.0023 | H5 | 2.0085 | 2.4560 | 1.0031 | 3.1991 | | 1.7019 | 3.9779 | 3.4724 | H6 | 2.0674 | 3.1394 | 1.0023 | 2.5193 | 1.7019 | | 3.4724 | 2.3030 | H7 | 2.0085 | 2.4560 | 3.1991 | 1.0031 | 3.9779 | 3.4724 | | 1.7019 | H8 | 2.0674 | 3.1394 | 2.5193 | 1.0023 | 3.4724 | 2.3030 | 1.7019 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
114.190 |
|
C1 |
N3 |
H6 |
119.854 |
| C1 |
N4 |
H7 |
114.190 |
|
C1 |
N4 |
H8 |
119.854 |
| O2 |
C1 |
N3 |
122.537 |
|
O2 |
C1 |
N4 |
122.537 |
| N3 |
C1 |
N4 |
114.792 |
|
H5 |
N3 |
H6 |
116.132 |
| H7 |
N4 |
H8 |
116.132 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
S1C2
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -224.912902 |
| Energy at 298.15K | |
| HF Energy | -224.084069 |
| Nuclear repulsion energy | 124.434296 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1 |
3815 |
3651 |
74.01 |
|
|
|
| 2 |
A1 |
3683 |
3524 |
10.18 |
|
|
|
| 3 |
A1 |
1827 |
1749 |
568.01 |
|
|
|
| 4 |
A1 |
1656 |
1584 |
0.80 |
|
|
|
| 5 |
A1 |
1167 |
1117 |
1.17 |
|
|
|
| 6 |
A1 |
991 |
949 |
15.24 |
|
|
|
| 7 |
A1 |
486 |
465 |
2.78 |
|
|
|
| 8 |
A2 |
379 |
363 |
0.00 |
|
|
|
| 9 |
A2 |
411i |
393i |
0.00 |
|
|
|
| 10 |
B1 |
797 |
763 |
5.52 |
|
|
|
| 11 |
B1 |
574 |
549 |
7.48 |
|
|
|
| 12 |
B1 |
322i |
308i |
444.63 |
|
|
|
| 13 |
B2 |
3813 |
3649 |
37.56 |
|
|
|
| 14 |
B2 |
3676 |
3517 |
91.81 |
|
|
|
| 15 |
B2 |
1655 |
1584 |
279.01 |
|
|
|
| 16 |
B2 |
1454 |
1391 |
227.32 |
|
|
|
| 17 |
B2 |
1012 |
968 |
13.78 |
|
|
|
| 18 |
B2 |
577 |
552 |
15.01 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13413.4 cm
-1
Scaled (by 0.957) Zero Point Vibrational Energy (zpe) 12836.6 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is C2v
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.141 |
| O2 |
0.000 |
0.000 |
1.354 |
| N3 |
0.000 |
1.153 |
-0.594 |
| N4 |
0.000 |
-1.153 |
-0.594 |
| H5 |
0.000 |
2.016 |
-0.089 |
| H6 |
0.000 |
1.169 |
-1.593 |
| H7 |
0.000 |
-2.016 |
-0.089 |
| H8 |
0.000 |
-1.169 |
-1.593 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
N4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.2131 | 1.3679 | 1.3679 | 2.0295 | 2.0913 | 2.0295 | 2.0913 |
O2 | 1.2131 | | 2.2643 | 2.2643 | 2.4798 | 3.1706 | 2.4798 | 3.1706 | N3 | 1.3679 | 2.2643 | | 2.3068 | 1.0000 | 0.9990 | 3.2098 | 2.5279 | N4 | 1.3679 | 2.2643 | 2.3068 | | 3.2098 | 2.5279 | 1.0000 | 0.9990 | H5 | 2.0295 | 2.4798 | 1.0000 | 3.2098 | | 1.7264 | 4.0328 | 3.5225 | H6 | 2.0913 | 3.1706 | 0.9990 | 2.5279 | 1.7264 | | 3.5225 | 2.3376 | H7 | 2.0295 | 2.4798 | 3.2098 | 1.0000 | 4.0328 | 3.5225 | | 1.7264 | H8 | 2.0913 | 3.1706 | 2.5279 | 0.9990 | 3.5225 | 2.3376 | 1.7264 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
117.140 |
|
C1 |
N3 |
H6 |
123.402 |
| C1 |
N4 |
H7 |
117.140 |
|
C1 |
N4 |
H8 |
123.402 |
| O2 |
C1 |
N3 |
122.519 |
|
O2 |
C1 |
N4 |
122.519 |
| N3 |
C1 |
N4 |
114.962 |
|
H5 |
N3 |
H6 |
119.458 |
| H7 |
N4 |
H8 |
119.458 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
