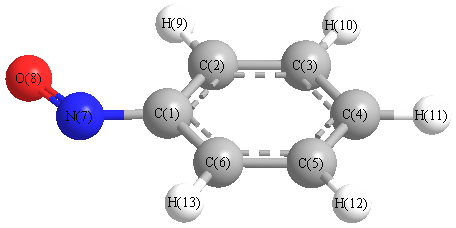
Vibrational Frequencies calculated at B1B95/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3233 |
3099 |
4.79 |
|
|
|
| 2 |
A' |
3224 |
3091 |
8.93 |
|
|
|
| 3 |
A' |
3214 |
3081 |
7.62 |
|
|
|
| 4 |
A' |
3208 |
3074 |
3.92 |
|
|
|
| 5 |
A' |
3196 |
3064 |
0.42 |
|
|
|
| 6 |
A' |
1684 |
1614 |
44.18 |
|
|
|
| 7 |
A' |
1666 |
1597 |
0.89 |
|
|
|
| 8 |
A' |
1621 |
1554 |
173.19 |
|
|
|
| 9 |
A' |
1514 |
1451 |
6.92 |
|
|
|
| 10 |
A' |
1498 |
1436 |
24.92 |
|
|
|
| 11 |
A' |
1384 |
1326 |
13.80 |
|
|
|
| 12 |
A' |
1343 |
1287 |
8.45 |
|
|
|
| 13 |
A' |
1206 |
1156 |
21.21 |
|
|
|
| 14 |
A' |
1181 |
1132 |
2.78 |
|
|
|
| 15 |
A' |
1144 |
1097 |
136.19 |
|
|
|
| 16 |
A' |
1108 |
1062 |
4.89 |
|
|
|
| 17 |
A' |
1050 |
1006 |
4.12 |
|
|
|
| 18 |
A' |
1023 |
980 |
0.59 |
|
|
|
| 19 |
A' |
840 |
805 |
38.11 |
|
|
|
| 20 |
A' |
680 |
652 |
9.21 |
|
|
|
| 21 |
A' |
614 |
588 |
0.07 |
|
|
|
| 22 |
A' |
448 |
429 |
1.02 |
|
|
|
| 23 |
A' |
263 |
252 |
1.85 |
|
|
|
| 24 |
A" |
1028 |
986 |
0.05 |
|
|
|
| 25 |
A" |
1012 |
970 |
0.01 |
|
|
|
| 26 |
A" |
971 |
931 |
4.01 |
|
|
|
| 27 |
A" |
872 |
836 |
0.02 |
|
|
|
| 28 |
A" |
785 |
752 |
59.01 |
|
|
|
| 29 |
A" |
695 |
666 |
22.28 |
|
|
|
| 30 |
A" |
470 |
450 |
2.20 |
|
|
|
| 31 |
A" |
416 |
399 |
0.00 |
|
|
|
| 32 |
A" |
246 |
236 |
0.31 |
|
|
|
| 33 |
A" |
106 |
102 |
0.10 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 21470.9 cm
-1
Scaled (by 0.9585) Zero Point Vibrational Energy (zpe) 20579.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B1B95/aug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.949 |
|
|
|
| 2 |
C |
-0.686 |
|
|
|
| 3 |
C |
-0.449 |
|
|
|
| 4 |
C |
-0.529 |
|
|
|
| 5 |
C |
-0.517 |
|
|
|
| 6 |
C |
-0.905 |
|
|
|
| 7 |
N |
-0.382 |
|
|
|
| 8 |
O |
-0.393 |
|
|
|
| 9 |
H |
0.697 |
|
|
|
| 10 |
H |
0.495 |
|
|
|
| 11 |
H |
0.556 |
|
|
|
| 12 |
H |
0.430 |
|
|
|
| 13 |
H |
0.733 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
1.111 |
-3.535 |
0.000 |
3.705 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-41.759 |
2.313 |
0.000 |
| y |
2.313 |
-48.611 |
0.000 |
| z |
0.000 |
0.000 |
-47.532 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
6.312 |
2.313 |
0.000 |
| y |
2.313 |
-3.965 |
0.000 |
| z |
0.000 |
0.000 |
-2.347 |
|
| Polar |
|---|
| 3z2-r2 | -4.694 |
|---|
| x2-y2 | 6.852 |
|---|
| xy | 2.313 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
13.548 |
-1.649 |
0.000 |
| y |
-1.649 |
16.416 |
-0.000 |
| z |
0.000 |
-0.000 |
6.972 |
<r2> (average value of r
2) Å
2
| <r2> |
246.333 |
| (<r2>)1/2 |
15.695 |