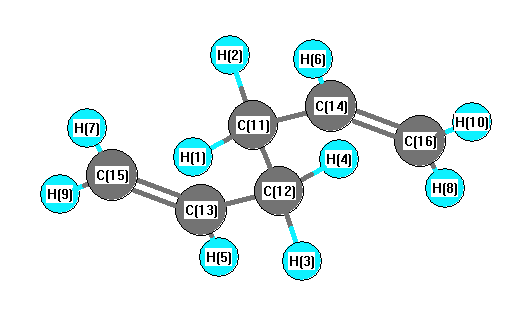
All results from a given calculation for C6H10 (1,5-Hexadiene)
using model chemistry: CCD/aug-cc-pVTZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
C2 |
1Ag |
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -234.130639 |
| Energy at 298.15K | |
| HF Energy | -233.069982 |
| Nuclear repulsion energy | 212.775611 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is C2
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.424 |
0.641 |
-0.302 |
| C2 |
0.424 |
-0.641 |
-0.302 |
| C3 |
0.424 |
1.878 |
-0.340 |
| C4 |
-0.424 |
-1.878 |
-0.340 |
| C5 |
0.442 |
2.814 |
0.606 |
| C6 |
-0.442 |
-2.814 |
0.606 |
| H7 |
-1.085 |
0.624 |
-1.174 |
| H8 |
-1.062 |
0.655 |
0.584 |
| H9 |
1.085 |
-0.624 |
-1.174 |
| H10 |
1.062 |
-0.655 |
0.584 |
| H11 |
1.073 |
1.989 |
-1.204 |
| H12 |
-1.073 |
-1.989 |
-1.204 |
| H13 |
1.083 |
3.682 |
0.530 |
| H14 |
-0.187 |
2.738 |
1.484 |
| H15 |
-1.083 |
-3.682 |
0.530 |
| H16 |
0.187 |
-2.738 |
1.484 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
C6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
H14 |
H15 |
H16 |
| C1 | | 1.5362 | 1.5002 | 2.5190 | 2.5097 | 3.5721 | 1.0947 | 1.0921 | 2.1533 | 2.1616 | 2.2070 | 2.8548 | 3.4947 | 2.7649 | 4.4513 | 3.8701 |
C2 | 1.5362 | | 2.5190 | 1.5002 | 3.5721 | 2.5097 | 2.1533 | 2.1616 | 1.0947 | 1.0921 | 2.8548 | 2.2070 | 4.4513 | 3.8701 | 3.4947 | 2.7649 | C3 | 1.5002 | 2.5190 | | 3.8506 | 1.3308 | 4.8643 | 2.1322 | 2.1347 | 2.7190 | 2.7711 | 1.0865 | 4.2356 | 2.1086 | 2.1071 | 5.8261 | 4.9689 | C4 | 2.5190 | 1.5002 | 3.8506 | | 4.8643 | 1.3308 | 2.7190 | 2.7711 | 2.1322 | 2.1347 | 4.2356 | 1.0865 | 5.8261 | 4.9689 | 2.1086 | 2.1071 | C5 | 2.5097 | 3.5721 | 1.3308 | 4.8643 | | 5.6973 | 3.2093 | 2.6316 | 3.9244 | 3.5244 | 2.0868 | 5.3518 | 1.0815 | 1.0834 | 6.6734 | 5.6267 | C6 | 3.5721 | 2.5097 | 4.8643 | 1.3308 | 5.6973 | | 3.9244 | 3.5244 | 3.2093 | 2.6316 | 5.3518 | 2.0868 | 6.6734 | 5.6267 | 1.0815 | 1.0834 | H7 | 1.0947 | 2.1533 | 2.1322 | 2.7190 | 3.2093 | 3.9244 | | 1.7582 | 2.5038 | 3.0559 | 2.5538 | 2.6130 | 4.1181 | 3.5132 | 4.6309 | 4.4705 | H8 | 1.0921 | 2.1616 | 2.1347 | 2.7711 | 2.6316 | 3.5244 | 1.7582 | | 3.0559 | 2.4964 | 3.0877 | 3.1920 | 3.7105 | 2.4319 | 4.3379 | 3.7261 | H9 | 2.1533 | 1.0947 | 2.7190 | 2.1322 | 3.9244 | 3.2093 | 2.5038 | 3.0559 | | 1.7582 | 2.6130 | 2.5538 | 4.6309 | 4.4705 | 4.1181 | 3.5132 | H10 | 2.1616 | 1.0921 | 2.7711 | 2.1347 | 3.5244 | 2.6316 | 3.0559 | 2.4964 | 1.7582 | | 3.1920 | 3.0877 | 4.3379 | 3.7261 | 3.7105 | 2.4319 | H11 | 2.2070 | 2.8548 | 1.0865 | 4.2356 | 2.0868 | 5.3518 | 2.5538 | 3.0877 | 2.6130 | 3.1920 | | 4.5195 | 2.4238 | 3.0620 | 6.3100 | 5.5095 | H12 | 2.8548 | 2.2070 | 4.2356 | 1.0865 | 5.3518 | 2.0868 | 2.6130 | 3.1920 | 2.5538 | 3.0877 | 4.5195 | | 6.3100 | 5.5095 | 2.4238 | 3.0620 | H13 | 3.4947 | 4.4513 | 2.1086 | 5.8261 | 1.0815 | 6.6734 | 4.1181 | 3.7105 | 4.6309 | 4.3379 | 2.4238 | 6.3100 | | 1.8481 | 7.6763 | 6.5520 | H14 | 2.7649 | 3.8701 | 2.1071 | 4.9689 | 1.0834 | 5.6267 | 3.5132 | 2.4319 | 4.4705 | 3.7261 | 3.0620 | 5.5095 | 1.8481 | | 6.5520 | 5.4882 | H15 | 4.4513 | 3.4947 | 5.8261 | 2.1086 | 6.6734 | 1.0815 | 4.6309 | 4.3379 | 4.1181 | 3.7105 | 6.3100 | 2.4238 | 7.6763 | 6.5520 | | 1.8481 | H16 | 3.8701 | 2.7649 | 4.9689 | 2.1071 | 5.6267 | 1.0834 | 4.4705 | 3.7261 | 3.5132 | 2.4319 | 5.5095 | 3.0620 | 6.5520 | 5.4882 | 1.8481 | |
 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C4 |
112.110 |
|
C1 |
C2 |
H9 |
108.697 |
| C1 |
C2 |
H10 |
109.495 |
|
C1 |
C3 |
C5 |
124.757 |
| C1 |
C3 |
H11 |
116.209 |
|
C2 |
C1 |
C3 |
112.110 |
| C2 |
C1 |
H7 |
108.697 |
|
C2 |
C1 |
H8 |
109.495 |
| C2 |
C4 |
C6 |
124.757 |
|
C2 |
C4 |
H12 |
116.209 |
| C3 |
C1 |
H7 |
109.511 |
|
C3 |
C1 |
H8 |
109.865 |
| C3 |
C5 |
H13 |
121.534 |
|
C3 |
C5 |
H14 |
121.234 |
| C4 |
C2 |
H9 |
109.511 |
|
C4 |
C2 |
H10 |
109.865 |
| C4 |
C6 |
H15 |
121.534 |
|
C4 |
C6 |
H16 |
121.234 |
| C5 |
C3 |
H11 |
119.027 |
|
C6 |
C4 |
H12 |
119.027 |
| H7 |
C1 |
H8 |
107.024 |
|
H9 |
C2 |
H10 |
107.024 |
| H13 |
C5 |
H14 |
117.232 |
|
H15 |
C6 |
H16 |
117.232 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
