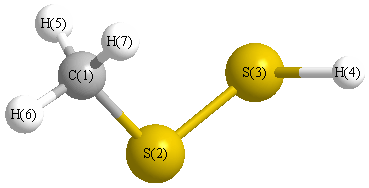
Vibrational Frequencies calculated at B2PLYP/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3164 |
3041 |
4.49 |
54.64 |
0.75 |
0.86 |
| 2 |
A |
3147 |
3024 |
4.72 |
77.81 |
0.75 |
0.86 |
| 3 |
A |
3059 |
2940 |
16.52 |
196.62 |
0.03 |
0.05 |
| 4 |
A |
2658 |
2554 |
3.34 |
161.35 |
0.21 |
0.35 |
| 5 |
A |
1493 |
1435 |
8.21 |
4.03 |
0.71 |
0.83 |
| 6 |
A |
1472 |
1415 |
8.40 |
5.19 |
0.75 |
0.86 |
| 7 |
A |
1355 |
1302 |
3.04 |
0.78 |
0.13 |
0.23 |
| 8 |
A |
984 |
946 |
4.67 |
1.28 |
0.08 |
0.14 |
| 9 |
A |
981 |
942 |
2.66 |
0.51 |
0.12 |
0.21 |
| 10 |
A |
886 |
851 |
4.61 |
9.53 |
0.40 |
0.57 |
| 11 |
A |
700 |
672 |
1.30 |
14.64 |
0.23 |
0.38 |
| 12 |
A |
509 |
489 |
0.64 |
13.61 |
0.13 |
0.22 |
| 13 |
A |
320 |
307 |
12.13 |
0.76 |
0.75 |
0.86 |
| 14 |
A |
239 |
230 |
0.15 |
5.26 |
0.50 |
0.67 |
| 15 |
A |
170 |
164 |
0.32 |
0.01 |
0.70 |
0.82 |
Unscaled Zero Point Vibrational Energy (zpe) 10567.2 cm
-1
Scaled (by 0.961) Zero Point Vibrational Energy (zpe) 10155.1 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B2PLYP/aug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
-0.702 |
|
|
|
| 2 |
S |
-0.031 |
|
|
|
| 3 |
S |
-0.199 |
|
|
|
| 4 |
H |
0.137 |
|
|
|
| 5 |
H |
0.280 |
|
|
|
| 6 |
H |
0.251 |
|
|
|
| 7 |
H |
0.265 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
-1.051 |
1.182 |
0.752 |
1.751 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-31.265 |
-0.262 |
1.547 |
| y |
-0.262 |
-35.377 |
0.586 |
| z |
1.547 |
0.586 |
-33.531 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
3.189 |
-0.262 |
1.547 |
| y |
-0.262 |
-2.979 |
0.586 |
| z |
1.547 |
0.586 |
-0.210 |
|
| Polar |
|---|
| 3z2-r2 | -0.420 |
|---|
| x2-y2 | 4.112 |
|---|
| xy | -0.262 |
|---|
| xz | 1.547 |
|---|
| yz | 0.586 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
10.890 |
0.184 |
0.151 |
| y |
0.184 |
8.036 |
0.048 |
| z |
0.151 |
0.048 |
7.013 |
<r2> (average value of r
2) Å
2
| <r2> |
102.870 |
| (<r2>)1/2 |
10.142 |