All results from a given calculation for C5H8 (Ethenylcyclopropane)
using model chemistry: CCD/aug-cc-pVTZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
CS |
1A' |
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -194.890056 |
| Energy at 298.15K | |
| HF Energy | -194.010580 |
| Nuclear repulsion energy | 163.359964 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
-0.396 |
0.290 |
0.000 |
| C2 |
0.255 |
-1.036 |
0.000 |
| C3 |
-0.390 |
-2.201 |
0.000 |
| C4 |
0.255 |
1.428 |
0.750 |
| C5 |
0.255 |
1.428 |
-0.750 |
| H6 |
-1.478 |
0.273 |
0.000 |
| H7 |
1.342 |
-1.030 |
0.000 |
| H8 |
0.145 |
-3.140 |
0.000 |
| H9 |
-1.472 |
-2.245 |
0.000 |
| H10 |
-0.391 |
2.127 |
1.262 |
| H11 |
1.186 |
1.216 |
1.256 |
| H12 |
-0.391 |
2.127 |
-1.262 |
| H13 |
1.186 |
1.216 |
-1.256 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
C3 |
C4 |
C5 |
H6 |
H7 |
H8 |
H9 |
H10 |
H11 |
H12 |
H13 |
| C1 | | 1.4764 | 2.4905 | 1.5112 | 1.5112 | 1.0819 | 2.1817 | 3.4717 | 2.7538 | 2.2288 | 2.2224 | 2.2288 | 2.2224 |
C2 | 1.4764 | | 1.3320 | 2.5757 | 2.5757 | 2.1715 | 1.0867 | 2.1069 | 2.1087 | 3.4655 | 2.7412 | 3.4655 | 2.7412 | C3 | 2.4905 | 1.3320 | | 3.7617 | 3.7617 | 2.7026 | 2.0907 | 1.0809 | 1.0829 | 4.5078 | 3.9670 | 4.5078 | 3.9670 | C4 | 1.5112 | 2.5757 | 3.7617 | | 1.5009 | 2.2135 | 2.7904 | 4.6305 | 4.1281 | 1.0799 | 1.0807 | 2.2257 | 2.2221 | C5 | 1.5112 | 2.5757 | 3.7617 | 1.5009 | | 2.2135 | 2.7904 | 4.6305 | 4.1281 | 2.2257 | 2.2221 | 1.0799 | 1.0807 | H6 | 1.0819 | 2.1715 | 2.7026 | 2.2135 | 2.2135 | | 3.1060 | 3.7793 | 2.5185 | 2.4919 | 3.0925 | 2.4919 | 3.0925 | H7 | 2.1817 | 1.0867 | 2.0907 | 2.7904 | 2.7904 | 3.1060 | | 2.4254 | 3.0652 | 3.8153 | 2.5777 | 3.8153 | 2.5777 | H8 | 3.4717 | 2.1069 | 1.0809 | 4.6305 | 4.6305 | 3.7793 | 2.4254 | | 1.8485 | 5.4420 | 4.6510 | 5.4420 | 4.6510 | H9 | 2.7538 | 2.1087 | 1.0829 | 4.1281 | 4.1281 | 2.5185 | 3.0652 | 1.8485 | | 4.6772 | 4.5415 | 4.6772 | 4.5415 | H10 | 2.2288 | 3.4655 | 4.5078 | 1.0799 | 2.2257 | 2.4919 | 3.8153 | 5.4420 | 4.6772 | | 1.8212 | 2.5236 | 3.1073 | H11 | 2.2224 | 2.7412 | 3.9670 | 1.0807 | 2.2221 | 3.0925 | 2.5777 | 4.6510 | 4.5415 | 1.8212 | | 3.1073 | 2.5116 | H12 | 2.2288 | 3.4655 | 4.5078 | 2.2257 | 1.0799 | 2.4919 | 3.8153 | 5.4420 | 4.6772 | 2.5236 | 3.1073 | | 1.8212 | H13 | 2.2224 | 2.7412 | 3.9670 | 2.2221 | 1.0807 | 3.0925 | 2.5777 | 4.6510 | 4.5415 | 3.1073 | 2.5116 | 1.8212 | |
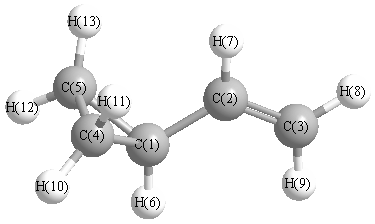 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
C3 |
124.867 |
|
C1 |
C2 |
H7 |
115.854 |
| C1 |
C4 |
C5 |
60.224 |
|
C1 |
C4 |
H10 |
117.709 |
| C1 |
C4 |
H11 |
117.088 |
|
C1 |
C5 |
C4 |
60.224 |
| C1 |
C5 |
H12 |
117.709 |
|
C1 |
C5 |
H13 |
117.088 |
| C2 |
C1 |
C4 |
119.107 |
|
C2 |
C1 |
C5 |
119.107 |
| C2 |
C1 |
H6 |
115.297 |
|
C2 |
C3 |
H8 |
121.317 |
| C2 |
C3 |
H9 |
121.320 |
|
C3 |
C2 |
H7 |
119.279 |
| C4 |
C1 |
H6 |
116.235 |
|
C4 |
C5 |
H12 |
118.262 |
| C4 |
C5 |
H13 |
117.877 |
|
C5 |
C1 |
H6 |
116.235 |
| C5 |
C4 |
H10 |
118.262 |
|
C5 |
C4 |
H11 |
117.877 |
| H8 |
C3 |
H9 |
117.363 |
|
H10 |
C4 |
H11 |
114.897 |
| H12 |
C5 |
H13 |
114.897 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
