Jump to
S1C2
Energy calculated at QCISD(T)/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -79.680126 |
| Energy at 298.15K | -79.686036 |
| HF Energy | -79.260157 |
| Nuclear repulsion energy | 42.196122 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
3032 |
2931 |
|
|
|
|
| 2 |
A1g |
1428 |
1381 |
|
|
|
|
| 3 |
A1g |
1010 |
977 |
|
|
|
|
| 4 |
A1u |
307 |
297 |
|
|
|
|
| 5 |
A2u |
3031 |
2930 |
|
|
|
|
| 6 |
A2u |
1412 |
1365 |
|
|
|
|
| 7 |
Eg |
3089 |
2987 |
|
|
|
|
| 7 |
Eg |
3089 |
2987 |
|
|
|
|
| 8 |
Eg |
1514 |
1464 |
|
|
|
|
| 8 |
Eg |
1514 |
1463 |
|
|
|
|
| 9 |
Eg |
1226 |
1185 |
|
|
|
|
| 9 |
Eg |
1226 |
1185 |
|
|
|
|
| 10 |
Eu |
3113 |
3009 |
|
|
|
|
| 10 |
Eu |
3113 |
3009 |
|
|
|
|
| 11 |
Eu |
1515 |
1465 |
|
|
|
|
| 11 |
Eu |
1515 |
1465 |
|
|
|
|
| 12 |
Eu |
823 |
795 |
|
|
|
|
| 12 |
Eu |
822 |
795 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16389.1 cm
-1
Scaled (by 0.9668) Zero Point Vibrational Energy (zpe) 15845.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/aug-cc-pVTZ
Point Group is D3d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.765 |
| C2 |
0.000 |
0.000 |
-0.765 |
| H3 |
0.000 |
1.019 |
1.160 |
| H4 |
-0.883 |
-0.510 |
1.160 |
| H5 |
0.883 |
-0.510 |
1.160 |
| H6 |
0.000 |
-1.019 |
-1.160 |
| H7 |
-0.883 |
0.510 |
-1.160 |
| H8 |
0.883 |
0.510 |
-1.160 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5303 | 1.0928 | 1.0928 | 1.0928 | 2.1781 | 2.1781 | 2.1781 |
C2 | 1.5303 | | 2.1781 | 2.1781 | 2.1781 | 1.0928 | 1.0928 | 1.0928 | H3 | 1.0928 | 2.1781 | | 1.7650 | 1.7650 | 3.0879 | 2.5337 | 2.5337 | H4 | 1.0928 | 2.1781 | 1.7650 | | 1.7650 | 2.5337 | 2.5337 | 3.0879 | H5 | 1.0928 | 2.1781 | 1.7650 | 1.7650 | | 2.5337 | 3.0879 | 2.5337 | H6 | 2.1781 | 1.0928 | 3.0879 | 2.5337 | 2.5337 | | 1.7650 | 1.7650 | H7 | 2.1781 | 1.0928 | 2.5337 | 2.5337 | 3.0879 | 1.7650 | | 1.7650 | H8 | 2.1781 | 1.0928 | 2.5337 | 3.0879 | 2.5337 | 1.7650 | 1.7650 | |
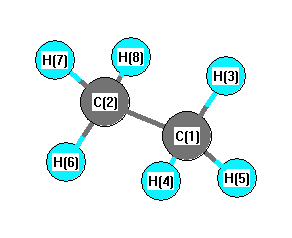 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.176 |
|
C1 |
C2 |
H7 |
111.176 |
| C1 |
C2 |
H8 |
111.176 |
|
C2 |
C1 |
H3 |
111.176 |
| C2 |
C1 |
H4 |
111.176 |
|
C2 |
C1 |
H5 |
111.176 |
| H3 |
C1 |
H4 |
107.714 |
|
H3 |
C1 |
H5 |
107.714 |
| H4 |
C1 |
H5 |
107.714 |
|
H6 |
C2 |
H7 |
107.714 |
| H6 |
C2 |
H8 |
107.714 |
|
H7 |
C2 |
H8 |
107.714 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -79.675704 |
| Energy at 298.15K | |
| HF Energy | -79.255309 |
| Nuclear repulsion energy | 42.031183 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
3045 |
2944 |
|
|
|
|
| 2 |
A1' |
1443 |
1395 |
|
|
|
|
| 3 |
A1' |
1007 |
973 |
|
|
|
|
| 4 |
A1" |
310i |
300i |
|
|
|
|
| 5 |
A2" |
3037 |
2936 |
|
|
|
|
| 6 |
A2" |
1405 |
1358 |
|
|
|
|
| 7 |
E' |
3119 |
3015 |
|
|
|
|
| 7 |
E' |
3119 |
3015 |
|
|
|
|
| 8 |
E' |
1521 |
1471 |
|
|
|
|
| 8 |
E' |
1521 |
1471 |
|
|
|
|
| 9 |
E' |
897 |
867 |
|
|
|
|
| 9 |
E' |
897 |
867 |
|
|
|
|
| 10 |
E" |
3097 |
2994 |
|
|
|
|
| 10 |
E" |
3097 |
2994 |
|
|
|
|
| 11 |
E" |
1511 |
1461 |
|
|
|
|
| 11 |
E" |
1510 |
1460 |
|
|
|
|
| 12 |
E" |
1175 |
1136 |
|
|
|
|
| 12 |
E" |
1175 |
1136 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16132.4 cm
-1
Scaled (by 0.9668) Zero Point Vibrational Energy (zpe) 15596.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/aug-cc-pVTZ
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.772 |
| C2 |
0.000 |
0.000 |
-0.772 |
| H3 |
0.000 |
1.015 |
1.174 |
| H4 |
-0.879 |
-0.508 |
1.174 |
| H5 |
0.879 |
-0.508 |
1.174 |
| H6 |
0.000 |
1.015 |
-1.174 |
| H7 |
0.879 |
-0.508 |
-1.174 |
| H8 |
-0.879 |
-0.508 |
-1.174 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5442 | 1.0918 | 1.0918 | 1.0918 | 2.1950 | 2.1950 | 2.1950 |
C2 | 1.5442 | | 2.1950 | 2.1950 | 2.1950 | 1.0918 | 1.0918 | 1.0918 | H3 | 1.0918 | 2.1950 | | 1.7582 | 1.7582 | 2.3481 | 2.9334 | 2.9334 | H4 | 1.0918 | 2.1950 | 1.7582 | | 1.7582 | 2.9334 | 2.9334 | 2.3481 | H5 | 1.0918 | 2.1950 | 1.7582 | 1.7582 | | 2.9334 | 2.3481 | 2.9334 | H6 | 2.1950 | 1.0918 | 2.3481 | 2.9334 | 2.9334 | | 1.7582 | 1.7582 | H7 | 2.1950 | 1.0918 | 2.9334 | 2.9334 | 2.3481 | 1.7582 | | 1.7582 | H8 | 2.1950 | 1.0918 | 2.9334 | 2.3481 | 2.9334 | 1.7582 | 1.7582 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.601 |
|
C1 |
C2 |
H7 |
111.601 |
| C1 |
C2 |
H8 |
111.601 |
|
C2 |
C1 |
H3 |
111.601 |
| C2 |
C1 |
H4 |
111.601 |
|
C2 |
C1 |
H5 |
111.601 |
| H3 |
C1 |
H4 |
107.260 |
|
H3 |
C1 |
H5 |
107.260 |
| H4 |
C1 |
H5 |
107.260 |
|
H6 |
C2 |
H7 |
107.260 |
| H6 |
C2 |
H8 |
107.260 |
|
H7 |
C2 |
H8 |
107.260 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
