Jump to
S1C2
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -79.664287 |
| Energy at 298.15K | -79.670217 |
| HF Energy | -79.260307 |
| Nuclear repulsion energy | 42.279610 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1g |
3058 |
2927 |
0.00 |
|
|
|
| 2 |
A1g |
1448 |
1386 |
0.00 |
|
|
|
| 3 |
A1g |
1023 |
979 |
0.00 |
|
|
|
| 4 |
A1u |
311 |
297 |
0.00 |
|
|
|
| 5 |
A2u |
3058 |
2927 |
50.90 |
|
|
|
| 6 |
A2u |
1430 |
1368 |
0.83 |
|
|
|
| 7 |
Eg |
3119 |
2984 |
0.00 |
|
|
|
| 7 |
Eg |
3119 |
2984 |
0.00 |
|
|
|
| 8 |
Eg |
1529 |
1464 |
0.00 |
|
|
|
| 8 |
Eg |
1529 |
1464 |
0.00 |
|
|
|
| 9 |
Eg |
1241 |
1188 |
0.00 |
|
|
|
| 9 |
Eg |
1241 |
1188 |
0.00 |
|
|
|
| 10 |
Eu |
3141 |
3006 |
56.54 |
|
|
|
| 10 |
Eu |
3141 |
3006 |
56.54 |
|
|
|
| 11 |
Eu |
1531 |
1465 |
7.98 |
|
|
|
| 11 |
Eu |
1531 |
1465 |
7.98 |
|
|
|
| 12 |
Eu |
832 |
797 |
2.69 |
|
|
|
| 12 |
Eu |
832 |
797 |
2.68 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16556.6 cm
-1
Scaled (by 0.957) Zero Point Vibrational Energy (zpe) 15844.7 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is D3d
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.764 |
| C2 |
0.000 |
0.000 |
-0.764 |
| H3 |
0.000 |
1.017 |
1.158 |
| H4 |
-0.881 |
-0.508 |
1.158 |
| H5 |
0.881 |
-0.508 |
1.158 |
| H6 |
0.000 |
-1.017 |
-1.158 |
| H7 |
-0.881 |
0.508 |
-1.158 |
| H8 |
0.881 |
0.508 |
-1.158 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5270 | 1.0906 | 1.0906 | 1.0906 | 2.1738 | 2.1738 | 2.1738 |
C2 | 1.5270 | | 2.1738 | 2.1738 | 2.1738 | 1.0906 | 1.0906 | 1.0906 | H3 | 1.0906 | 2.1738 | | 1.7613 | 1.7613 | 3.0819 | 2.5290 | 2.5290 | H4 | 1.0906 | 2.1738 | 1.7613 | | 1.7613 | 2.5290 | 2.5290 | 3.0819 | H5 | 1.0906 | 2.1738 | 1.7613 | 1.7613 | | 2.5290 | 3.0819 | 2.5290 | H6 | 2.1738 | 1.0906 | 3.0819 | 2.5290 | 2.5290 | | 1.7613 | 1.7613 | H7 | 2.1738 | 1.0906 | 2.5290 | 2.5290 | 3.0819 | 1.7613 | | 1.7613 | H8 | 2.1738 | 1.0906 | 2.5290 | 3.0819 | 2.5290 | 1.7613 | 1.7613 | |
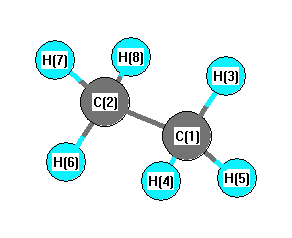 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.192 |
|
C1 |
C2 |
H7 |
111.192 |
| C1 |
C2 |
H8 |
111.192 |
|
C2 |
C1 |
H3 |
111.192 |
| C2 |
C1 |
H4 |
111.192 |
|
C2 |
C1 |
H5 |
111.192 |
| H3 |
C1 |
H4 |
107.697 |
|
H3 |
C1 |
H5 |
107.697 |
| H4 |
C1 |
H5 |
107.697 |
|
H6 |
C2 |
H7 |
107.697 |
| H6 |
C2 |
H8 |
107.697 |
|
H7 |
C2 |
H8 |
107.697 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -79.659828 |
| Energy at 298.15K | |
| HF Energy | -79.255461 |
| Nuclear repulsion energy | 42.116162 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A1' |
3072 |
2940 |
0.00 |
|
|
|
| 2 |
A1' |
1462 |
1399 |
0.00 |
|
|
|
| 3 |
A1' |
1020 |
977 |
0.00 |
|
|
|
| 4 |
A1" |
308i |
294i |
0.00 |
|
|
|
| 5 |
A2" |
3065 |
2933 |
53.17 |
|
|
|
| 6 |
A2" |
1423 |
1362 |
0.45 |
|
|
|
| 7 |
E' |
3147 |
3012 |
48.75 |
|
|
|
| 7 |
E' |
3147 |
3012 |
48.76 |
|
|
|
| 8 |
E' |
1537 |
1471 |
9.43 |
|
|
|
| 8 |
E' |
1537 |
1471 |
9.43 |
|
|
|
| 9 |
E' |
907 |
868 |
2.01 |
|
|
|
| 9 |
E' |
907 |
868 |
2.01 |
|
|
|
| 10 |
E" |
3126 |
2992 |
0.00 |
|
|
|
| 10 |
E" |
3126 |
2992 |
0.00 |
|
|
|
| 11 |
E" |
1526 |
1460 |
0.00 |
|
|
|
| 11 |
E" |
1526 |
1460 |
0.00 |
|
|
|
| 12 |
E" |
1190 |
1139 |
0.00 |
|
|
|
| 12 |
E" |
1190 |
1139 |
0.00 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 16300.9 cm
-1
Scaled (by 0.957) Zero Point Vibrational Energy (zpe) 15599.9 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is D3h
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.000 |
0.770 |
| C2 |
0.000 |
0.000 |
-0.770 |
| H3 |
0.000 |
1.013 |
1.172 |
| H4 |
-0.877 |
-0.506 |
1.172 |
| H5 |
0.877 |
-0.506 |
1.172 |
| H6 |
0.000 |
1.013 |
-1.172 |
| H7 |
0.877 |
-0.506 |
-1.172 |
| H8 |
-0.877 |
-0.506 |
-1.172 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
H3 |
H4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5409 | 1.0895 | 1.0895 | 1.0895 | 2.1906 | 2.1906 | 2.1906 |
C2 | 1.5409 | | 2.1906 | 2.1906 | 2.1906 | 1.0895 | 1.0895 | 1.0895 | H3 | 1.0895 | 2.1906 | | 1.7544 | 1.7544 | 2.3437 | 2.9276 | 2.9276 | H4 | 1.0895 | 2.1906 | 1.7544 | | 1.7544 | 2.9276 | 2.9276 | 2.3437 | H5 | 1.0895 | 2.1906 | 1.7544 | 1.7544 | | 2.9276 | 2.3437 | 2.9276 | H6 | 2.1906 | 1.0895 | 2.3437 | 2.9276 | 2.9276 | | 1.7544 | 1.7544 | H7 | 2.1906 | 1.0895 | 2.9276 | 2.9276 | 2.3437 | 1.7544 | | 1.7544 | H8 | 2.1906 | 1.0895 | 2.9276 | 2.3437 | 2.9276 | 1.7544 | 1.7544 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H6 |
111.618 |
|
C1 |
C2 |
H7 |
111.618 |
| C1 |
C2 |
H8 |
111.618 |
|
C2 |
C1 |
H3 |
111.618 |
| C2 |
C1 |
H4 |
111.618 |
|
C2 |
C1 |
H5 |
111.618 |
| H3 |
C1 |
H4 |
107.241 |
|
H3 |
C1 |
H5 |
107.241 |
| H4 |
C1 |
H5 |
107.241 |
|
H6 |
C2 |
H7 |
107.241 |
| H6 |
C2 |
H8 |
107.241 |
|
H7 |
C2 |
H8 |
107.241 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
