All results from a given calculation for CH3CHO (Acetaldehyde)
using model chemistry: QCISD(T)/aug-cc-pVTZ
19 10 17 12 22
States and conformations
| State |
Conformation |
minimum conformation |
conformer description |
state description |
| 1 |
1 |
yes |
CS |
1A' |
Energy calculated at QCISD(T)/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -153.599240 |
| Energy at 298.15K | |
| HF Energy | -152.976245 |
| Nuclear repulsion energy | 69.538396 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVTZ
Geometric Data calculated at QCISD(T)/aug-cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.231 |
0.403 |
-0.000 |
| C2 |
-1.170 |
-0.150 |
-0.000 |
| O3 |
1.236 |
-0.278 |
0.000 |
| H4 |
0.308 |
1.508 |
-0.000 |
| H5 |
-1.151 |
-1.239 |
-0.001 |
| H6 |
-1.704 |
0.219 |
0.881 |
| H7 |
-1.705 |
0.220 |
-0.880 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
O3 |
H4 |
H5 |
H6 |
H7 |
| C1 | | 1.5053 | 1.2141 | 1.1079 | 2.1456 | 2.1337 | 2.1337 |
C2 | 1.5053 | | 2.4091 | 2.2206 | 1.0892 | 1.0942 | 1.0942 | O3 | 1.2141 | 2.4091 | | 2.0125 | 2.5733 | 3.1093 | 3.1097 | H4 | 1.1079 | 2.2206 | 2.0125 | | 3.1101 | 2.5466 | 2.5462 | H5 | 2.1456 | 1.0892 | 2.5733 | 3.1101 | | 1.7911 | 1.7911 | H6 | 2.1337 | 1.0942 | 3.1093 | 2.5466 | 1.7911 | | 1.7606 | H7 | 2.1337 | 1.0942 | 3.1097 | 2.5462 | 1.7911 | 1.7606 | |
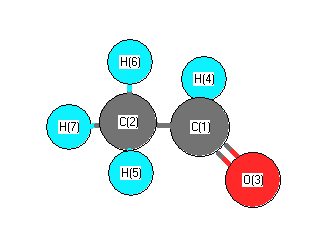 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
H5 |
110.554 |
|
C1 |
C2 |
H6 |
109.313 |
| C1 |
C2 |
H7 |
109.313 |
|
C2 |
C1 |
O3 |
124.381 |
| C2 |
C1 |
H4 |
115.532 |
|
O3 |
C1 |
H4 |
120.087 |
| H5 |
C2 |
H6 |
110.233 |
|
H5 |
C2 |
H7 |
110.233 |
| H6 |
C2 |
H7 |
107.123 |
|
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
