Jump to
S1C2
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -169.624237 |
| Energy at 298.15K | -169.627608 |
| HF Energy | -169.005636 |
| Nuclear repulsion energy | 71.478674 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3789 |
3626 |
55.17 |
|
|
|
| 2 |
A' |
3648 |
3491 |
53.13 |
|
|
|
| 3 |
A' |
3026 |
2896 |
68.36 |
|
|
|
| 4 |
A' |
1845 |
1766 |
461.67 |
|
|
|
| 5 |
A' |
1648 |
1577 |
64.60 |
|
|
|
| 6 |
A' |
1446 |
1384 |
4.54 |
|
|
|
| 7 |
A' |
1292 |
1236 |
134.57 |
|
|
|
| 8 |
A' |
1074 |
1028 |
7.11 |
|
|
|
| 9 |
A' |
576 |
551 |
11.99 |
|
|
|
| 10 |
A" |
1076 |
1029 |
2.49 |
|
|
|
| 11 |
A" |
632 |
605 |
13.17 |
|
|
|
| 12 |
A" |
23 |
22 |
219.09 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10036.5 cm
-1
Scaled (by 0.957) Zero Point Vibrational Energy (zpe) 9605.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.419 |
0.000 |
| O2 |
1.193 |
0.234 |
0.000 |
| N3 |
-0.935 |
-0.563 |
0.000 |
| H4 |
-0.443 |
1.425 |
0.000 |
| H5 |
-0.641 |
-1.523 |
0.000 |
| H6 |
-1.912 |
-0.347 |
0.000 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2068 | 1.3562 | 1.0996 | 2.0450 | 2.0593 |
O2 | 1.2068 | | 2.2721 | 2.0234 | 2.5399 | 3.1580 | N3 | 1.3562 | 2.2721 | | 2.0487 | 1.0032 | 1.0004 | H4 | 1.0996 | 2.0234 | 2.0487 | | 2.9548 | 2.3015 | H5 | 2.0450 | 2.5399 | 1.0032 | 2.9548 | | 1.7309 | H6 | 2.0593 | 3.1580 | 1.0004 | 2.3015 | 1.7309 | |
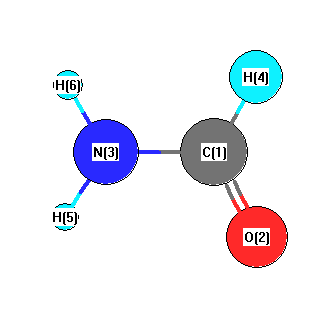 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.417 |
|
C1 |
N3 |
H6 |
121.073 |
| O2 |
C1 |
N3 |
124.772 |
|
O2 |
C1 |
H4 |
122.572 |
| N3 |
C1 |
H4 |
112.656 |
|
H5 |
N3 |
H6 |
119.511 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at CCD/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -169.624237 |
| Energy at 298.15K | -169.627606 |
| HF Energy | -169.005636 |
| Nuclear repulsion energy | 71.480331 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A |
3789 |
3626 |
55.18 |
|
|
|
| 2 |
A |
3648 |
3491 |
53.16 |
|
|
|
| 3 |
A |
3026 |
2896 |
68.31 |
|
|
|
| 4 |
A |
1845 |
1765 |
461.77 |
|
|
|
| 5 |
A |
1648 |
1577 |
64.57 |
|
|
|
| 6 |
A |
1446 |
1384 |
4.53 |
|
|
|
| 7 |
A |
1292 |
1237 |
134.50 |
|
|
|
| 8 |
A |
1075 |
1029 |
2.54 |
|
|
|
| 9 |
A |
1074 |
1028 |
7.01 |
|
|
|
| 10 |
A |
632 |
605 |
13.23 |
|
|
|
| 11 |
A |
576 |
551 |
11.98 |
|
|
|
| 12 |
A |
22 |
21 |
219.08 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 10035.9 cm
-1
Scaled (by 0.957) Zero Point Vibrational Energy (zpe) 9604.4 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at CCD/aug-cc-pVTZ
Point Group is C1
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.162 |
0.386 |
0.000 |
| O2 |
1.190 |
-0.246 |
0.000 |
| N3 |
-1.080 |
-0.157 |
-0.001 |
| H4 |
0.144 |
1.486 |
0.000 |
| H5 |
-1.181 |
-1.155 |
0.001 |
| H6 |
-1.897 |
0.421 |
0.002 |
Atom - Atom Distances (Å)
| |
C1 |
O2 |
N3 |
H4 |
H5 |
H6 |
| C1 | | 1.2068 | 1.3561 | 1.0996 | 2.0448 | 2.0593 |
O2 | 1.2068 | | 2.2719 | 2.0234 | 2.5393 | 3.1580 | N3 | 1.3561 | 2.2719 | | 2.0489 | 1.0032 | 1.0004 | H4 | 1.0996 | 2.0234 | 2.0489 | | 2.9548 | 2.3019 | H5 | 2.0448 | 2.5393 | 1.0032 | 2.9548 | | 1.7309 | H6 | 2.0593 | 3.1580 | 1.0004 | 2.3019 | 1.7309 | |
More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
N3 |
H5 |
119.400 |
|
C1 |
N3 |
H6 |
121.081 |
| O2 |
C1 |
N3 |
124.758 |
|
O2 |
C1 |
H4 |
122.565 |
| N3 |
C1 |
H4 |
112.677 |
|
H5 |
N3 |
H6 |
119.518 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
