return to home page Computational Chemistry Comparison and Benchmark DataBase Release 22 (May 2022) Standard Reference Database 101 National Institute of Standards and Technology
You are here: Calculated > Energy > Optimized > Energy

All results from a given calculation for CH3CHClCH3 (Propane, 2-chloro-)

using model chemistry: CCD/aug-cc-pVTZ

19 10 17 12 22

States and conformations

State Conformation minimum conformation conformer description state description
1 1 yes CS 1A'
Energy calculated at CCD/aug-cc-pVTZ
 hartrees
Energy at 0K-578.032890
Energy at 298.15K-578.040665
HF Energy-577.251243
Nuclear repulsion energy166.371659
The energy at 298.15K was derived from the energy at 0K and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at CCD/aug-cc-pVTZ
Mode Number Symmetry Frequency
(cm-1)
Scaled Frequency
(cm-1)
IR Intensities
(km mol-1)
Raman Act
(Å4/u)
Dep P Dep U
1 A' 3159 3023 21.26      
2 A' 3140 3005 33.08      
3 A' 3117 2983 0.27      
4 A' 3060 2929 18.84      
5 A' 1532 1466 6.53      
6 A' 1518 1453 8.25      
7 A' 1445 1383 5.33      
8 A' 1319 1262 32.15      
9 A' 1211 1159 13.41      
10 A' 1105 1057 11.95      
11 A' 922 882 8.92      
12 A' 659 631 22.92      
13 A' 426 408 1.48      
14 A' 340 325 1.05      
15 A' 274 262 0.09      
16 A" 3154 3019 11.61      
17 A" 3133 2998 1.02      
18 A" 3057 2926 10.96      
19 A" 1512 1447 0.23      
20 A" 1507 1442 2.21      
21 A" 1433 1372 9.98      
22 A" 1386 1326 0.55      
23 A" 1174 1123 1.76      
24 A" 975 933 0.03      
25 A" 955 914 0.84      
26 A" 328 314 1.42      
27 A" 243 233 0.02      

Unscaled Zero Point Vibrational Energy (zpe) 21040.7 cm-1
Scaled (by 0.957) Zero Point Vibrational Energy (zpe) 20135.9 cm-1
See section III.C.1 List or set vibrational scaling factors to change the scale factors used here.
See section III.C.2 Calculate a vibrational scaling factor for a given set of molecules to determine the least squares best scaling factor.
Rotational Constants (cm-1) from geometry optimized at CCD/aug-cc-pVTZ
ABC
0.27056 0.15255 0.10726

See section I.F.4 to change rotational constant units
Geometric Data calculated at CCD/aug-cc-pVTZ

Point Group is Cs

Cartesians (Å)
Atom x (Å) y (Å) z (Å)
Cl1 -0.887 1.012 0.000
C2 0.569 -0.061 0.000
C3 0.569 -0.898 1.265
C4 0.569 -0.898 -1.265
H5 1.413 0.625 0.000
H6 1.465 -1.521 1.290
H7 1.465 -1.521 -1.290
H8 0.557 -0.268 2.153
H9 0.557 -0.268 -2.153
H10 -0.306 -1.548 1.289
H11 -0.306 -1.548 -1.289

Atom - Atom Distances (Å)
  Cl1 C2 C3 C4 H5 H6 H7 H8 H9 H10 H11
Cl11.80822.71442.71442.33223.68963.68962.89072.89072.92452.9245
C21.80821.51731.51731.08792.14492.14492.16282.16282.15382.1538
C32.71441.51732.53092.15271.09212.77931.08863.47601.09022.7774
C42.71441.51732.53092.15272.77931.09213.47601.08862.77741.0902
H52.33221.08792.15272.15272.50442.50442.48282.48283.05623.0562
H63.68962.14491.09212.77932.50442.58081.77203.77521.77203.1295
H73.68962.14492.77931.09212.50442.58083.77521.77203.12951.7720
H82.89072.16281.08863.47602.48281.77203.77524.30571.76943.7725
H92.89072.16283.47601.08862.48283.77521.77204.30573.77251.7694
H102.92452.15381.09022.77743.05621.77203.12951.76943.77252.5783
H112.92452.15382.77741.09023.05623.12951.77203.77251.76942.5783

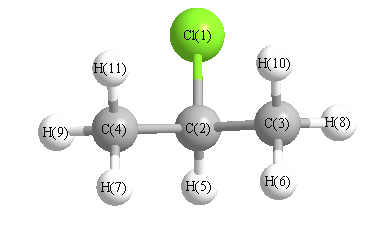
picture of Propane, 2-chloro- state 1 conformation 1
More geometry information
Calculated Bond Angles
atom1 atom2 atom3 angle atom1 atom2 atom3 angle
Cl1 C2 C3 109.109 Cl1 C2 C4 109.109
Cl1 C3 H5 55.835 C2 C3 H6 109.487
C2 C3 H8 111.116 C2 C3 H10 110.308
C2 C4 H7 109.487 C2 C4 H9 111.116
C2 C4 H11 110.308 C3 C2 C4 113.029
C3 C2 H5 110.357 C4 C2 H5 110.357
H6 C3 H8 108.698 H6 C3 H10 108.578
H7 C4 H9 108.698 H7 C4 H11 108.578
H8 C3 H10 108.598 H9 C4 H11 108.598
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability