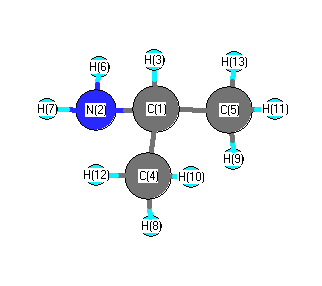
Vibrational Frequencies calculated at B3PW91/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3507 |
3385 |
0.99 |
|
|
|
| 2 |
A' |
3114 |
3005 |
50.10 |
|
|
|
| 3 |
A' |
3098 |
2990 |
44.15 |
|
|
|
| 4 |
A' |
3031 |
2926 |
5.58 |
|
|
|
| 5 |
A' |
2931 |
2829 |
67.67 |
|
|
|
| 6 |
A' |
1649 |
1592 |
33.24 |
|
|
|
| 7 |
A' |
1500 |
1447 |
7.16 |
|
|
|
| 8 |
A' |
1487 |
1435 |
6.42 |
|
|
|
| 9 |
A' |
1407 |
1358 |
14.97 |
|
|
|
| 10 |
A' |
1367 |
1319 |
21.43 |
|
|
|
| 11 |
A' |
1200 |
1158 |
8.89 |
|
|
|
| 12 |
A' |
1157 |
1116 |
13.17 |
|
|
|
| 13 |
A' |
994 |
959 |
3.94 |
|
|
|
| 14 |
A' |
831 |
802 |
70.50 |
|
|
|
| 15 |
A' |
821 |
792 |
46.58 |
|
|
|
| 16 |
A' |
470 |
454 |
11.26 |
|
|
|
| 17 |
A' |
358 |
345 |
0.12 |
|
|
|
| 18 |
A' |
257 |
248 |
0.26 |
|
|
|
| 19 |
A" |
3589 |
3463 |
0.43 |
|
|
|
| 20 |
A" |
3109 |
3000 |
0.06 |
|
|
|
| 21 |
A" |
3096 |
2988 |
18.53 |
|
|
|
| 22 |
A" |
3028 |
2922 |
41.18 |
|
|
|
| 23 |
A" |
1481 |
1429 |
0.27 |
|
|
|
| 24 |
A" |
1476 |
1425 |
0.45 |
|
|
|
| 25 |
A" |
1408 |
1359 |
17.05 |
|
|
|
| 26 |
A" |
1380 |
1332 |
0.64 |
|
|
|
| 27 |
A" |
1263 |
1219 |
0.28 |
|
|
|
| 28 |
A" |
1036 |
1000 |
1.17 |
|
|
|
| 29 |
A" |
961 |
927 |
0.32 |
|
|
|
| 30 |
A" |
926 |
893 |
0.35 |
|
|
|
| 31 |
A" |
402 |
388 |
6.32 |
|
|
|
| 32 |
A" |
272 |
263 |
27.23 |
|
|
|
| 33 |
A" |
209 |
202 |
7.13 |
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 26405.6 cm
-1
Scaled (by 0.9651) Zero Point Vibrational Energy (zpe) 25484.0 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Charges, Dipole, Quadrupole and Polarizability
Charges from optimized geometry at B3PW91/aug-cc-pVTZ
Charges (e)
| Number |
Element |
Mulliken |
CHELPG |
AIM |
ESP |
| 1 |
C |
0.700 |
|
|
|
| 2 |
N |
-0.523 |
|
|
|
| 3 |
H |
0.155 |
|
|
|
| 4 |
C |
-0.809 |
|
|
|
| 5 |
C |
-0.809 |
|
|
|
| 6 |
H |
0.037 |
|
|
|
| 7 |
H |
0.037 |
|
|
|
| 8 |
H |
0.218 |
|
|
|
| 9 |
H |
0.218 |
|
|
|
| 10 |
H |
0.207 |
|
|
|
| 11 |
H |
0.207 |
|
|
|
| 12 |
H |
0.182 |
|
|
|
| 13 |
H |
0.182 |
|
|
|
Electric dipole moments
Electric dipole components in Debye
(What's a Debye? See section
VII.A.3)
| |
x |
y |
z |
Total |
| |
0.987 |
0.769 |
0.000 |
1.251 |
| CHELPG |
|
|
|
|
| AIM |
|
|
|
|
| ESP |
|
|
|
|
Electric Quadrupole moment
Quadrupole components in D Å
| Primitive |
|---|
| | x | y | z |
|---|
| x |
-30.126 |
-0.548 |
0.000 |
| y |
-0.548 |
-25.134 |
0.000 |
| z |
0.000 |
0.000 |
-26.537 |
|
| Traceless |
|---|
| | x | y | z |
|---|
| x |
-4.290 |
-0.548 |
0.000 |
| y |
-0.548 |
3.197 |
0.000 |
| z |
0.000 |
0.000 |
1.093 |
|
| Polar |
|---|
| 3z2-r2 | 2.187 |
|---|
| x2-y2 | -4.991 |
|---|
| xy | -0.548 |
|---|
| xz | 0.000 |
|---|
| yz | 0.000 |
|---|
|
Polarizabilities
Components of the polarizability tensor.
Units are
Å
3 (Angstrom cubed)
Change units.
| |
x |
y |
z |
| x |
7.221 |
-0.389 |
0.000 |
| y |
-0.389 |
7.482 |
0.000 |
| z |
0.000 |
0.000 |
7.802 |
<r2> (average value of r
2) Å
2
| <r2> |
93.127 |
| (<r2>)1/2 |
9.650 |