Jump to
S1C2
Energy calculated at QCISD(T)/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -637.958376 |
| Energy at 298.15K | -637.963262 |
| HF Energy | -637.088346 |
| Nuclear repulsion energy | 156.979922 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVTZ
| Mode Number |
Symmetry |
Frequency
(cm-1) |
Scaled Frequency
(cm-1) |
IR Intensities
(km mol-1) |
Raman Act
(Å4/u) |
Dep P |
Dep U |
|---|
| 1 |
A' |
3099 |
2996 |
|
|
|
|
| 2 |
A' |
3067 |
2966 |
|
|
|
|
| 3 |
A' |
1528 |
1477 |
|
|
|
|
| 4 |
A' |
1505 |
1455 |
|
|
|
|
| 5 |
A' |
1418 |
1371 |
|
|
|
|
| 6 |
A' |
1283 |
1240 |
|
|
|
|
| 7 |
A' |
1092 |
1056 |
|
|
|
|
| 8 |
A' |
1054 |
1019 |
|
|
|
|
| 9 |
A' |
784 |
758 |
|
|
|
|
| 10 |
A' |
384 |
372 |
|
|
|
|
| 11 |
A' |
240 |
232 |
|
|
|
|
| 12 |
A" |
3167 |
3062 |
|
|
|
|
| 13 |
A" |
3124 |
3020 |
|
|
|
|
| 14 |
A" |
1305 |
1262 |
|
|
|
|
| 15 |
A" |
1220 |
1179 |
|
|
|
|
| 16 |
A" |
1065 |
1029 |
|
|
|
|
| 17 |
A" |
794 |
768 |
|
|
|
|
| 18 |
A" |
130 |
125 |
|
|
|
|
Unscaled Zero Point Vibrational Energy (zpe) 13128.7 cm
-1
Scaled (by 0.9668) Zero Point Vibrational Energy (zpe) 12692.8 cm
-1
See section
III.C.1 List or set vibrational scaling factors
to change the scale factors used here.
See section
III.C.2
Calculate a vibrational scaling factor for a given set of molecules
to determine the least squares best scaling factor.
Geometric Data calculated at QCISD(T)/aug-cc-pVTZ
Point Group is Cs
Cartesians (Å)
| Atom |
x (Å) |
y (Å) |
z (Å) |
|---|
| C1 |
0.000 |
0.626 |
0.000 |
| C2 |
0.982 |
-0.528 |
0.000 |
| Cl3 |
-1.663 |
-0.047 |
0.000 |
| F4 |
2.270 |
0.002 |
0.000 |
| H5 |
0.110 |
1.239 |
0.892 |
| H6 |
0.110 |
1.239 |
-0.892 |
| H7 |
0.864 |
-1.143 |
0.893 |
| H8 |
0.864 |
-1.143 |
-0.893 |
Atom - Atom Distances (Å)
| |
C1 |
C2 |
Cl3 |
F4 |
H5 |
H6 |
H7 |
H8 |
| C1 | | 1.5155 | 1.7943 | 2.3547 | 1.0880 | 1.0880 | 2.1624 | 2.1624 |
C2 | 1.5155 | | 2.6888 | 1.3928 | 2.1628 | 2.1628 | 1.0914 | 1.0914 | Cl3 | 1.7943 | 2.6888 | | 3.9339 | 2.3655 | 2.3655 | 2.8958 | 2.8958 | F4 | 2.3547 | 1.3928 | 3.9339 | | 2.6442 | 2.6442 | 2.0221 | 2.0221 | H5 | 1.0880 | 2.1628 | 2.3655 | 2.6442 | | 1.7847 | 2.4985 | 3.0711 | H6 | 1.0880 | 2.1628 | 2.3655 | 2.6442 | 1.7847 | | 3.0711 | 2.4985 | H7 | 2.1624 | 1.0914 | 2.8958 | 2.0221 | 2.4985 | 3.0711 | | 1.7870 | H8 | 2.1624 | 1.0914 | 2.8958 | 2.0221 | 3.0711 | 2.4985 | 1.7870 | |
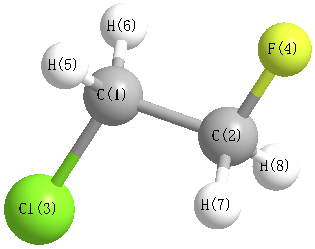 More geometry information
Calculated Bond Angles
| atom1 |
atom2 |
atom3 |
angle |
|
atom1 |
atom2 |
atom3 |
angle |
| C1 |
C2 |
F4 |
108.043 |
|
C1 |
C2 |
H7 |
111.042 |
| C1 |
C2 |
H8 |
111.042 |
|
C2 |
C1 |
Cl3 |
108.365 |
| C2 |
C1 |
H5 |
111.283 |
|
C2 |
C1 |
H6 |
111.283 |
| Cl3 |
C1 |
H5 |
107.779 |
|
Cl3 |
C1 |
H6 |
107.779 |
| F4 |
C2 |
H7 |
108.360 |
|
F4 |
C2 |
H8 |
108.360 |
| H5 |
C1 |
H6 |
110.204 |
|
H7 |
C2 |
H8 |
109.897 |
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
Jump to
S1C1
Energy calculated at QCISD(T)/aug-cc-pVTZ
| | hartrees |
|---|
| Energy at 0K | -637.957423 |
| Energy at 298.15K | |
| HF Energy | -637.087115 |
| Nuclear repulsion energy | 161.125550 |
The energy at 298.15K was derived from the energy at 0K
and an integrated heat capacity that used the calculated vibrational frequencies.
Vibrational Frequencies calculated at QCISD(T)/aug-cc-pVTZ
Geometric Data calculated at QCISD(T)/aug-cc-pVTZ
Point Group is C1
Electronic energy levels
Charges, Dipole, Quadrupole and Polarizability
